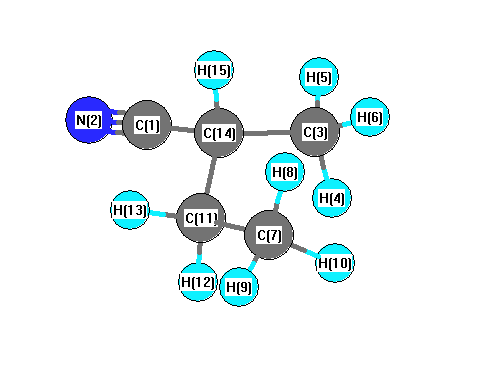
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3087 |
3036 |
15.00 |
|
|
|
| 2 |
A |
3081 |
3030 |
16.45 |
|
|
|
| 3 |
A |
3070 |
3019 |
28.65 |
|
|
|
| 4 |
A |
3061 |
3011 |
33.04 |
|
|
|
| 5 |
A |
3029 |
2979 |
5.94 |
|
|
|
| 6 |
A |
3001 |
2952 |
13.59 |
|
|
|
| 7 |
A |
2990 |
2941 |
29.36 |
|
|
|
| 8 |
A |
2986 |
2937 |
8.79 |
|
|
|
| 9 |
A |
2958 |
2909 |
4.96 |
|
|
|
| 10 |
A |
2274 |
2236 |
9.08 |
|
|
|
| 11 |
A |
1492 |
1468 |
3.24 |
|
|
|
| 12 |
A |
1485 |
1461 |
9.92 |
|
|
|
| 13 |
A |
1480 |
1456 |
6.40 |
|
|
|
| 14 |
A |
1474 |
1450 |
9.38 |
|
|
|
| 15 |
A |
1465 |
1440 |
0.21 |
|
|
|
| 16 |
A |
1393 |
1370 |
2.41 |
|
|
|
| 17 |
A |
1385 |
1362 |
5.72 |
|
|
|
| 18 |
A |
1347 |
1324 |
0.44 |
|
|
|
| 19 |
A |
1314 |
1292 |
1.34 |
|
|
|
| 20 |
A |
1287 |
1265 |
2.67 |
|
|
|
| 21 |
A |
1253 |
1233 |
1.54 |
|
|
|
| 22 |
A |
1159 |
1140 |
2.49 |
|
|
|
| 23 |
A |
1114 |
1095 |
0.17 |
|
|
|
| 24 |
A |
1092 |
1074 |
5.96 |
|
|
|
| 25 |
A |
1034 |
1017 |
1.12 |
|
|
|
| 26 |
A |
977 |
961 |
2.02 |
|
|
|
| 27 |
A |
955 |
939 |
6.24 |
|
|
|
| 28 |
A |
885 |
870 |
0.89 |
|
|
|
| 29 |
A |
796 |
783 |
0.70 |
|
|
|
| 30 |
A |
753 |
740 |
3.93 |
|
|
|
| 31 |
A |
551 |
542 |
0.14 |
|
|
|
| 32 |
A |
525 |
517 |
1.05 |
|
|
|
| 33 |
A |
387 |
381 |
0.26 |
|
|
|
| 34 |
A |
318 |
312 |
0.49 |
|
|
|
| 35 |
A |
272 |
268 |
0.12 |
|
|
|
| 36 |
A |
214 |
210 |
0.12 |
|
|
|
| 37 |
A |
199 |
196 |
2.00 |
|
|
|
| 38 |
A |
157 |
155 |
4.84 |
|
|
|
| 39 |
A |
87 |
85 |
1.38 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28191.9 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 27726.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.359 |
|
|
|
| 2 |
N |
-0.474 |
|
|
|
| 3 |
C |
-0.487 |
|
|
|
| 4 |
H |
0.179 |
|
|
|
| 5 |
H |
0.183 |
|
|
|
| 6 |
H |
0.174 |
|
|
|
| 7 |
C |
-0.494 |
|
|
|
| 8 |
H |
0.163 |
|
|
|
| 9 |
H |
0.172 |
|
|
|
| 10 |
H |
0.167 |
|
|
|
| 11 |
C |
-0.283 |
|
|
|
| 12 |
H |
0.169 |
|
|
|
| 13 |
H |
0.174 |
|
|
|
| 14 |
C |
-0.195 |
|
|
|
| 15 |
H |
0.192 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
3.777 |
1.273 |
0.470 |
4.013 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-47.768 |
-3.749 |
-0.829 |
| y |
-3.749 |
-36.973 |
-0.104 |
| z |
-0.829 |
-0.104 |
-36.430 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-11.067 |
-3.749 |
-0.829 |
| y |
-3.749 |
5.126 |
-0.104 |
| z |
-0.829 |
-0.104 |
5.940 |
|
| Polar |
|---|
| 3z2-r2 | 11.880 |
|---|
| x2-y2 | -10.795 |
|---|
| xy | -3.749 |
|---|
| xz | -0.829 |
|---|
| yz | -0.104 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.522 |
0.524 |
0.261 |
| y |
0.524 |
7.873 |
0.117 |
| z |
0.261 |
0.117 |
6.836 |
<r2> (average value of r
2) Å
2
| <r2> |
200.853 |
| (<r2>)1/2 |
14.172 |