Jump to
S1C2
S1C3
Energy calculated at MP4/6-31G*
| | hartrees |
|---|
| Energy at 0K | -1590.521973 |
| Energy at 298.15K | |
| HF Energy | -1589.975570 |
| Nuclear repulsion energy | 346.235451 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at MP4/6-31G*
Point Group is D2d
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
1.449 |
0.316 |
| S2 |
0.000 |
-1.449 |
0.316 |
| S3 |
-1.449 |
0.000 |
-0.316 |
| S4 |
1.449 |
0.000 |
-0.316 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
S4 |
| S1 | | 2.8974 | 2.1442 | 2.1442 |
S2 | 2.8974 | | 2.1442 | 2.1442 | S3 | 2.1442 | 2.1442 | | 2.8974 | S4 | 2.1442 | 2.1442 | 2.8974 | |
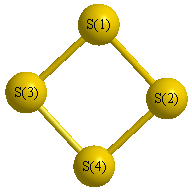 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
S4 |
47.497 |
|
S1 |
S3 |
S4 |
47.497 |
| S2 |
S1 |
S3 |
47.497 |
|
S2 |
S4 |
S3 |
47.497 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Energy calculated at MP4/6-31G*
| | hartrees |
|---|
| Energy at 0K | -1590.580103 |
| Energy at 298.15K | |
| HF Energy | -1589.904531 |
| Nuclear repulsion energy | 314.921809 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
600 |
573 |
0.00 |
|
|
|
| 2 |
A1 |
169 |
161 |
0.00 |
|
|
|
| 3 |
A1 |
88 |
84 |
0.00 |
|
|
|
| 4 |
A2 |
182 |
174 |
0.00 |
|
|
|
| 5 |
B2 |
582 |
555 |
0.00 |
|
|
|
| 6 |
B2 |
251 |
240 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 936.1 cm
-1
Scaled (by 0.9548) Zero Point Vibrational Energy (zpe) 893.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP4/6-31G*
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
1.398 |
0.977 |
| S2 |
0.000 |
-1.398 |
0.977 |
| S3 |
0.000 |
1.398 |
-0.977 |
| S4 |
0.000 |
-1.398 |
-0.977 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
S4 |
| S1 | | 2.7968 | 1.9543 | 3.4117 |
S2 | 2.7968 | | 3.4117 | 1.9543 | S3 | 1.9543 | 3.4117 | | 2.7963 | S4 | 3.4117 | 1.9543 | 2.7963 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
S4 |
89.994 |
|
S1 |
S3 |
S4 |
90.006 |
| S2 |
S1 |
S3 |
89.994 |
|
S2 |
S4 |
S3 |
90.006 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at MP4/6-31G*
| | hartrees |
|---|
| Energy at 0K | -1590.580103 |
| Energy at 298.15K | |
| HF Energy | -1589.904491 |
| Nuclear repulsion energy | 314.910150 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at MP4/6-31G*
Point Group is D2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
0.977 |
1.398 |
| S2 |
0.000 |
0.977 |
-1.398 |
| S3 |
0.000 |
-0.977 |
1.398 |
| S4 |
0.000 |
-0.977 |
-1.398 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
S4 |
| S1 | | 2.7966 | 1.9545 | 3.4119 |
S2 | 2.7966 | | 3.4119 | 1.9545 | S3 | 1.9545 | 3.4119 | | 2.7966 | S4 | 3.4119 | 1.9545 | 2.7966 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
S4 |
90.000 |
|
S1 |
S3 |
S4 |
90.000 |
| S2 |
S1 |
S3 |
90.000 |
|
S2 |
S4 |
S3 |
90.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
