return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H6 (3-Penten-1-yne, (E)-)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-193.819588
Energy at 298.15K-193.823888
HF Energy-193.607221
Nuclear repulsion energy137.888613
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3514 3335 63.88      
2 A' 3202 3039 21.21      
3 A' 3185 3023 8.75      
4 A' 3158 2998 14.09      
5 A' 3069 2913 26.70      
6 A' 2192 2081 0.42      
7 A' 1721 1633 2.36      
8 A' 1539 1461 9.84      
9 A' 1461 1387 1.44      
10 A' 1356 1287 2.46      
11 A' 1340 1272 1.56      
12 A' 1154 1095 0.67      
13 A' 1063 1009 9.73      
14 A' 931 883 5.77      
15 A' 615 583 46.25      
16 A' 543 515 0.37      
17 A' 392 372 1.39      
18 A' 172 164 0.87      
19 A" 3125 2966 19.66      
20 A" 1527 1450 6.66      
21 A" 1089 1034 0.09      
22 A" 1001 950 33.57      
23 A" 811 770 0.41      
24 A" 540 513 44.54      
25 A" 439 417 5.47      
26 A" 202 192 0.17      
27 A" 170 161 1.55      

Unscaled Zero Point Vibrational Energy (zpe) 19754.6 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 18751.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
1.29417 0.07451 0.07138

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -2.253 -0.525 0.000
H2 -2.662 -1.019 0.883
H3 -2.662 -1.019 -0.883
H4 -2.581 0.516 0.000
C5 -0.785 -0.597 0.000
H6 -0.329 -1.535 0.000
C7 0.000 0.502 0.000
H8 -0.466 1.440 0.000
C9 1.343 0.437 0.000
C10 2.551 0.393 0.000
H11 3.561 0.357 0.000

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 H6 C7 H8 C9 C10 H11
C11.09131.09131.09191.46972.17322.47602.65563.72234.89115.8807
H21.09131.76521.77292.11722.54793.19113.41304.35245.47326.4350
H31.09131.76521.77292.11722.54793.19113.41304.35245.47326.4350
H41.09191.77291.77292.11363.04692.58152.30813.92535.13436.1449
C51.46972.11722.11722.11361.04321.35082.06142.36583.47994.4497
H62.17322.54792.54793.04691.04322.06392.97792.58543.46584.3259
C72.47603.19113.19112.58151.35082.06391.04661.34462.55373.5643
H82.65563.41303.41302.30812.06142.97791.04662.06823.19384.1703
C93.72234.35244.35243.92532.36582.58541.34462.06821.20922.2198
C104.89115.47325.47325.13433.47993.46582.55373.19381.20921.0106
H115.88076.43506.43506.14494.44974.32593.56434.17032.21981.0106

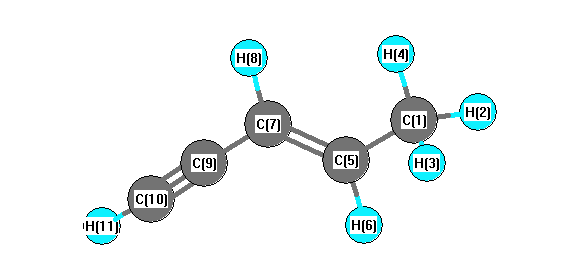
picture of 3-Penten-1-yne, (E)- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C5 H6 118.740 C1 C5 C7 122.713
H2 C1 H3 107.946 H2 C1 H4 108.596
H2 C1 C5 110.652 H3 C1 H4 108.596
H3 C1 C5 110.652 H4 C1 C5 110.324
C5 C7 H8 118.037 C5 C7 C9 122.734
H6 C5 C7 118.548 C7 C9 C10 179.312
H8 C7 C9 119.229 C9 C10 H11 179.920
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.505      
2 H 0.171      
3 H 0.171      
4 H 0.167      
5 C -0.092      
6 H 0.169      
7 C -0.234      
8 H 0.177      
9 C 0.228      
10 C -0.486      
11 H 0.234      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 12.848 1.507 0.000
y 1.507 5.993 0.000
z 0.000 0.000 3.660


<r2> (average value of r2) Å2
<r2> 159.214
(<r2>)1/2 12.618