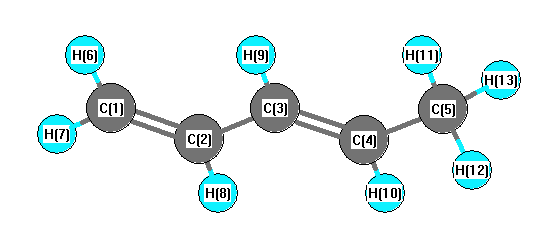
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3259 |
3118 |
15.85 |
|
|
|
| 2 |
A |
3173 |
3036 |
4.55 |
|
|
|
| 3 |
A |
3163 |
3026 |
56.54 |
|
|
|
| 4 |
A |
3156 |
3019 |
11.62 |
|
|
|
| 5 |
A |
3152 |
3015 |
0.95 |
|
|
|
| 6 |
A |
3133 |
2998 |
6.75 |
|
|
|
| 7 |
A |
3038 |
2907 |
37.24 |
|
|
|
| 8 |
A |
1749 |
1673 |
14.49 |
|
|
|
| 9 |
A |
1696 |
1622 |
4.11 |
|
|
|
| 10 |
A |
1515 |
1449 |
8.86 |
|
|
|
| 11 |
A |
1469 |
1406 |
3.72 |
|
|
|
| 12 |
A |
1434 |
1372 |
3.21 |
|
|
|
| 13 |
A |
1347 |
1288 |
2.50 |
|
|
|
| 14 |
A |
1331 |
1274 |
0.29 |
|
|
|
| 15 |
A |
1305 |
1249 |
2.30 |
|
|
|
| 16 |
A |
1224 |
1171 |
1.17 |
|
|
|
| 17 |
A |
1117 |
1068 |
0.08 |
|
|
|
| 18 |
A |
998 |
954 |
11.14 |
|
|
|
| 19 |
A |
917 |
878 |
6.50 |
|
|
|
| 20 |
A |
484 |
463 |
0.00 |
|
|
|
| 21 |
A |
451 |
431 |
0.39 |
|
|
|
| 22 |
A |
195 |
186 |
1.72 |
|
|
|
| 23 |
A |
3092 |
2958 |
21.81 |
|
|
|
| 24 |
A |
1501 |
1436 |
7.52 |
|
|
|
| 25 |
A |
1069 |
1023 |
0.07 |
|
|
|
| 26 |
A |
1052 |
1007 |
38.12 |
|
|
|
| 27 |
A |
987 |
944 |
7.69 |
|
|
|
| 28 |
A |
919 |
879 |
41.14 |
|
|
|
| 29 |
A |
845 |
809 |
7.04 |
|
|
|
| 30 |
A |
646 |
618 |
1.27 |
|
|
|
| 31 |
A |
260 |
248 |
2.16 |
|
|
|
| 32 |
A |
211 |
202 |
0.06 |
|
|
|
| 33 |
A |
137 |
131 |
1.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25010.7 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 23927.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.408 |
|
|
-0.400 |
| 2 |
C |
-0.093 |
|
|
-0.074 |
| 3 |
C |
-0.132 |
|
|
-0.240 |
| 4 |
C |
-0.136 |
|
|
-0.017 |
| 5 |
C |
-0.559 |
|
|
-0.296 |
| 6 |
H |
0.163 |
|
|
0.167 |
| 7 |
H |
0.166 |
|
|
0.164 |
| 8 |
H |
0.154 |
|
|
0.130 |
| 9 |
H |
0.148 |
|
|
0.178 |
| 10 |
H |
0.154 |
|
|
0.099 |
| 11 |
H |
0.179 |
|
|
0.091 |
| 12 |
H |
0.182 |
|
|
0.099 |
| 13 |
H |
0.182 |
|
|
0.099 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.560 |
-0.208 |
0.000 |
0.597 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
0.567 |
-0.180 |
0.000 |
0.595 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-28.772 |
-0.177 |
0.000 |
| y |
-0.177 |
-28.792 |
0.000 |
| z |
0.000 |
0.000 |
-34.215 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.732 |
-0.177 |
0.000 |
| y |
-0.177 |
2.702 |
0.000 |
| z |
0.000 |
0.000 |
-5.434 |
|
| Polar |
|---|
| 3z2-r2 | -10.867 |
|---|
| x2-y2 | 0.020 |
|---|
| xy | -0.177 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
173.008 |
| (<r2>)1/2 |
13.153 |