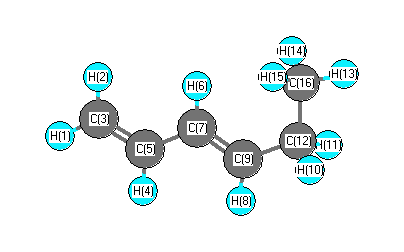
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3258 |
3117 |
16.36 |
|
|
|
| 2 |
A' |
3176 |
3038 |
23.66 |
|
|
|
| 3 |
A' |
3173 |
3035 |
4.94 |
|
|
|
| 4 |
A' |
3158 |
3021 |
14.33 |
|
|
|
| 5 |
A' |
3146 |
3010 |
19.07 |
|
|
|
| 6 |
A' |
3139 |
3003 |
32.68 |
|
|
|
| 7 |
A' |
3060 |
2928 |
20.65 |
|
|
|
| 8 |
A' |
3020 |
2889 |
35.21 |
|
|
|
| 9 |
A' |
1746 |
1670 |
20.69 |
|
|
|
| 10 |
A' |
1694 |
1621 |
3.55 |
|
|
|
| 11 |
A' |
1534 |
1467 |
5.23 |
|
|
|
| 12 |
A' |
1497 |
1432 |
4.91 |
|
|
|
| 13 |
A' |
1470 |
1407 |
1.92 |
|
|
|
| 14 |
A' |
1435 |
1373 |
7.12 |
|
|
|
| 15 |
A' |
1394 |
1333 |
16.03 |
|
|
|
| 16 |
A' |
1350 |
1292 |
4.05 |
|
|
|
| 17 |
A' |
1331 |
1274 |
0.57 |
|
|
|
| 18 |
A' |
1300 |
1243 |
0.18 |
|
|
|
| 19 |
A' |
1214 |
1161 |
4.85 |
|
|
|
| 20 |
A' |
1111 |
1063 |
4.40 |
|
|
|
| 21 |
A' |
1049 |
1004 |
2.97 |
|
|
|
| 22 |
A' |
962 |
920 |
1.98 |
|
|
|
| 23 |
A' |
904 |
865 |
2.61 |
|
|
|
| 24 |
A' |
570 |
545 |
3.38 |
|
|
|
| 25 |
A' |
468 |
448 |
0.37 |
|
|
|
| 26 |
A' |
306 |
293 |
0.54 |
|
|
|
| 27 |
A' |
149 |
142 |
0.69 |
|
|
|
| 28 |
A" |
3135 |
2999 |
30.92 |
|
|
|
| 29 |
A" |
3043 |
2911 |
19.86 |
|
|
|
| 30 |
A" |
1524 |
1458 |
7.09 |
|
|
|
| 31 |
A" |
1300 |
1244 |
0.06 |
|
|
|
| 32 |
A" |
1109 |
1061 |
1.18 |
|
|
|
| 33 |
A" |
1054 |
1008 |
37.18 |
|
|
|
| 34 |
A" |
990 |
947 |
5.69 |
|
|
|
| 35 |
A" |
918 |
878 |
41.45 |
|
|
|
| 36 |
A" |
874 |
837 |
1.45 |
|
|
|
| 37 |
A" |
770 |
737 |
8.22 |
|
|
|
| 38 |
A" |
636 |
609 |
0.62 |
|
|
|
| 39 |
A" |
307 |
293 |
1.75 |
|
|
|
| 40 |
A" |
238 |
227 |
0.41 |
|
|
|
| 41 |
A" |
160 |
153 |
0.91 |
|
|
|
| 42 |
A" |
115 |
110 |
0.24 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31393.3 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 30034.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.165 |
|
|
0.166 |
| 2 |
H |
0.162 |
|
|
0.172 |
| 3 |
C |
-0.410 |
|
|
-0.419 |
| 4 |
H |
0.152 |
|
|
0.135 |
| 5 |
C |
-0.091 |
|
|
-0.079 |
| 6 |
H |
0.152 |
|
|
0.146 |
| 7 |
C |
-0.132 |
|
|
-0.130 |
| 8 |
H |
0.151 |
|
|
0.113 |
| 9 |
C |
-0.150 |
|
|
-0.220 |
| 10 |
H |
0.176 |
|
|
0.015 |
| 11 |
H |
0.176 |
|
|
0.015 |
| 12 |
C |
-0.351 |
|
|
0.148 |
| 13 |
H |
0.170 |
|
|
0.071 |
| 14 |
H |
0.175 |
|
|
0.077 |
| 15 |
H |
0.175 |
|
|
0.077 |
| 16 |
C |
-0.520 |
|
|
-0.286 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.143 |
-0.700 |
0.000 |
0.715 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-0.117 |
-0.669 |
0.000 |
0.680 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.540 |
0.259 |
0.000 |
| y |
0.259 |
-35.630 |
0.000 |
| z |
0.000 |
0.000 |
-40.582 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.566 |
0.259 |
0.000 |
| y |
0.259 |
2.431 |
0.000 |
| z |
0.000 |
0.000 |
-4.997 |
|
| Polar |
|---|
| 3z2-r2 | -9.994 |
|---|
| x2-y2 | 0.090 |
|---|
| xy | 0.259 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
249.025 |
| (<r2>)1/2 |
15.781 |