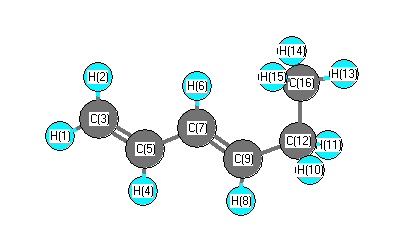
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3180 |
3128 |
16.03 |
|
|
|
| 2 |
A' |
3094 |
3043 |
5.48 |
|
|
|
| 3 |
A' |
3090 |
3039 |
26.75 |
|
|
|
| 4 |
A' |
3073 |
3022 |
13.24 |
|
|
|
| 5 |
A' |
3067 |
3016 |
29.96 |
|
|
|
| 6 |
A' |
3064 |
3013 |
18.68 |
|
|
|
| 7 |
A' |
2988 |
2938 |
19.86 |
|
|
|
| 8 |
A' |
2940 |
2891 |
33.91 |
|
|
|
| 9 |
A' |
1681 |
1654 |
21.29 |
|
|
|
| 10 |
A' |
1638 |
1611 |
3.09 |
|
|
|
| 11 |
A' |
1490 |
1466 |
5.66 |
|
|
|
| 12 |
A' |
1451 |
1427 |
6.14 |
|
|
|
| 13 |
A' |
1427 |
1404 |
2.09 |
|
|
|
| 14 |
A' |
1389 |
1366 |
6.29 |
|
|
|
| 15 |
A' |
1347 |
1325 |
17.71 |
|
|
|
| 16 |
A' |
1309 |
1287 |
4.04 |
|
|
|
| 17 |
A' |
1291 |
1270 |
0.46 |
|
|
|
| 18 |
A' |
1260 |
1239 |
0.26 |
|
|
|
| 19 |
A' |
1186 |
1166 |
4.13 |
|
|
|
| 20 |
A' |
1084 |
1067 |
5.57 |
|
|
|
| 21 |
A' |
1019 |
1002 |
3.38 |
|
|
|
| 22 |
A' |
937 |
922 |
2.03 |
|
|
|
| 23 |
A' |
883 |
868 |
2.92 |
|
|
|
| 24 |
A' |
556 |
547 |
3.64 |
|
|
|
| 25 |
A' |
457 |
450 |
0.40 |
|
|
|
| 26 |
A' |
300 |
295 |
0.42 |
|
|
|
| 27 |
A' |
145 |
143 |
0.64 |
|
|
|
| 28 |
A" |
3060 |
3009 |
28.42 |
|
|
|
| 29 |
A" |
2961 |
2912 |
18.84 |
|
|
|
| 30 |
A" |
1480 |
1456 |
6.83 |
|
|
|
| 31 |
A" |
1258 |
1238 |
0.03 |
|
|
|
| 32 |
A" |
1072 |
1054 |
0.91 |
|
|
|
| 33 |
A" |
1020 |
1003 |
33.59 |
|
|
|
| 34 |
A" |
951 |
935 |
3.75 |
|
|
|
| 35 |
A" |
868 |
854 |
37.08 |
|
|
|
| 36 |
A" |
840 |
826 |
1.89 |
|
|
|
| 37 |
A" |
747 |
735 |
8.42 |
|
|
|
| 38 |
A" |
618 |
608 |
0.37 |
|
|
|
| 39 |
A" |
301 |
296 |
1.73 |
|
|
|
| 40 |
A" |
233 |
229 |
0.25 |
|
|
|
| 41 |
A" |
158 |
155 |
0.95 |
|
|
|
| 42 |
A" |
115 |
113 |
0.28 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30512.5 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 30009.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.150 |
|
|
|
| 2 |
H |
0.146 |
|
|
|
| 3 |
C |
-0.380 |
|
|
|
| 4 |
H |
0.130 |
|
|
|
| 5 |
C |
-0.069 |
|
|
|
| 6 |
H |
0.128 |
|
|
|
| 7 |
C |
-0.106 |
|
|
|
| 8 |
H |
0.130 |
|
|
|
| 9 |
C |
-0.127 |
|
|
|
| 10 |
H |
0.165 |
|
|
|
| 11 |
H |
0.165 |
|
|
|
| 12 |
C |
-0.332 |
|
|
|
| 13 |
H |
0.159 |
|
|
|
| 14 |
H |
0.166 |
|
|
|
| 15 |
H |
0.166 |
|
|
|
| 16 |
C |
-0.491 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.138 |
-0.752 |
0.000 |
0.765 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.852 |
0.223 |
0.000 |
| y |
0.223 |
-35.833 |
0.000 |
| z |
0.000 |
0.000 |
-40.640 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.385 |
0.223 |
0.000 |
| y |
0.223 |
2.413 |
0.000 |
| z |
0.000 |
0.000 |
-4.798 |
|
| Polar |
|---|
| 3z2-r2 | -9.596 |
|---|
| x2-y2 | -0.019 |
|---|
| xy | 0.223 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.511 |
1.714 |
0.000 |
| y |
1.714 |
16.626 |
0.000 |
| z |
0.000 |
0.000 |
5.313 |
<r2> (average value of r
2) Å
2
| <r2> |
251.139 |
| (<r2>)1/2 |
15.847 |