Jump to
S1C2
Energy calculated at B3PW91/6-31G*
| | hartrees |
|---|
| Energy at 0K | -133.900578 |
| Energy at 298.15K | -133.906041 |
| HF Energy | -133.900579 |
| Nuclear repulsion energy | 70.837576 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3447 |
3298 |
5.30 |
|
|
|
| 2 |
A' |
3172 |
3035 |
10.54 |
|
|
|
| 3 |
A' |
3053 |
2921 |
11.86 |
|
|
|
| 4 |
A' |
3020 |
2890 |
71.43 |
|
|
|
| 5 |
A' |
1759 |
1683 |
67.59 |
|
|
|
| 6 |
A' |
1494 |
1430 |
13.18 |
|
|
|
| 7 |
A' |
1457 |
1393 |
21.75 |
|
|
|
| 8 |
A' |
1406 |
1346 |
12.30 |
|
|
|
| 9 |
A' |
1290 |
1234 |
36.52 |
|
|
|
| 10 |
A' |
1070 |
1024 |
26.08 |
|
|
|
| 11 |
A' |
938 |
897 |
7.69 |
|
|
|
| 12 |
A' |
488 |
467 |
17.17 |
|
|
|
| 13 |
A" |
3113 |
2978 |
15.01 |
|
|
|
| 14 |
A" |
1502 |
1437 |
9.91 |
|
|
|
| 15 |
A" |
1133 |
1084 |
9.26 |
|
|
|
| 16 |
A" |
1085 |
1038 |
11.04 |
|
|
|
| 17 |
A" |
684 |
655 |
59.14 |
|
|
|
| 18 |
A" |
196 |
188 |
1.04 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 15154.4 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 14498.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.032 |
-0.634 |
0.000 |
| C2 |
0.000 |
0.450 |
0.000 |
| N3 |
1.244 |
0.183 |
0.000 |
| H4 |
-0.547 |
-1.613 |
0.000 |
| H5 |
-1.681 |
-0.548 |
0.881 |
| H6 |
-1.681 |
-0.548 |
-0.881 |
| H7 |
-0.397 |
1.478 |
0.000 |
| H8 |
1.791 |
1.049 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4964 | 2.4182 | 1.0928 | 1.0977 | 1.0977 | 2.2051 | 3.2868 |
C2 | 1.4964 | | 1.2722 | 2.1343 | 2.1443 | 2.1443 | 1.1024 | 1.8888 | N3 | 2.4182 | 1.2722 | | 2.5369 | 3.1410 | 3.1410 | 2.0908 | 1.0244 | H4 | 1.0928 | 2.1343 | 2.5369 | | 1.7876 | 1.7876 | 3.0949 | 3.5436 | H5 | 1.0977 | 2.1443 | 3.1410 | 1.7876 | | 1.7627 | 2.5553 | 3.9221 | H6 | 1.0977 | 2.1443 | 3.1410 | 1.7876 | 1.7627 | | 2.5553 | 3.9221 | H7 | 2.2051 | 1.1024 | 2.0908 | 3.0949 | 2.5553 | 2.5553 | | 2.2303 | H8 | 3.2868 | 1.8888 | 1.0244 | 3.5436 | 3.9221 | 3.9221 | 2.2303 | |
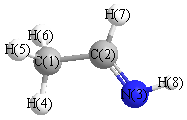 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
121.507 |
|
C1 |
C2 |
H7 |
115.260 |
| C2 |
C1 |
H4 |
110.065 |
|
C2 |
C1 |
H5 |
110.558 |
| C2 |
C1 |
H6 |
110.558 |
|
C2 |
N3 |
H8 |
110.188 |
| N3 |
C2 |
H7 |
123.232 |
|
H4 |
C1 |
H5 |
109.390 |
| H4 |
C1 |
H6 |
109.390 |
|
H5 |
C1 |
H6 |
106.813 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.554 |
|
|
|
| 2 |
C |
0.068 |
|
|
|
| 3 |
N |
-0.529 |
|
|
|
| 4 |
H |
0.203 |
|
|
|
| 5 |
H |
0.184 |
|
|
|
| 6 |
H |
0.184 |
|
|
|
| 7 |
H |
0.140 |
|
|
|
| 8 |
H |
0.303 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.421 |
1.424 |
0.000 |
2.012 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-19.329 |
2.559 |
0.000 |
| y |
2.559 |
-17.384 |
0.000 |
| z |
0.000 |
0.000 |
-19.469 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.903 |
2.559 |
0.000 |
| y |
2.559 |
2.015 |
0.000 |
| z |
0.000 |
0.000 |
-1.113 |
|
| Polar |
|---|
| 3z2-r2 | -2.225 |
|---|
| x2-y2 | -1.945 |
|---|
| xy | 2.559 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.614 |
0.329 |
0.000 |
| y |
0.329 |
4.077 |
0.000 |
| z |
0.000 |
0.000 |
2.717 |
<r2> (average value of r
2) Å
2
| <r2> |
50.138 |
| (<r2>)1/2 |
7.081 |
Jump to
S1C1
Energy calculated at B3PW91/6-31G*
| | hartrees |
|---|
| Energy at 0K | -133.900030 |
| Energy at 298.15K | -133.905466 |
| HF Energy | -133.900030 |
| Nuclear repulsion energy | 70.753061 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3404 |
3257 |
11.39 |
|
|
|
| 2 |
A' |
3144 |
3008 |
28.31 |
|
|
|
| 3 |
A' |
3098 |
2964 |
30.32 |
|
|
|
| 4 |
A' |
3048 |
2916 |
9.06 |
|
|
|
| 5 |
A' |
1759 |
1683 |
66.73 |
|
|
|
| 6 |
A' |
1503 |
1438 |
26.39 |
|
|
|
| 7 |
A' |
1445 |
1383 |
23.65 |
|
|
|
| 8 |
A' |
1415 |
1353 |
5.92 |
|
|
|
| 9 |
A' |
1296 |
1240 |
58.32 |
|
|
|
| 10 |
A' |
1077 |
1030 |
21.12 |
|
|
|
| 11 |
A' |
920 |
880 |
4.18 |
|
|
|
| 12 |
A' |
492 |
471 |
7.28 |
|
|
|
| 13 |
A" |
3116 |
2981 |
11.33 |
|
|
|
| 14 |
A" |
1495 |
1430 |
9.48 |
|
|
|
| 15 |
A" |
1157 |
1107 |
46.35 |
|
|
|
| 16 |
A" |
1084 |
1037 |
12.20 |
|
|
|
| 17 |
A" |
689 |
660 |
8.85 |
|
|
|
| 18 |
A" |
175 |
167 |
0.92 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 15157.4 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 14501.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.020 |
-0.624 |
0.000 |
| C2 |
0.000 |
0.479 |
0.000 |
| N3 |
1.268 |
0.383 |
0.000 |
| H4 |
-0.548 |
-1.612 |
0.000 |
| H5 |
-1.671 |
-0.545 |
0.881 |
| H6 |
-1.671 |
-0.545 |
-0.881 |
| H7 |
-0.410 |
1.496 |
0.000 |
| H8 |
1.545 |
-0.606 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5021 | 2.4996 | 1.0954 | 1.0977 | 1.0977 | 2.2063 | 2.5647 |
C2 | 1.5021 | | 1.2714 | 2.1619 | 2.1480 | 2.1480 | 1.0971 | 1.8878 | N3 | 2.4996 | 1.2714 | | 2.6980 | 3.2048 | 3.2048 | 2.0137 | 1.0275 | H4 | 1.0954 | 2.1619 | 2.6980 | | 1.7819 | 1.7819 | 3.1119 | 2.3220 | H5 | 1.0977 | 2.1480 | 3.2048 | 1.7819 | | 1.7612 | 2.5555 | 3.3342 | H6 | 1.0977 | 2.1480 | 3.2048 | 1.7819 | 1.7612 | | 2.5555 | 3.3342 | H7 | 2.2063 | 1.0971 | 2.0137 | 3.1119 | 2.5555 | 2.5555 | | 2.8712 | H8 | 2.5647 | 1.8878 | 1.0275 | 2.3220 | 3.3342 | 3.3342 | 2.8712 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
128.448 |
|
C1 |
C2 |
H7 |
115.279 |
| C2 |
C1 |
H4 |
111.712 |
|
C2 |
C1 |
H5 |
110.459 |
| C2 |
C1 |
H6 |
110.459 |
|
C2 |
N3 |
H8 |
109.951 |
| N3 |
C2 |
H7 |
116.273 |
|
H4 |
C1 |
H5 |
108.684 |
| H4 |
C1 |
H6 |
108.684 |
|
H5 |
C1 |
H6 |
106.686 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.568 |
|
|
|
| 2 |
C |
0.068 |
|
|
|
| 3 |
N |
-0.524 |
|
|
|
| 4 |
H |
0.178 |
|
|
|
| 5 |
H |
0.195 |
|
|
|
| 6 |
H |
0.195 |
|
|
|
| 7 |
H |
0.165 |
|
|
|
| 8 |
H |
0.292 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.016 |
-1.490 |
0.000 |
2.507 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-20.861 |
-3.214 |
0.000 |
| y |
-3.214 |
-18.695 |
0.000 |
| z |
0.000 |
0.000 |
-19.438 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.795 |
-3.214 |
0.000 |
| y |
-3.214 |
1.455 |
0.000 |
| z |
0.000 |
0.000 |
0.340 |
|
| Polar |
|---|
| 3z2-r2 | 0.680 |
|---|
| x2-y2 | -2.166 |
|---|
| xy | -3.214 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.459 |
-0.112 |
0.000 |
| y |
-0.112 |
4.104 |
0.000 |
| z |
0.000 |
0.000 |
2.709 |
<r2> (average value of r
2) Å
2
| <r2> |
50.302 |
| (<r2>)1/2 |
7.092 |
