Jump to
S1C2
Energy calculated at QCISD/6-31G*
| | hartrees |
|---|
| Energy at 0K | -133.524330 |
| Energy at 298.15K | -133.529796 |
| HF Energy | -133.071767 |
| Nuclear repulsion energy | 70.600007 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3431 |
3268 |
7.10 |
|
|
|
| 2 |
A' |
3177 |
3025 |
11.77 |
|
|
|
| 3 |
A' |
3068 |
2922 |
23.30 |
|
|
|
| 4 |
A' |
3057 |
2911 |
45.58 |
|
|
|
| 5 |
A' |
1751 |
1668 |
45.66 |
|
|
|
| 6 |
A' |
1527 |
1455 |
8.35 |
|
|
|
| 7 |
A' |
1479 |
1409 |
23.34 |
|
|
|
| 8 |
A' |
1440 |
1371 |
11.68 |
|
|
|
| 9 |
A' |
1313 |
1250 |
33.17 |
|
|
|
| 10 |
A' |
1092 |
1040 |
25.07 |
|
|
|
| 11 |
A' |
955 |
909 |
5.09 |
|
|
|
| 12 |
A' |
495 |
471 |
17.51 |
|
|
|
| 13 |
A" |
3129 |
2980 |
18.73 |
|
|
|
| 14 |
A" |
1529 |
1456 |
8.36 |
|
|
|
| 15 |
A" |
1141 |
1087 |
4.31 |
|
|
|
| 16 |
A" |
1107 |
1054 |
16.97 |
|
|
|
| 17 |
A" |
695 |
662 |
57.31 |
|
|
|
| 18 |
A" |
181 |
173 |
1.86 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 15283.0 cm
-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 14554.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.041 |
-0.627 |
0.000 |
| C2 |
0.000 |
0.455 |
0.000 |
| N3 |
1.249 |
0.171 |
0.000 |
| H4 |
-0.551 |
-1.606 |
0.000 |
| H5 |
-1.686 |
-0.542 |
0.885 |
| H6 |
-1.686 |
-0.542 |
-0.885 |
| H7 |
-0.382 |
1.488 |
0.000 |
| H8 |
1.805 |
1.036 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5014 | 2.4255 | 1.0940 | 1.0982 | 1.0982 | 2.2152 | 3.2960 |
C2 | 1.5014 | | 1.2811 | 2.1331 | 2.1491 | 2.1491 | 1.1013 | 1.8959 | N3 | 2.4255 | 1.2811 | | 2.5301 | 3.1477 | 3.1477 | 2.0962 | 1.0276 | H4 | 1.0940 | 2.1331 | 2.5301 | | 1.7892 | 1.7892 | 3.0981 | 3.5399 | H5 | 1.0982 | 2.1491 | 3.1477 | 1.7892 | | 1.7694 | 2.5696 | 3.9316 | H6 | 1.0982 | 2.1491 | 3.1477 | 1.7892 | 1.7694 | | 2.5696 | 3.9316 | H7 | 2.2152 | 1.1013 | 2.0962 | 3.0981 | 2.5696 | 2.5696 | | 2.2328 | H8 | 3.2960 | 1.8959 | 1.0276 | 3.5399 | 3.9316 | 3.9316 | 2.2328 | |
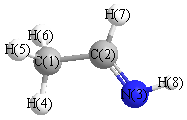 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
121.107 |
|
C1 |
C2 |
H7 |
115.816 |
| C2 |
C1 |
H4 |
109.542 |
|
C2 |
C1 |
H5 |
110.565 |
| C2 |
C1 |
H6 |
110.565 |
|
C2 |
N3 |
H8 |
109.927 |
| N3 |
C2 |
H7 |
123.077 |
|
H4 |
C1 |
H5 |
109.400 |
| H4 |
C1 |
H6 |
109.400 |
|
H5 |
C1 |
H6 |
107.331 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/6-31G*
| | hartrees |
|---|
| Energy at 0K | -133.523449 |
| Energy at 298.15K | -133.528887 |
| HF Energy | -133.070720 |
| Nuclear repulsion energy | 70.514739 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3391 |
3229 |
11.83 |
|
|
|
| 2 |
A' |
3151 |
3001 |
37.04 |
|
|
|
| 3 |
A' |
3129 |
2980 |
16.06 |
|
|
|
| 4 |
A' |
3061 |
2915 |
9.75 |
|
|
|
| 5 |
A' |
1748 |
1665 |
47.06 |
|
|
|
| 6 |
A' |
1533 |
1460 |
21.60 |
|
|
|
| 7 |
A' |
1467 |
1397 |
21.68 |
|
|
|
| 8 |
A' |
1448 |
1379 |
4.67 |
|
|
|
| 9 |
A' |
1319 |
1256 |
54.95 |
|
|
|
| 10 |
A' |
1095 |
1043 |
21.94 |
|
|
|
| 11 |
A' |
937 |
892 |
2.02 |
|
|
|
| 12 |
A' |
500 |
477 |
8.54 |
|
|
|
| 13 |
A" |
3133 |
2983 |
14.29 |
|
|
|
| 14 |
A" |
1522 |
1450 |
7.76 |
|
|
|
| 15 |
A" |
1163 |
1108 |
44.70 |
|
|
|
| 16 |
A" |
1105 |
1052 |
13.35 |
|
|
|
| 17 |
A" |
697 |
663 |
11.40 |
|
|
|
| 18 |
A" |
160 |
152 |
0.54 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 15279.1 cm
-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 14550.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.028 |
-0.618 |
0.000 |
| C2 |
0.000 |
0.485 |
0.000 |
| N3 |
1.276 |
0.371 |
0.000 |
| H4 |
-0.550 |
-1.605 |
0.000 |
| H5 |
-1.675 |
-0.539 |
0.884 |
| H6 |
-1.675 |
-0.539 |
-0.884 |
| H7 |
-0.397 |
1.506 |
0.000 |
| H8 |
1.536 |
-0.625 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5074 | 2.5073 | 1.0968 | 1.0980 | 1.0980 | 2.2158 | 2.5648 |
C2 | 1.5074 | | 1.2806 | 2.1604 | 2.1525 | 2.1525 | 1.0961 | 1.8954 | N3 | 2.5073 | 1.2806 | | 2.6904 | 3.2115 | 3.2115 | 2.0212 | 1.0304 | H4 | 1.0968 | 2.1604 | 2.6904 | | 1.7840 | 1.7840 | 3.1147 | 2.3049 | H5 | 1.0980 | 2.1525 | 3.2115 | 1.7840 | | 1.7683 | 2.5683 | 3.3317 | H6 | 1.0980 | 2.1525 | 3.2115 | 1.7840 | 1.7683 | | 2.5683 | 3.3317 | H7 | 2.2158 | 1.0961 | 2.0212 | 3.1147 | 2.5683 | 2.5683 | | 2.8779 | H8 | 2.5648 | 1.8954 | 1.0304 | 2.3049 | 3.3317 | 3.3317 | 2.8779 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
127.946 |
|
C1 |
C2 |
H7 |
115.756 |
| C2 |
C1 |
H4 |
111.136 |
|
C2 |
C1 |
H5 |
110.422 |
| C2 |
C1 |
H6 |
110.422 |
|
C2 |
N3 |
H8 |
109.730 |
| N3 |
C2 |
H7 |
116.299 |
|
H4 |
C1 |
H5 |
108.748 |
| H4 |
C1 |
H6 |
108.748 |
|
H5 |
C1 |
H6 |
107.259 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
