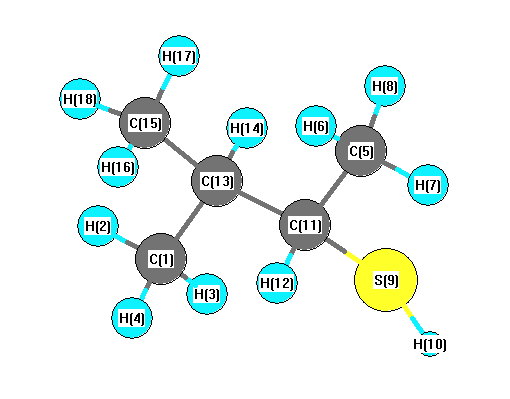
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3148 |
3011 |
24.21 |
|
|
|
| 2 |
A |
3145 |
3009 |
31.49 |
|
|
|
| 3 |
A |
3143 |
3007 |
23.92 |
|
|
|
| 4 |
A |
3132 |
2997 |
32.09 |
|
|
|
| 5 |
A |
3128 |
2993 |
52.79 |
|
|
|
| 6 |
A |
3123 |
2988 |
0.87 |
|
|
|
| 7 |
A |
3064 |
2931 |
33.72 |
|
|
|
| 8 |
A |
3060 |
2927 |
19.90 |
|
|
|
| 9 |
A |
3055 |
2923 |
19.45 |
|
|
|
| 10 |
A |
3052 |
2919 |
16.79 |
|
|
|
| 11 |
A |
3036 |
2905 |
3.89 |
|
|
|
| 12 |
A |
2707 |
2589 |
26.84 |
|
|
|
| 13 |
A |
1546 |
1479 |
3.18 |
|
|
|
| 14 |
A |
1530 |
1464 |
5.53 |
|
|
|
| 15 |
A |
1527 |
1460 |
21.61 |
|
|
|
| 16 |
A |
1516 |
1450 |
2.40 |
|
|
|
| 17 |
A |
1514 |
1449 |
3.76 |
|
|
|
| 18 |
A |
1508 |
1443 |
1.18 |
|
|
|
| 19 |
A |
1446 |
1383 |
6.50 |
|
|
|
| 20 |
A |
1435 |
1373 |
9.92 |
|
|
|
| 21 |
A |
1425 |
1364 |
3.64 |
|
|
|
| 22 |
A |
1398 |
1337 |
0.83 |
|
|
|
| 23 |
A |
1362 |
1303 |
0.47 |
|
|
|
| 24 |
A |
1334 |
1276 |
4.08 |
|
|
|
| 25 |
A |
1268 |
1213 |
30.78 |
|
|
|
| 26 |
A |
1211 |
1159 |
2.98 |
|
|
|
| 27 |
A |
1189 |
1138 |
7.93 |
|
|
|
| 28 |
A |
1166 |
1115 |
0.76 |
|
|
|
| 29 |
A |
1100 |
1053 |
1.55 |
|
|
|
| 30 |
A |
1052 |
1006 |
11.02 |
|
|
|
| 31 |
A |
996 |
952 |
3.40 |
|
|
|
| 32 |
A |
979 |
936 |
0.59 |
|
|
|
| 33 |
A |
940 |
899 |
0.95 |
|
|
|
| 34 |
A |
937 |
896 |
2.50 |
|
|
|
| 35 |
A |
906 |
867 |
4.07 |
|
|
|
| 36 |
A |
799 |
765 |
2.98 |
|
|
|
| 37 |
A |
696 |
666 |
2.70 |
|
|
|
| 38 |
A |
478 |
457 |
0.51 |
|
|
|
| 39 |
A |
407 |
390 |
0.60 |
|
|
|
| 40 |
A |
375 |
358 |
0.16 |
|
|
|
| 41 |
A |
369 |
353 |
0.39 |
|
|
|
| 42 |
A |
334 |
319 |
1.38 |
|
|
|
| 43 |
A |
253 |
242 |
1.36 |
|
|
|
| 44 |
A |
240 |
230 |
2.92 |
|
|
|
| 45 |
A |
228 |
218 |
0.11 |
|
|
|
| 46 |
A |
198 |
190 |
3.27 |
|
|
|
| 47 |
A |
176 |
169 |
15.41 |
|
|
|
| 48 |
A |
61 |
58 |
1.86 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35344.3 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 33813.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.516 |
|
|
-0.626 |
| 2 |
H |
0.167 |
|
|
0.150 |
| 3 |
H |
0.195 |
|
|
0.189 |
| 4 |
H |
0.165 |
|
|
0.135 |
| 5 |
C |
-0.510 |
|
|
-0.281 |
| 6 |
H |
0.178 |
|
|
0.097 |
| 7 |
H |
0.170 |
|
|
0.099 |
| 8 |
H |
0.186 |
|
|
0.081 |
| 9 |
S |
-0.097 |
|
|
-0.401 |
| 10 |
H |
0.097 |
|
|
0.229 |
| 11 |
C |
-0.267 |
|
|
-0.160 |
| 12 |
H |
0.188 |
|
|
0.123 |
| 13 |
C |
-0.123 |
|
|
0.505 |
| 14 |
H |
0.171 |
|
|
0.003 |
| 15 |
C |
-0.514 |
|
|
-0.501 |
| 16 |
H |
0.165 |
|
|
0.131 |
| 17 |
H |
0.173 |
|
|
0.113 |
| 18 |
H |
0.173 |
|
|
0.116 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.949 |
1.444 |
0.629 |
1.839 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-0.986 |
1.477 |
0.681 |
1.902 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-46.226 |
2.805 |
1.775 |
| y |
2.805 |
-47.720 |
0.709 |
| z |
1.775 |
0.709 |
-47.656 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.462 |
2.805 |
1.775 |
| y |
2.805 |
-0.779 |
0.709 |
| z |
1.775 |
0.709 |
-0.683 |
|
| Polar |
|---|
| 3z2-r2 | -1.366 |
|---|
| x2-y2 | 1.494 |
|---|
| xy | 2.805 |
|---|
| xz | 1.775 |
|---|
| yz | 0.709 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
249.978 |
| (<r2>)1/2 |
15.811 |