Jump to
S1C2
Vibrational Frequencies calculated at CID/6-31G*
Geometric Data calculated at CID/6-31G*
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CID/6-31G*
| | hartrees |
|---|
| Energy at 0K | -192.731612 |
| Energy at 298.15K | -192.736552 |
| HF Energy | -192.192419 |
| Nuclear repulsion energy | 149.445137 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3373 |
3115 |
0.14 |
|
|
|
| 2 |
A1 |
3362 |
3106 |
11.40 |
|
|
|
| 3 |
A1 |
3335 |
3080 |
1.17 |
|
|
|
| 4 |
A1 |
1698 |
1568 |
0.07 |
|
|
|
| 5 |
A1 |
1555 |
1436 |
1.03 |
|
|
|
| 6 |
A1 |
1209 |
1117 |
0.00 |
|
|
|
| 7 |
A1 |
1147 |
1059 |
10.73 |
|
|
|
| 8 |
A1 |
1091 |
1008 |
5.58 |
|
|
|
| 9 |
A1 |
881 |
813 |
0.29 |
|
|
|
| 10 |
A2 |
937 |
865 |
0.00 |
|
|
|
| 11 |
A2 |
842 |
778 |
0.00 |
|
|
|
| 12 |
A2 |
546 |
504 |
0.00 |
|
|
|
| 13 |
B1 |
906 |
837 |
9.12 |
|
|
|
| 14 |
B1 |
759 |
701 |
0.35 |
|
|
|
| 15 |
B1 |
720 |
665 |
102.61 |
|
|
|
| 16 |
B1 |
517 |
477 |
9.94 |
|
|
|
| 17 |
B2 |
8703 |
8039 |
325396.10 |
|
|
|
| 18 |
B2 |
3342 |
3087 |
85.52 |
|
|
|
| 19 |
B2 |
3334 |
3079 |
36.95 |
|
|
|
| 20 |
B2 |
1459 |
1347 |
69.51 |
|
|
|
| 21 |
B2 |
1370 |
1265 |
0.27 |
|
|
|
| 22 |
B2 |
1222 |
1129 |
15.53 |
|
|
|
| 23 |
B2 |
1005 |
928 |
0.04 |
|
|
|
| 24 |
B2 |
894 |
826 |
22.07 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22102.1 cm
-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 20415.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CID/6-31G*
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
1.218 |
| C2 |
0.000 |
1.118 |
0.393 |
| C3 |
0.000 |
0.674 |
-1.002 |
| C4 |
0.000 |
-0.674 |
-1.002 |
| C5 |
0.000 |
-1.118 |
0.393 |
| H6 |
0.000 |
0.000 |
2.295 |
| H7 |
0.000 |
2.149 |
0.714 |
| H8 |
0.000 |
1.328 |
-1.859 |
| H9 |
0.000 |
-1.328 |
-1.859 |
| H10 |
0.000 |
-2.149 |
0.714 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.3895 | 2.3202 | 2.3202 | 1.3895 | 1.0767 | 2.2069 | 3.3514 | 3.3514 | 2.2069 |
C2 | 1.3895 | | 1.4640 | 2.2713 | 2.2362 | 2.2060 | 1.0792 | 2.2618 | 3.3250 | 3.2823 | C3 | 2.3202 | 1.4640 | | 1.3484 | 2.2713 | 3.3650 | 2.2622 | 1.0779 | 2.1780 | 3.3033 | C4 | 2.3202 | 2.2713 | 1.3484 | | 1.4640 | 3.3650 | 3.3033 | 2.1780 | 1.0779 | 2.2622 | C5 | 1.3895 | 2.2362 | 2.2713 | 1.4640 | | 2.2060 | 3.2823 | 3.3250 | 2.2618 | 1.0792 | H6 | 1.0767 | 2.2060 | 3.3650 | 3.3650 | 2.2060 | | 2.6676 | 4.3609 | 4.3609 | 2.6676 | H7 | 2.2069 | 1.0792 | 2.2622 | 3.3033 | 3.2823 | 2.6676 | | 2.7003 | 4.3250 | 4.2971 | H8 | 3.3514 | 2.2618 | 1.0779 | 2.1780 | 3.3250 | 4.3609 | 2.7003 | | 2.6563 | 4.3250 | H9 | 3.3514 | 3.3250 | 2.1780 | 1.0779 | 2.2618 | 4.3609 | 4.3250 | 2.6563 | | 2.7003 | H10 | 2.2069 | 3.2823 | 3.3033 | 2.2622 | 1.0792 | 2.6676 | 4.2971 | 4.3250 | 2.7003 | |
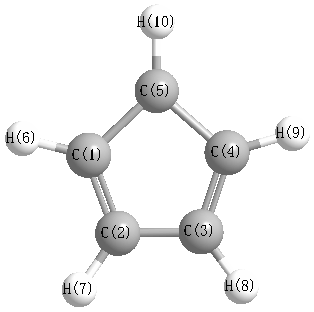 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
108.771 |
|
C1 |
C2 |
H7 |
126.296 |
| C1 |
C5 |
C4 |
108.771 |
|
C1 |
C5 |
H10 |
126.296 |
| C2 |
C1 |
C5 |
107.159 |
|
C2 |
C1 |
H6 |
126.421 |
| C2 |
C3 |
C4 |
107.649 |
|
C2 |
C3 |
H8 |
125.002 |
| C3 |
C2 |
H7 |
124.933 |
|
C3 |
C4 |
C5 |
107.649 |
| C3 |
C4 |
H9 |
127.348 |
|
C4 |
C3 |
H8 |
127.348 |
| C4 |
C5 |
H10 |
124.933 |
|
C5 |
C1 |
H6 |
126.421 |
| C5 |
C4 |
H9 |
125.002 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
