Jump to
S1C2
Vibrational Frequencies calculated at B3PW91/6-31G*
Geometric Data calculated at B3PW91/6-31G*
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B3PW91/6-31G*
| | hartrees |
|---|
| Energy at 0K | -193.389173 |
| Energy at 298.15K | |
| HF Energy | -193.389173 |
| Nuclear repulsion energy | 148.760151 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3276 |
3134 |
0.10 |
|
|
|
| 2 |
A1 |
3268 |
3127 |
12.38 |
|
|
|
| 3 |
A1 |
3235 |
3095 |
1.58 |
|
|
|
| 4 |
A1 |
1602 |
1533 |
0.04 |
|
|
|
| 5 |
A1 |
1480 |
1416 |
1.89 |
|
|
|
| 6 |
A1 |
1155 |
1105 |
0.00 |
|
|
|
| 7 |
A1 |
1097 |
1049 |
9.19 |
|
|
|
| 8 |
A1 |
1054 |
1009 |
9.81 |
|
|
|
| 9 |
A1 |
841 |
805 |
0.29 |
|
|
|
| 10 |
A2 |
902 |
863 |
0.00 |
|
|
|
| 11 |
A2 |
820 |
784 |
0.00 |
|
|
|
| 12 |
A2 |
526 |
503 |
0.00 |
|
|
|
| 13 |
B1 |
878 |
840 |
6.52 |
|
|
|
| 14 |
B1 |
719 |
688 |
1.62 |
|
|
|
| 15 |
B1 |
685 |
655 |
86.46 |
|
|
|
| 16 |
B1 |
489 |
468 |
11.23 |
|
|
|
| 17 |
B2 |
3251 |
3111 |
19.64 |
|
|
|
| 18 |
B2 |
3236 |
3096 |
9.78 |
|
|
|
| 19 |
B2 |
1417 |
1356 |
41.07 |
|
|
|
| 20 |
B2 |
1305 |
1249 |
0.00 |
|
|
|
| 21 |
B2 |
1227 |
1174 |
1.88 |
|
|
|
| 22 |
B2 |
982 |
939 |
0.22 |
|
|
|
| 23 |
B2 |
930 |
890 |
2.73 |
|
|
|
| 24 |
B2 |
37i |
36i |
4.20 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17168.1 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 16424.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-31G*
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
1.225 |
| C2 |
0.000 |
1.123 |
0.392 |
| C3 |
0.000 |
0.679 |
-1.005 |
| C4 |
0.000 |
-0.679 |
-1.005 |
| C5 |
0.000 |
-1.123 |
0.392 |
| H6 |
0.000 |
0.000 |
2.307 |
| H7 |
0.000 |
2.159 |
0.715 |
| H8 |
0.000 |
1.337 |
-1.866 |
| H9 |
0.000 |
-1.337 |
-1.866 |
| H10 |
0.000 |
-2.159 |
0.715 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.3977 | 2.3310 | 2.3310 | 1.3977 | 1.0823 | 2.2187 | 3.3677 | 3.3677 | 2.2187 |
C2 | 1.3977 | | 1.4662 | 2.2806 | 2.2457 | 2.2196 | 1.0853 | 2.2690 | 3.3393 | 3.2979 | C3 | 2.3310 | 1.4662 | | 1.3587 | 2.2806 | 3.3811 | 2.2689 | 1.0834 | 2.1922 | 3.3189 | C4 | 2.3310 | 2.2806 | 1.3587 | | 1.4662 | 3.3811 | 3.3189 | 2.1922 | 1.0834 | 2.2689 | C5 | 1.3977 | 2.2457 | 2.2806 | 1.4662 | | 2.2196 | 3.2979 | 3.3393 | 2.2690 | 1.0853 | H6 | 1.0823 | 2.2196 | 3.3811 | 3.3811 | 2.2196 | | 2.6829 | 4.3823 | 4.3823 | 2.6829 | H7 | 2.2187 | 1.0853 | 2.2689 | 3.3189 | 3.2979 | 2.6829 | | 2.7091 | 4.3454 | 4.3185 | H8 | 3.3677 | 2.2690 | 1.0834 | 2.1922 | 3.3393 | 4.3823 | 2.7091 | | 2.6730 | 4.3454 | H9 | 3.3677 | 3.3393 | 2.1922 | 1.0834 | 2.2690 | 4.3823 | 4.3454 | 2.6730 | | 2.7091 | H10 | 2.2187 | 3.2979 | 3.3189 | 2.2689 | 1.0853 | 2.6829 | 4.3185 | 4.3454 | 2.7091 | |
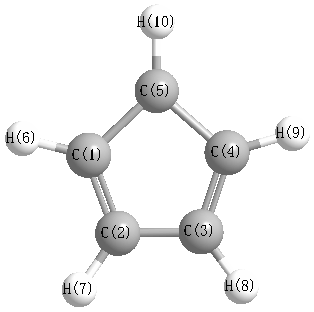 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
108.939 |
|
C1 |
C2 |
H7 |
126.181 |
| C1 |
C5 |
C4 |
108.939 |
|
C1 |
C5 |
H10 |
126.181 |
| C2 |
C1 |
C5 |
106.908 |
|
C2 |
C1 |
H6 |
126.546 |
| C2 |
C3 |
C4 |
107.607 |
|
C2 |
C3 |
H8 |
125.052 |
| C3 |
C2 |
H7 |
124.880 |
|
C3 |
C4 |
C5 |
107.607 |
| C3 |
C4 |
H9 |
127.341 |
|
C4 |
C3 |
H8 |
127.341 |
| C4 |
C5 |
H10 |
124.880 |
|
C5 |
C1 |
H6 |
126.546 |
| C5 |
C4 |
H9 |
125.052 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.189 |
|
|
|
| 2 |
C |
-0.146 |
|
|
|
| 3 |
C |
-0.174 |
|
|
|
| 4 |
C |
-0.174 |
|
|
|
| 5 |
C |
-0.146 |
|
|
|
| 6 |
H |
0.161 |
|
|
|
| 7 |
H |
0.170 |
|
|
|
| 8 |
H |
0.164 |
|
|
|
| 9 |
H |
0.164 |
|
|
|
| 10 |
H |
0.170 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.036 |
0.036 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.377 |
0.000 |
0.000 |
| y |
0.000 |
-25.016 |
0.000 |
| z |
0.000 |
0.000 |
-26.884 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.428 |
0.000 |
0.000 |
| y |
0.000 |
4.615 |
0.000 |
| z |
0.000 |
0.000 |
1.813 |
|
| Polar |
|---|
| 3z2-r2 | 3.626 |
|---|
| x2-y2 | -7.362 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.727 |
0.000 |
0.000 |
| y |
0.000 |
8.712 |
0.000 |
| z |
0.000 |
0.000 |
7.860 |
<r2> (average value of r
2) Å
2
| <r2> |
87.392 |
| (<r2>)1/2 |
9.348 |
