Jump to
S1C2
Vibrational Frequencies calculated at QCISD/6-31G*
Geometric Data calculated at QCISD/6-31G*
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/6-31G*
| | hartrees |
|---|
| Energy at 0K | -192.838330 |
| Energy at 298.15K | -192.843062 |
| HF Energy | -192.191621 |
| Nuclear repulsion energy | 148.345478 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3274 |
3118 |
0.13 |
|
|
|
| 2 |
A1 |
3266 |
3110 |
13.99 |
|
|
|
| 3 |
A1 |
3237 |
3082 |
1.33 |
|
|
|
| 4 |
A1 |
1622 |
1545 |
0.16 |
|
|
|
| 5 |
A1 |
1501 |
1429 |
0.47 |
|
|
|
| 6 |
A1 |
1167 |
1111 |
0.01 |
|
|
|
| 7 |
A1 |
1109 |
1056 |
7.73 |
|
|
|
| 8 |
A1 |
1059 |
1008 |
5.35 |
|
|
|
| 9 |
A1 |
851 |
811 |
0.29 |
|
|
|
| 10 |
A2 |
865 |
824 |
0.00 |
|
|
|
| 11 |
A2 |
773 |
736 |
0.00 |
|
|
|
| 12 |
A2 |
515 |
490 |
0.00 |
|
|
|
| 13 |
B1 |
826 |
787 |
10.30 |
|
|
|
| 14 |
B1 |
706 |
673 |
3.07 |
|
|
|
| 15 |
B1 |
673 |
641 |
90.00 |
|
|
|
| 16 |
B1 |
484 |
461 |
8.96 |
|
|
|
| 17 |
B2 |
3249 |
3094 |
18.59 |
|
|
|
| 18 |
B2 |
3237 |
3083 |
5.37 |
|
|
|
| 19 |
B2 |
1783 |
1698 |
289.71 |
|
|
|
| 20 |
B2 |
1416 |
1348 |
76.44 |
|
|
|
| 21 |
B2 |
1323 |
1260 |
0.03 |
|
|
|
| 22 |
B2 |
1155 |
1100 |
19.22 |
|
|
|
| 23 |
B2 |
980 |
933 |
0.01 |
|
|
|
| 24 |
B2 |
856 |
815 |
3.70 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17963.3 cm
-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 17106.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/6-31G*
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
1.226 |
| C2 |
0.000 |
1.128 |
0.395 |
| C3 |
0.000 |
0.680 |
-1.009 |
| C4 |
0.000 |
-0.680 |
-1.009 |
| C5 |
0.000 |
-1.128 |
0.395 |
| H6 |
0.000 |
0.000 |
2.310 |
| H7 |
0.000 |
2.166 |
0.719 |
| H8 |
0.000 |
1.338 |
-1.872 |
| H9 |
0.000 |
-1.338 |
-1.872 |
| H10 |
0.000 |
-2.166 |
0.719 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4005 | 2.3353 | 2.3353 | 1.4005 | 1.0844 | 2.2240 | 3.3739 | 3.3739 | 2.2240 |
C2 | 1.4005 | | 1.4740 | 2.2888 | 2.2560 | 2.2221 | 1.0868 | 2.2769 | 3.3496 | 3.3094 | C3 | 2.3353 | 1.4740 | | 1.3591 | 2.2888 | 3.3875 | 2.2787 | 1.0854 | 2.1941 | 3.3285 | C4 | 2.3353 | 2.2888 | 1.3591 | | 1.4740 | 3.3875 | 3.3285 | 2.1941 | 1.0854 | 2.2787 | C5 | 1.4005 | 2.2560 | 2.2888 | 1.4740 | | 2.2221 | 3.3094 | 3.3496 | 2.2769 | 1.0868 | H6 | 1.0844 | 2.2221 | 3.3875 | 3.3875 | 2.2221 | | 2.6872 | 4.3905 | 4.3905 | 2.6872 | H7 | 2.2240 | 1.0868 | 2.2787 | 3.3285 | 3.3094 | 2.6872 | | 2.7197 | 4.3571 | 4.3311 | H8 | 3.3739 | 2.2769 | 1.0854 | 2.1941 | 3.3496 | 4.3905 | 2.7197 | | 2.6753 | 4.3571 | H9 | 3.3739 | 3.3496 | 2.1941 | 1.0854 | 2.2769 | 4.3905 | 4.3571 | 2.6753 | | 2.7197 | H10 | 2.2240 | 3.3094 | 3.3285 | 2.2787 | 1.0868 | 2.6872 | 4.3311 | 4.3571 | 2.7197 | |
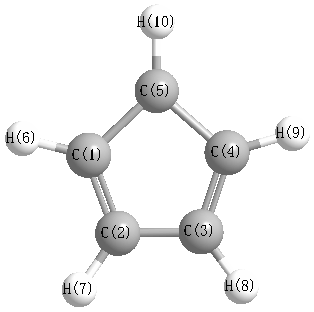 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
108.641 |
|
C1 |
C2 |
H7 |
126.335 |
| C1 |
C5 |
C4 |
108.641 |
|
C1 |
C5 |
H10 |
126.335 |
| C2 |
C1 |
C5 |
107.296 |
|
C2 |
C1 |
H6 |
126.352 |
| C2 |
C3 |
C4 |
107.711 |
|
C2 |
C3 |
H8 |
124.965 |
| C3 |
C2 |
H7 |
125.024 |
|
C3 |
C4 |
C5 |
107.711 |
| C3 |
C4 |
H9 |
127.324 |
|
C4 |
C3 |
H8 |
127.324 |
| C4 |
C5 |
H10 |
125.024 |
|
C5 |
C1 |
H6 |
126.352 |
| C5 |
C4 |
H9 |
124.965 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
