Jump to
S1C2
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
Geometric Data calculated at PBEPBEultrafine/6-31G*
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at PBEPBEultrafine/6-31G*
| | hartrees |
|---|
| Energy at 0K | -193.201384 |
| Energy at 298.15K | |
| HF Energy | -193.201384 |
| Nuclear repulsion energy | 147.880700 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3197 |
3145 |
0.18 |
|
|
|
| 2 |
A1 |
3190 |
3137 |
12.06 |
|
|
|
| 3 |
A1 |
3155 |
3103 |
1.89 |
|
|
|
| 4 |
A1 |
1549 |
1523 |
0.02 |
|
|
|
| 5 |
A1 |
1436 |
1412 |
2.41 |
|
|
|
| 6 |
A1 |
1123 |
1105 |
0.01 |
|
|
|
| 7 |
A1 |
1067 |
1049 |
7.91 |
|
|
|
| 8 |
A1 |
1029 |
1012 |
10.83 |
|
|
|
| 9 |
A1 |
816 |
803 |
0.29 |
|
|
|
| 10 |
A2 |
855 |
841 |
0.00 |
|
|
|
| 11 |
A2 |
792 |
779 |
0.00 |
|
|
|
| 12 |
A2 |
509 |
500 |
0.00 |
|
|
|
| 13 |
B1 |
841 |
827 |
5.62 |
|
|
|
| 14 |
B1 |
691 |
679 |
8.09 |
|
|
|
| 15 |
B1 |
657 |
646 |
69.61 |
|
|
|
| 16 |
B1 |
470 |
462 |
10.83 |
|
|
|
| 17 |
B2 |
3173 |
3121 |
18.24 |
|
|
|
| 18 |
B2 |
3156 |
3104 |
11.29 |
|
|
|
| 19 |
B2 |
1376 |
1353 |
36.87 |
|
|
|
| 20 |
B2 |
1262 |
1241 |
0.00 |
|
|
|
| 21 |
B2 |
1192 |
1172 |
2.19 |
|
|
|
| 22 |
B2 |
963 |
948 |
0.24 |
|
|
|
| 23 |
B2 |
906 |
891 |
2.99 |
|
|
|
| 24 |
B2 |
75i |
74i |
7.82 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16664.9 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 16390.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBEultrafine/6-31G*
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
1.232 |
| C2 |
0.000 |
1.129 |
0.394 |
| C3 |
0.000 |
0.685 |
-1.010 |
| C4 |
0.000 |
-0.685 |
-1.010 |
| C5 |
0.000 |
-1.129 |
0.394 |
| H6 |
0.000 |
0.000 |
2.322 |
| H7 |
0.000 |
2.173 |
0.719 |
| H8 |
0.000 |
1.347 |
-1.877 |
| H9 |
0.000 |
-1.347 |
-1.877 |
| H10 |
0.000 |
-2.173 |
0.719 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4059 | 2.3443 | 2.3443 | 1.4059 | 1.0899 | 2.2326 | 3.3883 | 3.3883 | 2.2326 |
C2 | 1.4059 | | 1.4727 | 2.2936 | 2.2577 | 2.2341 | 1.0933 | 2.2815 | 3.3599 | 3.3176 | C3 | 2.3443 | 1.4727 | | 1.3695 | 2.2936 | 3.4017 | 2.2811 | 1.0911 | 2.2092 | 3.3399 | C4 | 2.3443 | 2.2936 | 1.3695 | | 1.4727 | 3.4017 | 3.3399 | 2.2092 | 1.0911 | 2.2811 | C5 | 1.4059 | 2.2577 | 2.2936 | 1.4727 | | 2.2341 | 3.3176 | 3.3599 | 2.2815 | 1.0933 | H6 | 1.0899 | 2.2341 | 3.4017 | 3.4017 | 2.2341 | | 2.7003 | 4.4098 | 4.4098 | 2.7003 | H7 | 2.2326 | 1.0933 | 2.2811 | 3.3399 | 3.3176 | 2.7003 | | 2.7238 | 4.3737 | 4.3457 | H8 | 3.3883 | 2.2815 | 1.0911 | 2.2092 | 3.3599 | 4.4098 | 2.7238 | | 2.6946 | 4.3737 | H9 | 3.3883 | 3.3599 | 2.2092 | 1.0911 | 2.2815 | 4.4098 | 4.3737 | 2.6946 | | 2.7238 | H10 | 2.2326 | 3.3176 | 3.3399 | 2.2811 | 1.0933 | 2.7003 | 4.3457 | 4.3737 | 2.7238 | |
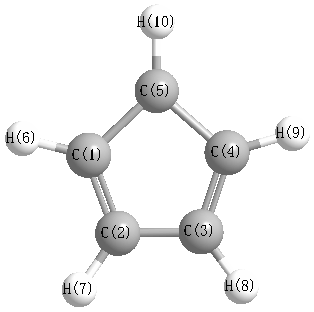 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
109.035 |
|
C1 |
C2 |
H7 |
126.136 |
| C1 |
C5 |
C4 |
109.035 |
|
C1 |
C5 |
H10 |
126.136 |
| C2 |
C1 |
C5 |
106.826 |
|
C2 |
C1 |
H6 |
126.587 |
| C2 |
C3 |
C4 |
107.553 |
|
C2 |
C3 |
H8 |
125.056 |
| C3 |
C2 |
H7 |
124.830 |
|
C3 |
C4 |
C5 |
107.553 |
| C3 |
C4 |
H9 |
127.392 |
|
C4 |
C3 |
H8 |
127.392 |
| C4 |
C5 |
H10 |
124.830 |
|
C5 |
C1 |
H6 |
126.587 |
| C5 |
C4 |
H9 |
125.056 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.155 |
|
|
|
| 2 |
C |
-0.129 |
|
|
|
| 3 |
C |
-0.148 |
|
|
|
| 4 |
C |
-0.148 |
|
|
|
| 5 |
C |
-0.129 |
|
|
|
| 6 |
H |
0.137 |
|
|
|
| 7 |
H |
0.147 |
|
|
|
| 8 |
H |
0.140 |
|
|
|
| 9 |
H |
0.140 |
|
|
|
| 10 |
H |
0.147 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.057 |
0.057 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.360 |
0.000 |
0.000 |
| y |
0.000 |
-25.323 |
0.000 |
| z |
0.000 |
0.000 |
-27.060 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.169 |
0.000 |
0.000 |
| y |
0.000 |
4.388 |
0.000 |
| z |
0.000 |
0.000 |
1.781 |
|
| Polar |
|---|
| 3z2-r2 | 3.563 |
|---|
| x2-y2 | -7.038 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.756 |
0.000 |
0.000 |
| y |
0.000 |
8.888 |
0.000 |
| z |
0.000 |
0.000 |
8.139 |
<r2> (average value of r
2) Å
2
| <r2> |
88.319 |
| (<r2>)1/2 |
9.398 |
