return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H7 (n-Propyl radical)

using model chemistry: CID/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 2A"
Energy calculated at CID/6-31G*
 hartrees
Energy at 0K-117.999121
Energy at 298.15K 
HF Energy-117.631501
Nuclear repulsion energy75.923745
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3353 3098 17.35      
2 A 3251 3003 14.32      
3 A 3217 2972 32.55      
4 A 3212 2967 39.50      
5 A 3148 2908 21.53      
6 A 3137 2897 24.21      
7 A 3076 2841 34.97      
8 A 1588 1467 3.65      
9 A 1581 1460 5.63      
10 A 1568 1449 0.74      
11 A 1545 1427 0.88      
12 A 1499 1384 2.50      
13 A 1440 1330 2.04      
14 A 1343 1240 0.24      
15 A 1237 1143 0.62      
16 A 1139 1052 0.05      
17 A 1117 1032 1.26      
18 A 965 891 1.41      
19 A 929 858 0.14      
20 A 796 735 0.93      
21 A 473 436 41.59      
22 A 382 352 5.53      
23 A 264 244 0.12      
24 A 144 133 0.59      

Unscaled Zero Point Vibrational Energy (zpe) 20201.4 cm-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 18660.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CID/6-31G*
ABC
1.09349 0.30033 0.26061

See section I.F.4 to change rotational constant units
Geometric Data calculated at CID/6-31G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.299 -0.295 -0.041
C2 0.075 0.558 0.058
C3 -1.220 -0.247 -0.040
H4 -2.094 0.403 0.035
H5 -1.275 -0.780 -0.991
H6 -1.281 -0.986 0.762
H7 0.079 1.114 1.006
H8 0.097 1.319 -0.728
H9 2.253 0.132 -0.314
H10 1.300 -1.296 0.369

Atom - Atom Distances (Å)
  C1 C2 C3 H4 H5 H6 H7 H8 H9 H10
C11.49452.51933.46442.78552.78792.13682.12621.08121.0821
C21.49451.52792.17482.17012.17131.09901.09392.25012.2441
C32.51931.52791.09121.09111.09192.15252.15813.50482.7611
H43.46442.17481.09121.76651.76542.48432.49404.36963.8107
H52.78552.17011.09111.76651.76423.06652.52053.70612.9578
H62.78792.17131.09191.76541.76422.51323.06993.85942.6292
H72.13681.09902.15252.48433.06652.51321.74552.72602.7758
H82.12621.09392.15812.49402.52053.06991.74552.49573.0808
H91.08122.25013.50484.36963.70613.85942.72602.49571.8484
H101.08212.24412.76113.81072.95782.62922.77583.08081.8484

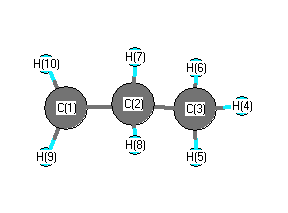
picture of n-Propyl radical state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 112.925 C1 C2 H7 110.019
C1 C2 H8 109.486 C2 C1 H9 120.927
C2 C1 H10 120.301 C2 C3 H4 111.179
C2 C3 H5 110.808 C2 C3 H6 110.858
C3 C2 H7 108.957 C3 C2 H8 109.694
H4 C3 H5 108.093 H4 C3 H6 107.935
H5 C3 H6 107.835 H7 C2 H8 105.491
H9 C1 H10 117.397
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability