Jump to
S2C1
Energy calculated at B3PW91/6-31G*
| | hartrees |
|---|
| Energy at 0K | -77.223146 |
| Energy at 298.15K | -77.222847 |
| Nuclear repulsion energy | 23.763482 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3159 |
3022 |
39.35 |
|
|
|
| 2 |
A1 |
1723 |
1649 |
83.94 |
|
|
|
| 3 |
A1 |
1238 |
1185 |
24.13 |
|
|
|
| 4 |
B1 |
755 |
722 |
100.79 |
|
|
|
| 5 |
B2 |
3236 |
3096 |
16.41 |
|
|
|
| 6 |
B2 |
353 |
338 |
1.97 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5232.4 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 5005.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-31G*
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-0.479 |
| C2 |
0.000 |
0.000 |
0.825 |
| H3 |
0.000 |
0.936 |
-1.038 |
| H4 |
0.000 |
-0.936 |
-1.038 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
H4 |
| C1 | | 1.3036 | 1.0907 | 1.0907 |
C2 | 1.3036 | | 2.0851 | 2.0851 | H3 | 1.0907 | 2.0851 | | 1.8727 | H4 | 1.0907 | 2.0851 | 1.8727 | |
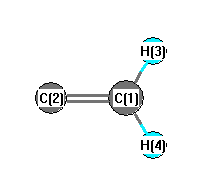 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C2 |
C1 |
H3 |
120.853 |
|
C2 |
C1 |
H4 |
120.853 |
| H3 |
C1 |
H4 |
118.293 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.498 |
|
|
|
| 2 |
C |
0.022 |
|
|
|
| 3 |
H |
0.238 |
|
|
|
| 4 |
H |
0.238 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-2.301 |
2.301 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-13.704 |
0.000 |
0.000 |
| y |
0.000 |
-9.927 |
0.000 |
| z |
0.000 |
0.000 |
-13.663 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.909 |
0.000 |
0.000 |
| y |
0.000 |
3.757 |
0.000 |
| z |
0.000 |
0.000 |
-1.848 |
|
| Polar |
|---|
| 3z2-r2 | -3.695 |
|---|
| x2-y2 | -3.777 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
1.442 |
0.000 |
0.000 |
| y |
0.000 |
2.456 |
0.000 |
| z |
0.000 |
0.000 |
3.769 |
<r2> (average value of r
2) Å
2
| <r2> |
17.131 |
| (<r2>)1/2 |
4.139 |
Jump to
S1C1
Energy calculated at B3PW91/6-31G*
| | hartrees |
|---|
| Energy at 0K | -77.158671 |
| Energy at 298.15K | -77.158736 |
| HF Energy | -77.158671 |
| Nuclear repulsion energy | 23.559191 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3070 |
2937 |
4.89 |
|
|
|
| 2 |
A1 |
1614 |
1544 |
4.36 |
|
|
|
| 3 |
A1 |
1421 |
1359 |
1.74 |
|
|
|
| 4 |
B1 |
781 |
747 |
12.86 |
|
|
|
| 5 |
B2 |
3140 |
3004 |
9.43 |
|
|
|
| 6 |
B2 |
1006 |
963 |
4.15 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5515.9 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 5277.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-31G*
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-0.483 |
| C2 |
0.000 |
0.000 |
0.834 |
| H3 |
0.000 |
0.934 |
-1.054 |
| H4 |
0.000 |
-0.934 |
-1.054 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
H4 |
| C1 | | 1.3174 | 1.0947 | 1.0947 |
C2 | 1.3174 | | 2.1066 | 2.1066 | H3 | 1.0947 | 2.1066 | | 1.8682 | H4 | 1.0947 | 2.1066 | 1.8682 | |
More geometry information
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.386 |
|
|
|
| 2 |
C |
0.004 |
|
|
|
| 3 |
H |
0.191 |
|
|
|
| 4 |
H |
0.191 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-0.541 |
0.541 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-13.413 |
0.000 |
0.000 |
| y |
0.000 |
-11.638 |
0.000 |
| z |
0.000 |
0.000 |
-10.841 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.173 |
0.000 |
0.000 |
| y |
0.000 |
0.489 |
0.000 |
| z |
0.000 |
0.000 |
1.684 |
|
| Polar |
|---|
| 3z2-r2 | 3.368 |
|---|
| x2-y2 | -1.775 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
1.586 |
0.000 |
0.000 |
| y |
0.000 |
2.134 |
0.000 |
| z |
0.000 |
0.000 |
3.897 |
<r2> (average value of r
2) Å
2
| <r2> |
17.016 |
| (<r2>)1/2 |
4.125 |
