return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H5CH2 (benzyl radical)

using model chemistry: QCISD/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 2B1
Energy calculated at QCISD/6-31G*
 hartrees
Energy at 0K-270.034316
Energy at 298.15K-270.041059
HF Energy-269.139439
Nuclear repulsion energy259.562897
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3223 3069 16.90      
2 A1 3203 3050 5.32      
3 A1 3189 3037 9.84      
4 A1 3181 3029 4.80      
5 A1 1642 1564 1.47      
6 A1 1537 1464 6.81      
7 A1 1528 1455 0.34      
8 A1 1308 1246 3.16      
9 A1 1211 1153 0.03      
10 A1 1058 1008 1.14      
11 A1 997 949 0.07      
12 A1 842 802 0.24      
13 A1 536 510 0.26      
14 A2 924 880 0.00      
15 A2 825 785 0.00      
16 A2 515 491 0.00      
17 A2 385 367 0.00      
18 B1 911 867 1.64      
19 B1 857 817 2.12      
20 B1 725 690 95.41      
21 B1 684 651 22.23      
22 B1 572 545 2.06      
23 B1 461 439 8.99      
24 B1 198 188 1.03      
25 B2 3276 3119 10.04      
26 B2 3209 3056 40.12      
27 B2 3190 3038 1.51      
28 B2 1628 1550 1.03      
29 B2 1509 1437 6.71      
30 B2 1375 1310 0.51      
31 B2 1318 1255 0.29      
32 B2 1190 1133 0.09      
33 B2 1144 1090 2.22      
34 B2 993 946 1.05      
35 B2 631 601 0.01      
36 B2 362 345 0.03      

Unscaled Zero Point Vibrational Energy (zpe) 25167.6 cm-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 23967.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G*
ABC
0.18408 0.09009 0.06049

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 0.994
C2 0.000 1.220 0.252
C3 0.000 1.213 -1.132
C4 0.000 0.000 -1.839
C5 0.000 -1.213 -1.132
C6 0.000 -1.220 0.252
C7 0.000 0.000 2.400
H8 0.000 2.165 0.795
H9 0.000 2.156 -1.677
H10 0.000 0.000 -2.928
H11 0.000 -2.156 -1.677
H12 0.000 -2.165 0.795
H13 0.000 0.929 2.964
H14 0.000 -0.929 2.964

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 H8 H9 H10 H11 H12 H13 H14
C11.42782.44812.83312.44811.42781.40552.17403.43283.92163.43282.17402.17812.1781
C21.42781.38462.42112.79922.43982.46971.09002.14453.40583.88853.42812.72733.4601
C32.44811.38461.40372.42572.79923.73452.15011.08932.16653.41293.88924.10624.6226
C42.83312.42111.40371.40372.42114.23863.40992.16241.08852.16243.40994.89214.8921
C52.44812.79922.42571.40371.38463.73453.88923.41292.16651.08932.15014.62264.1062
C61.42782.43982.79922.42111.38462.46973.42813.88853.40582.14451.09003.46012.7273
C71.40552.46973.73454.23863.73452.46972.69444.61175.32724.61172.69441.08721.0872
H82.17401.09002.15013.40993.88923.42812.69442.47254.30674.97854.32972.49583.7784
H93.43282.14451.08932.16243.41293.88854.61172.47252.49274.31264.97854.80045.5731
H103.92163.40582.16651.08852.16653.40585.32724.30672.49272.49274.30675.96435.9643
H113.43283.88853.41292.16241.08932.14454.61174.97854.31262.49272.47255.57314.8004
H122.17403.42813.88923.40992.15011.09002.69444.32974.97854.30672.47253.77842.4958
H132.17812.72734.10624.89214.62263.46011.08722.49584.80045.96435.57313.77841.8586
H142.17813.46014.62264.89214.10622.72731.08723.77845.57315.96434.80042.49581.8586

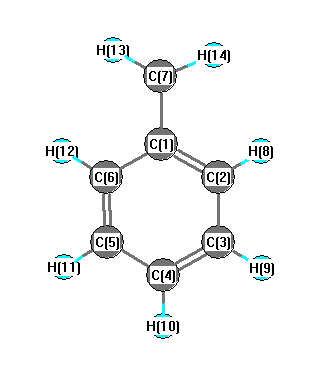
picture of benzyl radical state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 121.014 C1 C2 H8 118.802
C1 C6 C5 121.014 C1 C6 H12 118.802
C1 C7 H13 121.269 C1 C7 H14 121.269
C2 C1 C6 117.390 C2 C1 C7 121.305
C2 C3 C4 120.517 C2 C3 H9 119.704
C3 C2 H8 120.184 C3 C4 C5 119.549
C3 C4 H10 120.226 C4 C3 H9 119.779
C4 C5 C6 120.517 C4 C5 H11 119.779
C5 C1 C7 150.301 C5 C4 H10 120.226
C5 C6 H12 120.184 C6 C5 H11 119.704
H13 C7 H14 117.461
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability