return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H5CH2 (benzyl radical)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 2B1
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-270.591147
Energy at 298.15K-270.597996
HF Energy-270.308078
Nuclear repulsion energy260.118925
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3239 3075 16.59      
2 A1 3220 3057 5.75      
3 A1 3207 3044 14.46      
4 A1 3203 3040 0.30      
5 A1 1650 1566 1.02      
6 A1 1533 1455 6.17      
7 A1 1529 1451 0.79      
8 A1 1294 1228 2.44      
9 A1 1207 1145 0.01      
10 A1 1065 1011 1.45      
11 A1 1008 957 0.39      
12 A1 845 802 0.36      
13 A1 537 509 0.42      
14 A2 957 908 0.00      
15 A2 848 805 0.00      
16 A2 477 453 0.00      
17 A2 392 372 0.00      
18 B1 965 916 0.10      
19 B1 897 851 3.29      
20 B1 756 718 75.11      
21 B1 682 647 0.01      
22 B1 655 622 33.01      
23 B1 481 457 10.27      
24 B1 207 196 1.11      
25 B2 3302 3135 9.35      
26 B2 3226 3062 40.60      
27 B2 3208 3045 1.77      
28 B2 1635 1552 0.61      
29 B2 1511 1434 7.30      
30 B2 1377 1307 0.47      
31 B2 1351 1282 1.13      
32 B2 1199 1138 0.07      
33 B2 1142 1084 2.15      
34 B2 993 943 1.58      
35 B2 635 603 0.00      
36 B2 356 338 0.15      

Unscaled Zero Point Vibrational Energy (zpe) 25393.6 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 24103.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.18566 0.09016 0.06069

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 0.989
C2 0.000 1.214 0.252
C3 0.000 1.209 -1.133
C4 0.000 0.000 -1.836
C5 0.000 -1.209 -1.133
C6 0.000 -1.214 0.252
C7 0.000 0.000 2.401
H8 0.000 2.155 0.794
H9 0.000 2.148 -1.676
H10 0.000 0.000 -2.920
H11 0.000 -2.148 -1.676
H12 0.000 -2.155 0.794
H13 0.000 0.924 2.967
H14 0.000 -0.924 2.967

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 H8 H9 H10 H11 H12 H13 H14
C11.42062.44162.82422.44161.42061.41202.16343.42283.90893.42282.16342.18312.1831
C21.42061.38432.41482.79072.42892.46851.08522.14193.39643.87593.41242.73043.4561
C32.44161.38431.39822.41732.79073.73442.14601.08532.15783.40043.87594.10914.6209
C42.82422.41481.39821.39822.41484.23623.39922.15401.08462.15403.39924.89024.8902
C52.44162.79072.41731.39821.38433.73443.87593.40042.15781.08532.14604.62094.1091
C61.42062.42892.79072.41481.38432.46853.41243.87593.39642.14191.08523.45612.7304
C71.41202.46853.73444.23623.73442.46852.68804.60815.32094.60812.68801.08371.0837
H82.16341.08522.14603.39923.87593.41242.68802.46974.29364.96114.30932.49713.7685
H93.42282.14191.08532.15403.40043.87594.60812.46972.48234.29614.96114.80135.5672
H103.90893.39642.15781.08462.15783.39645.32094.29362.48232.48234.29365.95885.9588
H113.42283.87593.40042.15401.08532.14194.60814.96114.29612.48232.46975.56724.8013
H122.16343.41243.87593.39922.14601.08522.68804.30934.96114.29362.46973.76852.4971
H132.18312.73044.10914.89024.62093.45611.08372.49714.80135.95885.56723.76851.8485
H142.18313.45614.62094.89024.10912.73041.08373.76855.56725.95884.80132.49711.8485

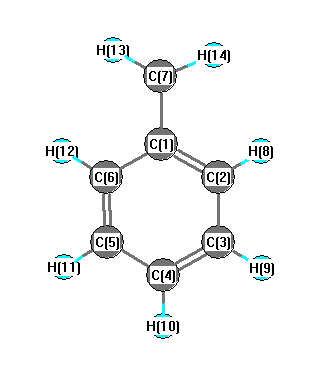
picture of benzyl radical state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 121.017 C1 C2 H8 118.784
C1 C6 C5 121.017 C1 C6 H12 118.784
C1 C7 H13 121.476 C1 C7 H14 121.476
C2 C1 C6 117.490 C2 C1 C7 121.255
C2 C3 C4 120.417 C2 C3 H9 119.811
C3 C2 H8 120.199 C3 C4 C5 119.643
C3 C4 H10 120.178 C4 C3 H9 119.772
C4 C5 C6 120.417 C4 C5 H11 119.772
C5 C1 C7 150.328 C5 C4 H10 120.178
C5 C6 H12 120.199 C6 C5 H11 119.811
H13 C7 H14 117.048
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C 0.162      
2 C -0.200      
3 C -0.155      
4 C -0.165      
5 C -0.155      
6 C -0.200      
7 C -0.408      
8 H 0.157      
9 H 0.160      
10 H 0.160      
11 H 0.160      
12 H 0.157      
13 H 0.163      
14 H 0.163      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 3.565 0.000 0.000
y 0.000 11.440 0.000
z 0.000 0.000 15.600


<r2> (average value of r2) Å2
<r2> 190.149
(<r2>)1/2 13.789