return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CHCH2CH3 (2-Butyl radical)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-157.568717
Energy at 298.15K-157.577302
HF Energy-157.403842
Nuclear repulsion energy121.987368
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3200 3038 31.28      
2 A 3158 2997 32.83      
3 A 3151 2991 36.72      
4 A 3144 2984 23.84      
5 A 3089 2932 29.77      
6 A 3080 2923 28.96      
7 A 3057 2902 24.65      
8 A 3004 2851 40.14      
9 A 2975 2824 37.38      
10 A 1556 1477 3.11      
11 A 1549 1470 5.46      
12 A 1540 1462 4.08      
13 A 1528 1450 5.29      
14 A 1525 1447 0.87      
15 A 1463 1388 2.27      
16 A 1458 1384 1.31      
17 A 1446 1373 0.30      
18 A 1348 1280 9.18      
19 A 1304 1238 0.06      
20 A 1211 1149 0.43      
21 A 1160 1101 0.14      
22 A 1105 1049 0.77      
23 A 1051 998 0.15      
24 A 1009 958 2.54      
25 A 1001 950 2.75      
26 A 866 822 0.22      
27 A 788 748 1.36      
28 A 435 413 2.66      
29 A 427 405 16.89      
30 A 266 253 1.18      
31 A 242 230 0.02      
32 A 126 120 0.04      
33 A 69 65 0.06      

Unscaled Zero Point Vibrational Energy (zpe) 26163.2 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 24834.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.88032 0.12131 0.11423

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.984 -0.047 -0.072
H2 2.067 -0.836 -0.828
H3 2.675 0.756 -0.341
H4 2.340 -0.479 0.877
C5 0.578 0.450 0.039
H6 0.417 1.464 0.400
C7 -0.571 -0.506 0.103
H8 -0.565 -1.033 1.073
H9 -0.433 -1.290 -0.653
C10 -1.934 0.167 -0.089
H11 -2.105 0.929 0.678
H12 -1.990 0.659 -1.065
H13 -2.748 -0.560 -0.027

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 H6 C7 H8 H9 C10 H11 H12 H13
C11.09601.09381.10181.49492.22702.60162.96282.77953.92304.26984.15584.7598
H21.09601.77311.76292.15013.08542.81683.25222.54704.18994.77364.32984.8889
H31.09381.77311.76682.15342.48023.51173.96263.73514.65334.89114.72215.5900
H41.10181.76291.76682.16082.77493.01232.96373.26974.42874.66674.87965.1686
C51.49492.15012.15342.16081.08781.49662.13922.12902.53082.79952.80303.4771
H62.22703.08542.48022.77491.08782.22362.76613.06872.72832.59302.93003.7809
C72.60162.81683.51173.01231.49662.22361.10371.09831.53192.17752.17562.1817
H82.96283.25223.96262.96372.13922.76611.10371.75022.16002.52503.07612.4904
H92.77952.54703.73513.26972.12903.06871.09831.75022.16643.08072.52782.5067
C103.92304.18994.65334.42872.53082.72831.53192.16002.16641.09451.09371.0939
H114.26984.77364.89114.66672.79952.59302.17752.52503.08071.09451.76781.7684
H124.15584.32984.72214.87962.80302.93002.17563.07612.52781.09371.76781.7710
H134.75984.88895.59005.16863.47713.78092.18172.49042.50671.09391.76841.7710

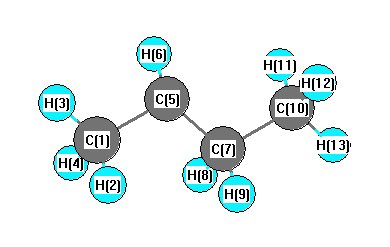
picture of 2-Butyl radical state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C5 H6 118.298 C1 C5 C7 120.839
H2 C1 H3 108.136 H2 C1 H4 106.667
H2 C1 C5 111.238 H3 C1 H4 107.167
H3 C1 C5 111.642 H4 C1 C5 111.743
C5 C7 H8 109.782 C5 C7 H9 109.295
C5 C7 C10 113.367 H6 C5 C7 117.854
C7 C10 H11 110.910 C7 C10 H12 110.816
C7 C10 H13 111.280 H8 C7 H9 105.276
H8 C7 C10 108.997 H9 C7 C10 109.810
H11 C10 H12 107.778 H11 C10 H13 107.811
H12 C10 H13 108.104
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.488      
2 H 0.155      
3 H 0.159      
4 H 0.155      
5 C -0.115      
6 H 0.141      
7 C -0.291      
8 H 0.146      
9 H 0.146      
10 C -0.462      
11 H 0.149      
12 H 0.155      
13 H 0.150      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 7.332 0.047 -0.055
y 0.047 6.064 0.092
z -0.055 0.092 5.286


<r2> (average value of r2) Å2
<r2> 116.628
(<r2>)1/2 10.799