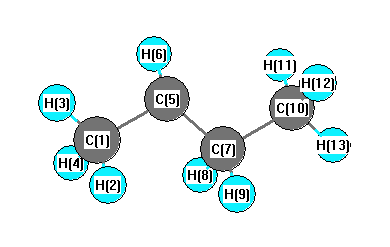
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3194 |
3038 |
29.09 |
|
|
|
| 2 |
A |
3152 |
2998 |
29.83 |
|
|
|
| 3 |
A |
3144 |
2990 |
34.47 |
|
|
|
| 4 |
A |
3138 |
2984 |
21.33 |
|
|
|
| 5 |
A |
3076 |
2925 |
28.10 |
|
|
|
| 6 |
A |
3069 |
2918 |
30.12 |
|
|
|
| 7 |
A |
3043 |
2894 |
25.15 |
|
|
|
| 8 |
A |
2992 |
2845 |
41.78 |
|
|
|
| 9 |
A |
2964 |
2819 |
36.34 |
|
|
|
| 10 |
A |
1533 |
1458 |
3.90 |
|
|
|
| 11 |
A |
1525 |
1451 |
6.98 |
|
|
|
| 12 |
A |
1516 |
1441 |
5.52 |
|
|
|
| 13 |
A |
1503 |
1430 |
6.87 |
|
|
|
| 14 |
A |
1498 |
1425 |
0.78 |
|
|
|
| 15 |
A |
1447 |
1377 |
1.46 |
|
|
|
| 16 |
A |
1437 |
1367 |
4.62 |
|
|
|
| 17 |
A |
1425 |
1355 |
0.45 |
|
|
|
| 18 |
A |
1327 |
1262 |
7.41 |
|
|
|
| 19 |
A |
1286 |
1223 |
0.06 |
|
|
|
| 20 |
A |
1200 |
1141 |
0.45 |
|
|
|
| 21 |
A |
1158 |
1101 |
0.20 |
|
|
|
| 22 |
A |
1086 |
1033 |
0.58 |
|
|
|
| 23 |
A |
1057 |
1005 |
0.26 |
|
|
|
| 24 |
A |
997 |
948 |
4.42 |
|
|
|
| 25 |
A |
984 |
936 |
2.72 |
|
|
|
| 26 |
A |
865 |
823 |
0.24 |
|
|
|
| 27 |
A |
775 |
737 |
1.82 |
|
|
|
| 28 |
A |
431 |
410 |
0.31 |
|
|
|
| 29 |
A |
405 |
385 |
20.51 |
|
|
|
| 30 |
A |
261 |
248 |
1.40 |
|
|
|
| 31 |
A |
239 |
227 |
0.04 |
|
|
|
| 32 |
A |
114 |
108 |
0.05 |
|
|
|
| 33 |
A |
64 |
61 |
0.05 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25952.9 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 24681.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.550 |
|
|
|
| 2 |
H |
0.173 |
|
|
|
| 3 |
H |
0.177 |
|
|
|
| 4 |
H |
0.174 |
|
|
|
| 5 |
C |
-0.115 |
|
|
|
| 6 |
H |
0.152 |
|
|
|
| 7 |
C |
-0.334 |
|
|
|
| 8 |
H |
0.166 |
|
|
|
| 9 |
H |
0.165 |
|
|
|
| 10 |
C |
-0.518 |
|
|
|
| 11 |
H |
0.168 |
|
|
|
| 12 |
H |
0.174 |
|
|
|
| 13 |
H |
0.168 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.069 |
-0.139 |
0.165 |
0.226 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-27.027 |
0.170 |
-0.041 |
| y |
0.170 |
-26.805 |
0.359 |
| z |
-0.041 |
0.359 |
-27.558 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.155 |
0.170 |
-0.041 |
| y |
0.170 |
0.488 |
0.359 |
| z |
-0.041 |
0.359 |
-0.643 |
|
| Polar |
|---|
| 3z2-r2 | -1.285 |
|---|
| x2-y2 | -0.222 |
|---|
| xy | 0.170 |
|---|
| xz | -0.041 |
|---|
| yz | 0.359 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.488 |
0.040 |
-0.048 |
| y |
0.040 |
6.148 |
0.078 |
| z |
-0.048 |
0.078 |
5.331 |
<r2> (average value of r
2) Å
2
| <r2> |
116.050 |
| (<r2>)1/2 |
10.773 |