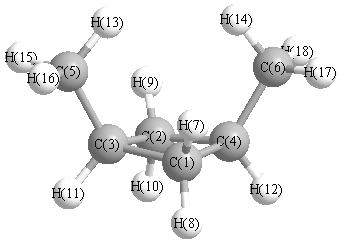
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3139 |
2985 |
105.42 |
|
|
|
| 2 |
A1 |
3128 |
2975 |
0.09 |
|
|
|
| 3 |
A1 |
3074 |
2924 |
44.00 |
|
|
|
| 4 |
A1 |
3063 |
2913 |
0.20 |
|
|
|
| 5 |
A1 |
3055 |
2905 |
8.96 |
|
|
|
| 6 |
A1 |
1531 |
1456 |
10.11 |
|
|
|
| 7 |
A1 |
1521 |
1447 |
5.85 |
|
|
|
| 8 |
A1 |
1441 |
1370 |
7.44 |
|
|
|
| 9 |
A1 |
1401 |
1332 |
0.43 |
|
|
|
| 10 |
A1 |
1212 |
1153 |
0.13 |
|
|
|
| 11 |
A1 |
1171 |
1114 |
0.07 |
|
|
|
| 12 |
A1 |
971 |
924 |
0.38 |
|
|
|
| 13 |
A1 |
907 |
862 |
0.42 |
|
|
|
| 14 |
A1 |
565 |
537 |
1.81 |
|
|
|
| 15 |
A1 |
432 |
410 |
0.06 |
|
|
|
| 16 |
A1 |
103 |
98 |
0.00 |
|
|
|
| 17 |
A2 |
3135 |
2981 |
0.00 |
|
|
|
| 18 |
A2 |
1523 |
1448 |
0.00 |
|
|
|
| 19 |
A2 |
1283 |
1221 |
0.00 |
|
|
|
| 20 |
A2 |
1243 |
1182 |
0.00 |
|
|
|
| 21 |
A2 |
1140 |
1085 |
0.00 |
|
|
|
| 22 |
A2 |
997 |
948 |
0.00 |
|
|
|
| 23 |
A2 |
918 |
873 |
0.00 |
|
|
|
| 24 |
A2 |
354 |
336 |
0.00 |
|
|
|
| 25 |
A2 |
233 |
221 |
0.00 |
|
|
|
| 26 |
B1 |
3135 |
2981 |
47.47 |
|
|
|
| 27 |
B1 |
3129 |
2975 |
47.08 |
|
|
|
| 28 |
B1 |
3061 |
2911 |
45.37 |
|
|
|
| 29 |
B1 |
1525 |
1450 |
5.34 |
|
|
|
| 30 |
B1 |
1500 |
1426 |
1.53 |
|
|
|
| 31 |
B1 |
1325 |
1260 |
0.24 |
|
|
|
| 32 |
B1 |
1184 |
1126 |
2.89 |
|
|
|
| 33 |
B1 |
923 |
878 |
0.07 |
|
|
|
| 34 |
B1 |
814 |
774 |
3.18 |
|
|
|
| 35 |
B1 |
286 |
272 |
0.08 |
|
|
|
| 36 |
B1 |
220 |
209 |
0.16 |
|
|
|
| 37 |
B2 |
3133 |
2980 |
28.53 |
|
|
|
| 38 |
B2 |
3067 |
2917 |
1.62 |
|
|
|
| 39 |
B2 |
3055 |
2906 |
48.62 |
|
|
|
| 40 |
B2 |
1522 |
1448 |
3.55 |
|
|
|
| 41 |
B2 |
1434 |
1364 |
2.64 |
|
|
|
| 42 |
B2 |
1392 |
1323 |
14.20 |
|
|
|
| 43 |
B2 |
1284 |
1221 |
0.74 |
|
|
|
| 44 |
B2 |
1269 |
1207 |
0.00 |
|
|
|
| 45 |
B2 |
1100 |
1046 |
0.11 |
|
|
|
| 46 |
B2 |
965 |
918 |
3.40 |
|
|
|
| 47 |
B2 |
789 |
750 |
0.17 |
|
|
|
| 48 |
B2 |
390 |
370 |
0.04 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 37017.5 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 35203.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.325 |
|
|
|
| 2 |
C |
-0.325 |
|
|
|
| 3 |
C |
-0.126 |
|
|
|
| 4 |
C |
-0.126 |
|
|
|
| 5 |
C |
-0.517 |
|
|
|
| 6 |
C |
-0.517 |
|
|
|
| 7 |
H |
0.155 |
|
|
|
| 8 |
H |
0.160 |
|
|
|
| 9 |
H |
0.155 |
|
|
|
| 10 |
H |
0.160 |
|
|
|
| 11 |
H |
0.157 |
|
|
|
| 12 |
H |
0.157 |
|
|
|
| 13 |
H |
0.161 |
|
|
|
| 14 |
H |
0.161 |
|
|
|
| 15 |
H |
0.168 |
|
|
|
| 16 |
H |
0.168 |
|
|
|
| 17 |
H |
0.168 |
|
|
|
| 18 |
H |
0.168 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-0.130 |
0.130 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.733 |
0.000 |
0.000 |
| y |
0.000 |
-40.158 |
0.000 |
| z |
0.000 |
0.000 |
-38.876 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.216 |
0.000 |
0.000 |
| y |
0.000 |
-0.854 |
0.000 |
| z |
0.000 |
0.000 |
1.070 |
|
| Polar |
|---|
| 3z2-r2 | 2.139 |
|---|
| x2-y2 | 0.425 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.833 |
0.000 |
0.000 |
| y |
0.000 |
10.324 |
0.000 |
| z |
0.000 |
0.000 |
8.352 |
<r2> (average value of r
2) Å
2
| <r2> |
197.078 |
| (<r2>)1/2 |
14.038 |