Jump to
S1C2
Vibrational Frequencies calculated at CID/6-31G*
Geometric Data calculated at CID/6-31G*
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CID/6-31G*
| | hartrees |
|---|
| Energy at 0K | -234.861628 |
| Energy at 298.15K | -234.875311 |
| HF Energy | -234.170055 |
| Nuclear repulsion energy | 250.697044 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3236 |
2989 |
115.25 |
|
|
|
| 2 |
A' |
3230 |
2984 |
21.31 |
|
|
|
| 3 |
A' |
3224 |
2978 |
22.46 |
|
|
|
| 4 |
A' |
3191 |
2947 |
19.63 |
|
|
|
| 5 |
A' |
3176 |
2934 |
12.27 |
|
|
|
| 6 |
A' |
3172 |
2930 |
9.35 |
|
|
|
| 7 |
A' |
3156 |
2915 |
26.25 |
|
|
|
| 8 |
A' |
3153 |
2912 |
26.06 |
|
|
|
| 9 |
A' |
1607 |
1485 |
4.80 |
|
|
|
| 10 |
A' |
1596 |
1474 |
2.16 |
|
|
|
| 11 |
A' |
1594 |
1472 |
5.72 |
|
|
|
| 12 |
A' |
1517 |
1402 |
8.61 |
|
|
|
| 13 |
A' |
1516 |
1400 |
1.02 |
|
|
|
| 14 |
A' |
1477 |
1365 |
1.43 |
|
|
|
| 15 |
A' |
1464 |
1352 |
6.97 |
|
|
|
| 16 |
A' |
1372 |
1268 |
1.52 |
|
|
|
| 17 |
A' |
1327 |
1226 |
0.58 |
|
|
|
| 18 |
A' |
1276 |
1178 |
0.10 |
|
|
|
| 19 |
A' |
1204 |
1112 |
0.29 |
|
|
|
| 20 |
A' |
1113 |
1028 |
0.13 |
|
|
|
| 21 |
A' |
1019 |
941 |
0.90 |
|
|
|
| 22 |
A' |
1009 |
932 |
0.68 |
|
|
|
| 23 |
A' |
917 |
847 |
0.94 |
|
|
|
| 24 |
A' |
852 |
787 |
0.99 |
|
|
|
| 25 |
A' |
683 |
631 |
1.62 |
|
|
|
| 26 |
A' |
458 |
423 |
0.06 |
|
|
|
| 27 |
A' |
344 |
318 |
0.10 |
|
|
|
| 28 |
A' |
117 |
108 |
0.01 |
|
|
|
| 29 |
A" |
3233 |
2986 |
37.78 |
|
|
|
| 30 |
A" |
3230 |
2983 |
29.42 |
|
|
|
| 31 |
A" |
3224 |
2978 |
9.19 |
|
|
|
| 32 |
A" |
3170 |
2928 |
63.57 |
|
|
|
| 33 |
A" |
1597 |
1475 |
2.43 |
|
|
|
| 34 |
A" |
1595 |
1473 |
0.42 |
|
|
|
| 35 |
A" |
1578 |
1458 |
3.83 |
|
|
|
| 36 |
A" |
1389 |
1283 |
0.60 |
|
|
|
| 37 |
A" |
1359 |
1256 |
0.00 |
|
|
|
| 38 |
A" |
1321 |
1220 |
0.08 |
|
|
|
| 39 |
A" |
1213 |
1121 |
0.53 |
|
|
|
| 40 |
A" |
1204 |
1112 |
0.10 |
|
|
|
| 41 |
A" |
1055 |
974 |
2.87 |
|
|
|
| 42 |
A" |
968 |
894 |
0.02 |
|
|
|
| 43 |
A" |
962 |
889 |
0.09 |
|
|
|
| 44 |
A" |
860 |
795 |
0.22 |
|
|
|
| 45 |
A" |
392 |
362 |
0.01 |
|
|
|
| 46 |
A" |
272 |
252 |
0.12 |
|
|
|
| 47 |
A" |
260 |
240 |
0.00 |
|
|
|
| 48 |
A" |
243 |
225 |
0.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 38561.6 cm
-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 35619.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CID/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.272 |
0.096 |
1.074 |
| C2 |
0.272 |
0.096 |
-1.074 |
| C3 |
-0.664 |
0.691 |
0.000 |
| C4 |
0.868 |
-0.842 |
0.000 |
| C5 |
-0.916 |
2.185 |
0.000 |
| C6 |
0.272 |
-2.239 |
0.000 |
| H7 |
0.998 |
0.832 |
1.418 |
| H8 |
-0.187 |
-0.369 |
1.946 |
| H9 |
0.998 |
0.832 |
-1.418 |
| H10 |
-0.187 |
-0.369 |
-1.946 |
| H11 |
-1.616 |
0.160 |
0.000 |
| H12 |
1.955 |
-0.912 |
0.000 |
| H13 |
0.025 |
2.734 |
0.000 |
| H14 |
-0.818 |
-2.205 |
0.000 |
| H15 |
-1.481 |
2.489 |
-0.881 |
| H16 |
-1.481 |
2.489 |
0.881 |
| H17 |
0.586 |
-2.797 |
-0.882 |
| H18 |
0.586 |
-2.797 |
0.882 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
| C1 | | 2.1470 | 1.5432 | 1.5449 | 2.6317 | 2.5697 | 1.0899 | 1.0895 | 2.6972 | 3.0890 | 2.1721 | 2.2368 | 2.8594 | 2.7626 | 3.5524 | 2.9726 | 3.5056 | 2.9162 |
C2 | 2.1470 | | 1.5432 | 1.5449 | 2.6317 | 2.5697 | 2.6972 | 3.0890 | 1.0899 | 1.0895 | 2.1721 | 2.2368 | 2.8594 | 2.7626 | 2.9726 | 3.5524 | 2.9162 | 3.5056 | C3 | 1.5432 | 1.5432 | | 2.1668 | 1.5147 | 3.0758 | 2.1887 | 2.2664 | 2.1887 | 2.2664 | 1.0902 | 3.0707 | 2.1563 | 2.9002 | 2.1627 | 2.1627 | 3.8087 | 3.8087 | C4 | 1.5449 | 1.5449 | 2.1668 | | 3.5129 | 1.5194 | 2.1969 | 2.2633 | 2.1969 | 2.2633 | 2.6782 | 1.0895 | 3.6740 | 2.1682 | 4.1698 | 4.1698 | 2.1636 | 2.1636 | C5 | 2.6317 | 2.6317 | 1.5147 | 3.5129 | | 4.5803 | 2.7392 | 3.2922 | 2.7392 | 3.2922 | 2.1420 | 4.2231 | 1.0901 | 4.3907 | 1.0899 | 1.0899 | 5.2774 | 5.2774 | C6 | 2.5697 | 2.5697 | 3.0758 | 1.5194 | 4.5803 | | 3.4593 | 2.7373 | 3.4593 | 2.7373 | 3.0526 | 2.1436 | 4.9796 | 1.0899 | 5.1190 | 5.1190 | 1.0897 | 1.0897 | H7 | 1.0899 | 2.6972 | 2.1887 | 2.1969 | 2.7392 | 3.4593 | | 1.7679 | 2.8352 | 3.7627 | 3.0483 | 2.4428 | 2.5644 | 3.8116 | 3.7650 | 3.0297 | 4.3156 | 3.6913 | H8 | 1.0895 | 3.0890 | 2.2664 | 2.2633 | 3.2922 | 2.7373 | 1.7679 | | 3.7627 | 3.8913 | 2.4711 | 2.9446 | 3.6692 | 2.7484 | 4.2230 | 3.3132 | 3.8060 | 2.7614 | H9 | 2.6972 | 1.0899 | 2.1887 | 2.1969 | 2.7392 | 3.4593 | 2.8352 | 3.7627 | | 1.7679 | 3.0483 | 2.4428 | 2.5644 | 3.8116 | 3.0297 | 3.7650 | 3.6913 | 4.3156 | H10 | 3.0890 | 1.0895 | 2.2664 | 2.2633 | 3.2922 | 2.7373 | 3.7627 | 3.8913 | 1.7679 | | 2.4711 | 2.9446 | 3.6692 | 2.7484 | 3.3132 | 4.2230 | 2.7614 | 3.8060 | H11 | 2.1721 | 2.1721 | 1.0902 | 2.6782 | 2.1420 | 3.0526 | 3.0483 | 2.4711 | 3.0483 | 2.4711 | | 3.7286 | 3.0528 | 2.4961 | 2.4938 | 2.4938 | 3.7908 | 3.7908 | H12 | 2.2368 | 2.2368 | 3.0707 | 1.0895 | 4.2231 | 2.1436 | 2.4428 | 2.9446 | 2.4428 | 2.9446 | 3.7286 | | 4.1259 | 3.0596 | 4.9145 | 4.9145 | 2.4910 | 2.4910 | H13 | 2.8594 | 2.8594 | 2.1563 | 3.6740 | 1.0901 | 4.9796 | 2.5644 | 3.6692 | 2.5644 | 3.6692 | 3.0528 | 4.1259 | | 5.0108 | 1.7619 | 1.7619 | 5.6294 | 5.6294 | H14 | 2.7626 | 2.7626 | 2.9002 | 2.1682 | 4.3907 | 1.0899 | 3.8116 | 2.7484 | 3.8116 | 2.7484 | 2.4961 | 3.0596 | 5.0108 | | 4.8220 | 4.8220 | 1.7602 | 1.7602 | H15 | 3.5524 | 2.9726 | 2.1627 | 4.1698 | 1.0899 | 5.1190 | 3.7650 | 4.2230 | 3.0297 | 3.3132 | 2.4938 | 4.9145 | 1.7619 | 4.8220 | | 1.7621 | 5.6760 | 5.9434 | H16 | 2.9726 | 3.5524 | 2.1627 | 4.1698 | 1.0899 | 5.1190 | 3.0297 | 3.3132 | 3.7650 | 4.2230 | 2.4938 | 4.9145 | 1.7619 | 4.8220 | 1.7621 | | 5.9434 | 5.6760 | H17 | 3.5056 | 2.9162 | 3.8087 | 2.1636 | 5.2774 | 1.0897 | 4.3156 | 3.8060 | 3.6913 | 2.7614 | 3.7908 | 2.4910 | 5.6294 | 1.7602 | 5.6760 | 5.9434 | | 1.7630 | H18 | 2.9162 | 3.5056 | 3.8087 | 2.1636 | 5.2774 | 1.0897 | 3.6913 | 2.7614 | 4.3156 | 3.8060 | 3.7908 | 2.4910 | 5.6294 | 1.7602 | 5.9434 | 5.6760 | 1.7630 | |
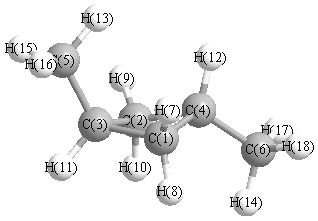 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C3 |
C2 |
88.159 |
|
C1 |
C3 |
C5 |
118.779 |
| C1 |
C3 |
H11 |
109.957 |
|
C1 |
C4 |
C2 |
88.033 |
| C1 |
C4 |
C6 |
113.978 |
|
C1 |
C4 |
H12 |
115.132 |
| C2 |
C3 |
C5 |
118.779 |
|
C2 |
C3 |
H11 |
109.957 |
| C2 |
C4 |
C6 |
113.978 |
|
C2 |
C4 |
H12 |
115.132 |
| C3 |
C1 |
C4 |
89.123 |
|
C3 |
C1 |
H7 |
111.283 |
| C3 |
C1 |
H8 |
117.799 |
|
C3 |
C2 |
C4 |
89.123 |
| C3 |
C2 |
H9 |
111.283 |
|
C3 |
C2 |
H10 |
117.799 |
| C3 |
C5 |
H13 |
110.699 |
|
C3 |
C5 |
H15 |
111.220 |
| C3 |
C5 |
H16 |
111.220 |
|
C4 |
C1 |
H7 |
111.818 |
| C4 |
C1 |
H8 |
117.385 |
|
C4 |
C2 |
H9 |
111.818 |
| C4 |
C2 |
H10 |
117.385 |
|
C4 |
C6 |
H14 |
111.323 |
| C4 |
C6 |
H17 |
110.968 |
|
C4 |
C6 |
H18 |
110.968 |
| C5 |
C3 |
H11 |
109.553 |
|
C6 |
C4 |
H12 |
109.392 |
| H7 |
C1 |
H8 |
108.421 |
|
H9 |
C2 |
H10 |
108.421 |
| H13 |
C5 |
H15 |
107.842 |
|
H13 |
C5 |
H16 |
107.842 |
| H14 |
C6 |
H17 |
107.718 |
|
H14 |
C6 |
H18 |
107.718 |
| H15 |
C5 |
H16 |
107.869 |
|
H17 |
C6 |
H18 |
107.991 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
