Jump to
S1C2
Vibrational Frequencies calculated at B3PW91/6-31G*
Geometric Data calculated at B3PW91/6-31G*
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B3PW91/6-31G*
| | hartrees |
|---|
| Energy at 0K | -235.764859 |
| Energy at 298.15K | -235.778219 |
| HF Energy | -235.764859 |
| Nuclear repulsion energy | 249.326013 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3123 |
2988 |
119.02 |
|
|
|
| 2 |
A' |
3118 |
2983 |
12.33 |
|
|
|
| 3 |
A' |
3113 |
2979 |
27.38 |
|
|
|
| 4 |
A' |
3075 |
2941 |
18.46 |
|
|
|
| 5 |
A' |
3062 |
2929 |
9.19 |
|
|
|
| 6 |
A' |
3058 |
2926 |
13.42 |
|
|
|
| 7 |
A' |
3043 |
2911 |
34.70 |
|
|
|
| 8 |
A' |
3042 |
2911 |
26.42 |
|
|
|
| 9 |
A' |
1527 |
1461 |
9.75 |
|
|
|
| 10 |
A' |
1522 |
1456 |
1.63 |
|
|
|
| 11 |
A' |
1517 |
1451 |
2.03 |
|
|
|
| 12 |
A' |
1434 |
1372 |
6.76 |
|
|
|
| 13 |
A' |
1431 |
1369 |
1.27 |
|
|
|
| 14 |
A' |
1395 |
1335 |
0.59 |
|
|
|
| 15 |
A' |
1378 |
1318 |
11.17 |
|
|
|
| 16 |
A' |
1293 |
1237 |
1.80 |
|
|
|
| 17 |
A' |
1265 |
1210 |
0.35 |
|
|
|
| 18 |
A' |
1211 |
1158 |
0.27 |
|
|
|
| 19 |
A' |
1147 |
1097 |
0.33 |
|
|
|
| 20 |
A' |
1062 |
1016 |
0.50 |
|
|
|
| 21 |
A' |
971 |
929 |
1.21 |
|
|
|
| 22 |
A' |
952 |
911 |
1.04 |
|
|
|
| 23 |
A' |
876 |
838 |
1.28 |
|
|
|
| 24 |
A' |
813 |
777 |
1.05 |
|
|
|
| 25 |
A' |
647 |
619 |
1.81 |
|
|
|
| 26 |
A' |
428 |
409 |
0.15 |
|
|
|
| 27 |
A' |
324 |
310 |
0.13 |
|
|
|
| 28 |
A' |
92 |
88 |
0.01 |
|
|
|
| 29 |
A" |
3121 |
2986 |
36.32 |
|
|
|
| 30 |
A" |
3118 |
2983 |
27.73 |
|
|
|
| 31 |
A" |
3114 |
2979 |
8.39 |
|
|
|
| 32 |
A" |
3059 |
2927 |
72.22 |
|
|
|
| 33 |
A" |
1524 |
1458 |
3.79 |
|
|
|
| 34 |
A" |
1522 |
1456 |
0.34 |
|
|
|
| 35 |
A" |
1495 |
1430 |
3.08 |
|
|
|
| 36 |
A" |
1308 |
1251 |
0.83 |
|
|
|
| 37 |
A" |
1284 |
1228 |
0.01 |
|
|
|
| 38 |
A" |
1244 |
1190 |
0.12 |
|
|
|
| 39 |
A" |
1155 |
1105 |
0.78 |
|
|
|
| 40 |
A" |
1144 |
1094 |
0.03 |
|
|
|
| 41 |
A" |
1007 |
963 |
3.85 |
|
|
|
| 42 |
A" |
923 |
883 |
0.00 |
|
|
|
| 43 |
A" |
916 |
876 |
0.00 |
|
|
|
| 44 |
A" |
815 |
780 |
0.35 |
|
|
|
| 45 |
A" |
375 |
359 |
0.01 |
|
|
|
| 46 |
A" |
259 |
248 |
0.15 |
|
|
|
| 47 |
A" |
256 |
245 |
0.00 |
|
|
|
| 48 |
A" |
238 |
228 |
0.07 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36896.4 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 35298.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.255 |
0.085 |
1.080 |
| C2 |
0.255 |
0.085 |
-1.080 |
| C3 |
-0.659 |
0.716 |
0.000 |
| C4 |
0.849 |
-0.859 |
0.000 |
| C5 |
-0.859 |
2.222 |
0.000 |
| C6 |
0.255 |
-2.262 |
0.000 |
| H7 |
0.997 |
0.803 |
1.450 |
| H8 |
-0.230 |
-0.386 |
1.943 |
| H9 |
0.997 |
0.803 |
-1.450 |
| H10 |
-0.230 |
-0.386 |
-1.943 |
| H11 |
-1.637 |
0.218 |
0.000 |
| H12 |
1.943 |
-0.928 |
0.000 |
| H13 |
0.107 |
2.744 |
0.000 |
| H14 |
-0.842 |
-2.233 |
0.000 |
| H15 |
-1.416 |
2.551 |
-0.886 |
| H16 |
-1.416 |
2.551 |
0.886 |
| H17 |
0.572 |
-2.825 |
-0.887 |
| H18 |
0.572 |
-2.825 |
0.887 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
| C1 | | 2.1604 | 1.5501 | 1.5524 | 2.6412 | 2.5835 | 1.0968 | 1.0966 | 2.7331 | 3.0983 | 2.1835 | 2.2452 | 2.8745 | 2.7826 | 3.5698 | 2.9858 | 3.5265 | 2.9334 |
C2 | 2.1604 | | 1.5501 | 1.5524 | 2.6412 | 2.5835 | 2.7331 | 3.0983 | 1.0968 | 1.0966 | 2.1835 | 2.2452 | 2.8745 | 2.7826 | 2.9858 | 3.5698 | 2.9334 | 3.5265 | C3 | 1.5501 | 1.5501 | | 2.1810 | 1.5188 | 3.1158 | 2.2032 | 2.2751 | 2.2032 | 2.2751 | 1.0976 | 3.0785 | 2.1678 | 2.9551 | 2.1735 | 2.1735 | 3.8528 | 3.8528 | C4 | 1.5524 | 1.5524 | 2.1810 | | 3.5227 | 1.5231 | 2.2113 | 2.2725 | 2.2113 | 2.2725 | 2.7096 | 1.0968 | 3.6793 | 2.1782 | 4.1887 | 4.1887 | 2.1742 | 2.1742 | C5 | 2.6412 | 2.6412 | 1.5188 | 3.5227 | | 4.6204 | 2.7495 | 3.3126 | 2.7495 | 3.3126 | 2.1497 | 4.2158 | 1.0975 | 4.4552 | 1.0972 | 1.0972 | 5.3204 | 5.3204 | C6 | 2.5835 | 2.5835 | 3.1158 | 1.5231 | 4.6204 | | 3.4713 | 2.7444 | 3.4713 | 2.7444 | 3.1202 | 2.1515 | 5.0087 | 1.0974 | 5.1718 | 5.1718 | 1.0970 | 1.0970 | H7 | 1.0968 | 2.7331 | 2.2032 | 2.2113 | 2.7495 | 3.4713 | | 1.7785 | 2.9008 | 3.7997 | 3.0635 | 2.4487 | 2.5815 | 3.8345 | 3.7863 | 3.0325 | 4.3368 | 3.6964 | H8 | 1.0966 | 3.0983 | 2.2751 | 2.2725 | 3.3126 | 2.7444 | 1.7785 | | 3.7997 | 3.8867 | 2.4743 | 2.9655 | 3.6999 | 2.7499 | 4.2472 | 3.3395 | 3.8212 | 2.7765 | H9 | 2.7331 | 1.0968 | 2.2032 | 2.2113 | 2.7495 | 3.4713 | 2.9008 | 3.7997 | | 1.7785 | 3.0635 | 2.4487 | 2.5815 | 3.8345 | 3.0325 | 3.7863 | 3.6964 | 4.3368 | H10 | 3.0983 | 1.0966 | 2.2751 | 2.2725 | 3.3126 | 2.7444 | 3.7997 | 3.8867 | 1.7785 | | 2.4743 | 2.9655 | 3.6999 | 2.7499 | 3.3395 | 4.2472 | 2.7765 | 3.8212 | H11 | 2.1835 | 2.1835 | 1.0976 | 2.7096 | 2.1497 | 3.1202 | 3.0635 | 2.4743 | 3.0635 | 2.4743 | | 3.7597 | 3.0696 | 2.5774 | 2.5053 | 2.5053 | 3.8640 | 3.8640 | H12 | 2.2452 | 2.2452 | 3.0785 | 1.0968 | 4.2158 | 2.1515 | 2.4487 | 2.9655 | 2.4487 | 2.9655 | 3.7597 | | 4.1060 | 3.0756 | 4.9167 | 4.9167 | 2.5033 | 2.5033 | H13 | 2.8745 | 2.8745 | 2.1678 | 3.6793 | 1.0975 | 5.0087 | 2.5815 | 3.6999 | 2.5815 | 3.6999 | 3.0696 | 4.1060 | | 5.0670 | 1.7720 | 1.7720 | 5.6588 | 5.6588 | H14 | 2.7826 | 2.7826 | 2.9551 | 2.1782 | 4.4552 | 1.0974 | 3.8345 | 2.7499 | 3.8345 | 2.7499 | 2.5774 | 3.0756 | 5.0670 | | 4.8995 | 4.8995 | 1.7705 | 1.7705 | H15 | 3.5698 | 2.9858 | 2.1735 | 4.1887 | 1.0972 | 5.1718 | 3.7863 | 4.2472 | 3.0325 | 3.3395 | 2.5053 | 4.9167 | 1.7720 | 4.8995 | | 1.7719 | 5.7321 | 6.0000 | H16 | 2.9858 | 3.5698 | 2.1735 | 4.1887 | 1.0972 | 5.1718 | 3.0325 | 3.3395 | 3.7863 | 4.2472 | 2.5053 | 4.9167 | 1.7720 | 4.8995 | 1.7719 | | 6.0000 | 5.7321 | H17 | 3.5265 | 2.9334 | 3.8528 | 2.1742 | 5.3204 | 1.0970 | 4.3368 | 3.8212 | 3.6964 | 2.7765 | 3.8640 | 2.5033 | 5.6588 | 1.7705 | 5.7321 | 6.0000 | | 1.7734 | H18 | 2.9334 | 3.5265 | 3.8528 | 2.1742 | 5.3204 | 1.0970 | 3.6964 | 2.7765 | 4.3368 | 3.8212 | 3.8640 | 2.5033 | 5.6588 | 1.7705 | 6.0000 | 5.7321 | 1.7734 | |
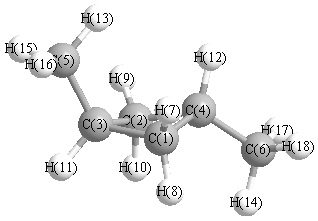 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C3 |
C2 |
88.354 |
|
C1 |
C3 |
C5 |
118.778 |
| C1 |
C3 |
H11 |
109.940 |
|
C1 |
C4 |
C2 |
88.186 |
| C1 |
C4 |
C6 |
114.285 |
|
C1 |
C4 |
H12 |
114.796 |
| C2 |
C3 |
C5 |
118.778 |
|
C2 |
C3 |
H11 |
109.940 |
| C2 |
C4 |
C6 |
114.285 |
|
C2 |
C4 |
H12 |
114.796 |
| C3 |
C1 |
C4 |
89.336 |
|
C3 |
C1 |
H7 |
111.544 |
| C3 |
C1 |
H8 |
117.523 |
|
C3 |
C2 |
C4 |
89.336 |
| C3 |
C2 |
H9 |
111.544 |
|
C3 |
C2 |
H10 |
117.523 |
| C3 |
C5 |
H13 |
110.881 |
|
C3 |
C5 |
H15 |
111.357 |
| C3 |
C5 |
H16 |
111.357 |
|
C4 |
C1 |
H7 |
112.020 |
| C4 |
C1 |
H8 |
117.118 |
|
C4 |
C2 |
H9 |
112.020 |
| C4 |
C2 |
H10 |
117.118 |
|
C4 |
C6 |
H14 |
111.410 |
| C4 |
C6 |
H17 |
111.120 |
|
C4 |
C6 |
H18 |
111.120 |
| C5 |
C3 |
H11 |
109.444 |
|
C6 |
C4 |
H12 |
109.339 |
| H7 |
C1 |
H8 |
108.357 |
|
H9 |
C2 |
H10 |
108.357 |
| H13 |
C5 |
H15 |
107.689 |
|
H13 |
C5 |
H16 |
107.689 |
| H14 |
C6 |
H17 |
107.575 |
|
H14 |
C6 |
H18 |
107.575 |
| H15 |
C5 |
H16 |
107.693 |
|
H17 |
C6 |
H18 |
107.862 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.322 |
|
|
|
| 2 |
C |
-0.322 |
|
|
|
| 3 |
C |
-0.115 |
|
|
|
| 4 |
C |
-0.132 |
|
|
|
| 5 |
C |
-0.515 |
|
|
|
| 6 |
C |
-0.512 |
|
|
|
| 7 |
H |
0.157 |
|
|
|
| 8 |
H |
0.156 |
|
|
|
| 9 |
H |
0.157 |
|
|
|
| 10 |
H |
0.156 |
|
|
|
| 11 |
H |
0.155 |
|
|
|
| 12 |
H |
0.151 |
|
|
|
| 13 |
H |
0.159 |
|
|
|
| 14 |
H |
0.163 |
|
|
|
| 15 |
H |
0.166 |
|
|
|
| 16 |
H |
0.166 |
|
|
|
| 17 |
H |
0.166 |
|
|
|
| 18 |
H |
0.166 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.020 |
-0.001 |
0.000 |
0.020 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.914 |
0.358 |
0.000 |
| y |
0.358 |
-39.909 |
0.000 |
| z |
0.000 |
0.000 |
-39.660 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.871 |
0.358 |
0.000 |
| y |
0.358 |
-0.622 |
0.000 |
| z |
0.000 |
0.000 |
-0.249 |
|
| Polar |
|---|
| 3z2-r2 | -0.498 |
|---|
| x2-y2 | 0.996 |
|---|
| xy | 0.358 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.499 |
-0.560 |
0.000 |
| y |
-0.560 |
10.092 |
0.000 |
| z |
0.000 |
0.000 |
8.818 |
<r2> (average value of r
2) Å
2
| <r2> |
191.954 |
| (<r2>)1/2 |
13.855 |
