Jump to
S1C2
Vibrational Frequencies calculated at QCISD/6-31G*
Geometric Data calculated at QCISD/6-31G*
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/6-31G*
| | hartrees |
|---|
| Energy at 0K | -235.026949 |
| Energy at 298.15K | -235.040457 |
| HF Energy | -234.168869 |
| Nuclear repulsion energy | 249.417095 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3130 |
2981 |
101.04 |
|
|
|
| 2 |
A' |
3126 |
2977 |
24.61 |
|
|
|
| 3 |
A' |
3118 |
2970 |
23.97 |
|
|
|
| 4 |
A' |
3083 |
2936 |
21.80 |
|
|
|
| 5 |
A' |
3069 |
2923 |
13.37 |
|
|
|
| 6 |
A' |
3065 |
2919 |
9.46 |
|
|
|
| 7 |
A' |
3053 |
2907 |
26.77 |
|
|
|
| 8 |
A' |
3050 |
2904 |
22.89 |
|
|
|
| 9 |
A' |
1556 |
1481 |
5.85 |
|
|
|
| 10 |
A' |
1546 |
1473 |
1.15 |
|
|
|
| 11 |
A' |
1544 |
1471 |
5.33 |
|
|
|
| 12 |
A' |
1465 |
1395 |
8.53 |
|
|
|
| 13 |
A' |
1463 |
1393 |
0.61 |
|
|
|
| 14 |
A' |
1428 |
1360 |
0.78 |
|
|
|
| 15 |
A' |
1414 |
1346 |
7.63 |
|
|
|
| 16 |
A' |
1318 |
1255 |
1.79 |
|
|
|
| 17 |
A' |
1282 |
1221 |
0.51 |
|
|
|
| 18 |
A' |
1235 |
1176 |
0.08 |
|
|
|
| 19 |
A' |
1171 |
1115 |
0.29 |
|
|
|
| 20 |
A' |
1079 |
1027 |
0.17 |
|
|
|
| 21 |
A' |
988 |
940 |
0.99 |
|
|
|
| 22 |
A' |
980 |
933 |
0.64 |
|
|
|
| 23 |
A' |
888 |
846 |
0.92 |
|
|
|
| 24 |
A' |
825 |
786 |
0.90 |
|
|
|
| 25 |
A' |
662 |
631 |
1.68 |
|
|
|
| 26 |
A' |
446 |
425 |
0.05 |
|
|
|
| 27 |
A' |
336 |
320 |
0.11 |
|
|
|
| 28 |
A' |
119 |
113 |
0.01 |
|
|
|
| 29 |
A" |
3129 |
2979 |
35.99 |
|
|
|
| 30 |
A" |
3126 |
2977 |
27.97 |
|
|
|
| 31 |
A" |
3117 |
2969 |
10.77 |
|
|
|
| 32 |
A" |
3065 |
2919 |
59.87 |
|
|
|
| 33 |
A" |
1547 |
1473 |
2.49 |
|
|
|
| 34 |
A" |
1545 |
1472 |
0.45 |
|
|
|
| 35 |
A" |
1528 |
1455 |
3.35 |
|
|
|
| 36 |
A" |
1337 |
1273 |
0.54 |
|
|
|
| 37 |
A" |
1309 |
1247 |
0.00 |
|
|
|
| 38 |
A" |
1267 |
1207 |
0.06 |
|
|
|
| 39 |
A" |
1170 |
1114 |
0.37 |
|
|
|
| 40 |
A" |
1162 |
1107 |
0.17 |
|
|
|
| 41 |
A" |
1019 |
970 |
2.75 |
|
|
|
| 42 |
A" |
933 |
889 |
0.01 |
|
|
|
| 43 |
A" |
928 |
884 |
0.08 |
|
|
|
| 44 |
A" |
833 |
794 |
0.24 |
|
|
|
| 45 |
A" |
379 |
361 |
0.01 |
|
|
|
| 46 |
A" |
264 |
251 |
0.12 |
|
|
|
| 47 |
A" |
252 |
240 |
0.00 |
|
|
|
| 48 |
A" |
237 |
226 |
0.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 37291.8 cm
-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 35513.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.267 |
0.093 |
1.081 |
| C2 |
0.267 |
0.093 |
-1.081 |
| C3 |
-0.667 |
0.700 |
0.000 |
| C4 |
0.864 |
-0.850 |
0.000 |
| C5 |
-0.898 |
2.204 |
0.000 |
| C6 |
0.267 |
-2.255 |
0.000 |
| H7 |
1.001 |
0.827 |
1.441 |
| H8 |
-0.207 |
-0.380 |
1.952 |
| H9 |
1.001 |
0.827 |
-1.441 |
| H10 |
-0.207 |
-0.380 |
-1.952 |
| H11 |
-1.635 |
0.179 |
0.000 |
| H12 |
1.960 |
-0.918 |
0.000 |
| H13 |
0.061 |
2.741 |
0.000 |
| H14 |
-0.831 |
-2.217 |
0.000 |
| H15 |
-1.462 |
2.518 |
-0.889 |
| H16 |
-1.462 |
2.518 |
0.889 |
| H17 |
0.585 |
-2.815 |
-0.889 |
| H18 |
0.585 |
-2.815 |
0.889 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
| C1 | | 2.1625 | 1.5522 | 1.5542 | 2.6422 | 2.5849 | 1.0988 | 1.0985 | 2.7279 | 3.1060 | 2.1896 | 2.2490 | 2.8676 | 2.7774 | 3.5710 | 2.9846 | 3.5279 | 2.9324 |
C2 | 2.1625 | | 1.5522 | 1.5542 | 2.6422 | 2.5849 | 2.7279 | 3.1060 | 1.0988 | 1.0985 | 2.1896 | 2.2490 | 2.8676 | 2.7774 | 2.9846 | 3.5710 | 2.9324 | 3.5279 | C3 | 1.5522 | 1.5522 | | 2.1784 | 1.5216 | 3.0987 | 2.2082 | 2.2775 | 2.2082 | 2.2775 | 1.0994 | 3.0852 | 2.1670 | 2.9218 | 2.1746 | 2.1746 | 3.8361 | 3.8361 | C4 | 1.5542 | 1.5542 | 2.1784 | | 3.5259 | 1.5256 | 2.2162 | 2.2750 | 2.2162 | 2.2750 | 2.7022 | 1.0985 | 3.6804 | 2.1772 | 4.1889 | 4.1889 | 2.1750 | 2.1750 | C5 | 2.6422 | 2.6422 | 1.5216 | 3.5259 | | 4.6082 | 2.7531 | 3.3112 | 2.7531 | 3.3112 | 2.1547 | 4.2326 | 1.0988 | 4.4215 | 1.0988 | 1.0988 | 5.3088 | 5.3088 | C6 | 2.5849 | 2.5849 | 3.0987 | 1.5256 | 4.6082 | | 3.4806 | 2.7470 | 3.4806 | 2.7470 | 3.0888 | 2.1568 | 5.0001 | 1.0990 | 5.1537 | 5.1537 | 1.0985 | 1.0985 | H7 | 1.0988 | 2.7279 | 2.2082 | 2.2162 | 2.7531 | 3.4806 | | 1.7829 | 2.8829 | 3.7988 | 3.0737 | 2.4584 | 2.5741 | 3.8347 | 3.7892 | 3.0387 | 4.3448 | 3.7079 | H8 | 1.0985 | 3.1060 | 2.2775 | 2.2750 | 3.3112 | 2.7470 | 1.7829 | | 3.7988 | 3.9031 | 2.4820 | 2.9655 | 3.6912 | 2.7517 | 4.2480 | 3.3327 | 3.8247 | 2.7722 | H9 | 2.7279 | 1.0988 | 2.2082 | 2.2162 | 2.7531 | 3.4806 | 2.8829 | 3.7988 | | 1.7829 | 3.0737 | 2.4584 | 2.5741 | 3.8347 | 3.0387 | 3.7892 | 3.7079 | 4.3448 | H10 | 3.1060 | 1.0985 | 2.2775 | 2.2750 | 3.3112 | 2.7470 | 3.7988 | 3.9031 | 1.7829 | | 2.4820 | 2.9655 | 3.6912 | 2.7517 | 3.3327 | 4.2480 | 2.7722 | 3.8247 | H11 | 2.1896 | 2.1896 | 1.0994 | 2.7022 | 2.1547 | 3.0888 | 3.0737 | 2.4820 | 3.0737 | 2.4820 | | 3.7586 | 3.0723 | 2.5274 | 2.5083 | 2.5083 | 3.8322 | 3.8322 | H12 | 2.2490 | 2.2490 | 3.0852 | 1.0985 | 4.2326 | 2.1568 | 2.4584 | 2.9655 | 2.4584 | 2.9655 | 3.7586 | | 4.1230 | 3.0786 | 4.9306 | 4.9306 | 2.5065 | 2.5065 | H13 | 2.8676 | 2.8676 | 2.1670 | 3.6804 | 1.0988 | 5.0001 | 2.5741 | 3.6912 | 2.5741 | 3.6912 | 3.0723 | 4.1230 | | 5.0380 | 1.7772 | 1.7772 | 5.6519 | 5.6519 | H14 | 2.7774 | 2.7774 | 2.9218 | 2.1772 | 4.4215 | 1.0990 | 3.8347 | 2.7517 | 3.8347 | 2.7517 | 2.5274 | 3.0786 | 5.0380 | | 4.8593 | 4.8593 | 1.7760 | 1.7760 | H15 | 3.5710 | 2.9846 | 2.1746 | 4.1889 | 1.0988 | 5.1537 | 3.7892 | 4.2480 | 3.0387 | 3.3327 | 2.5083 | 4.9306 | 1.7772 | 4.8593 | | 1.7777 | 5.7131 | 5.9834 | H16 | 2.9846 | 3.5710 | 2.1746 | 4.1889 | 1.0988 | 5.1537 | 3.0387 | 3.3327 | 3.7892 | 4.2480 | 2.5083 | 4.9306 | 1.7772 | 4.8593 | 1.7777 | | 5.9834 | 5.7131 | H17 | 3.5279 | 2.9324 | 3.8361 | 2.1750 | 5.3088 | 1.0985 | 4.3448 | 3.8247 | 3.7079 | 2.7722 | 3.8322 | 2.5065 | 5.6519 | 1.7760 | 5.7131 | 5.9834 | | 1.7790 | H18 | 2.9324 | 3.5279 | 3.8361 | 2.1750 | 5.3088 | 1.0985 | 3.7079 | 2.7722 | 4.3448 | 3.8247 | 3.8322 | 2.5065 | 5.6519 | 1.7760 | 5.9834 | 5.7131 | 1.7790 | |
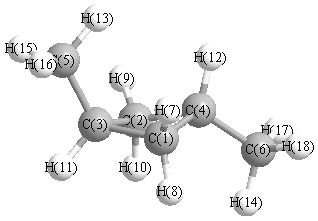 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C3 |
C2 |
88.308 |
|
C1 |
C3 |
C5 |
118.536 |
| C1 |
C3 |
H11 |
110.163 |
|
C1 |
C4 |
C2 |
88.167 |
| C1 |
C4 |
C6 |
114.137 |
|
C1 |
C4 |
H12 |
114.862 |
| C2 |
C3 |
C5 |
118.536 |
|
C2 |
C3 |
H11 |
110.163 |
| C2 |
C4 |
C6 |
114.137 |
|
C2 |
C4 |
H12 |
114.862 |
| C3 |
C1 |
C4 |
89.060 |
|
C3 |
C1 |
H7 |
111.670 |
| C3 |
C1 |
H8 |
117.431 |
|
C3 |
C2 |
C4 |
89.060 |
| C3 |
C2 |
H9 |
111.670 |
|
C3 |
C2 |
H10 |
117.431 |
| C3 |
C5 |
H13 |
110.539 |
|
C3 |
C5 |
H15 |
111.150 |
| C3 |
C5 |
H16 |
111.150 |
|
C4 |
C1 |
H7 |
112.168 |
| C4 |
C1 |
H8 |
117.064 |
|
C4 |
C2 |
H9 |
112.168 |
| C4 |
C2 |
H10 |
117.064 |
|
C4 |
C6 |
H14 |
111.060 |
| C4 |
C6 |
H17 |
110.915 |
|
C4 |
C6 |
H18 |
110.915 |
| C5 |
C3 |
H11 |
109.542 |
|
C6 |
C4 |
H12 |
109.478 |
| H7 |
C1 |
H8 |
108.466 |
|
H9 |
C2 |
H10 |
108.466 |
| H13 |
C5 |
H15 |
107.940 |
|
H13 |
C5 |
H16 |
107.940 |
| H14 |
C6 |
H17 |
107.837 |
|
H14 |
C6 |
H18 |
107.837 |
| H15 |
C5 |
H16 |
107.988 |
|
H17 |
C6 |
H18 |
108.142 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
