return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H12 ((1r,3r)-1,3-dimethylcyclobutane)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2H 1Ag
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-235.526680
Energy at 298.15K 
HF Energy-235.268173
Nuclear repulsion energy248.316198
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 Ag 3137 2977 0.00      
2 Ag 3095 2937 0.00      
3 Ag 3083 2926 0.00      
4 Ag 3064 2908 0.00      
5 Ag 1546 1468 0.00      
6 Ag 1542 1464 0.00      
7 Ag 1455 1381 0.00      
8 Ag 1405 1334 0.00      
9 Ag 1280 1215 0.00      
10 Ag 1138 1080 0.00      
11 Ag 978 928 0.00      
12 Ag 898 852 0.00      
13 Ag 669 635 0.00      
14 Ag 313 297 0.00      
15 Au 3139 2979 65.20      
16 Au 3083 2926 69.62      
17 Au 1548 1470 2.33      
18 Au 1522 1445 3.50      
19 Au 1324 1257 1.68      
20 Au 1186 1126 0.48      
21 Au 1029 977 2.77      
22 Au 915 869 0.01      
23 Au 265 252 0.03      
24 Au 239 227 0.10      
25 Bg 3140 2980 0.00      
26 Bg 3133 2974 0.00      
27 Bg 1546 1467 0.00      
28 Bg 1306 1240 0.00      
29 Bg 1270 1205 0.00      
30 Bg 1158 1100 0.00      
31 Bg 942 894 0.00      
32 Bg 822 781 0.00      
33 Bg 385 365 0.00      
34 Bg 265 251 0.00      
35 Bu 3144 2984 128.18      
36 Bu 3132 2973 38.14      
37 Bu 3084 2928 27.73      
38 Bu 3064 2908 57.04      
39 Bu 1546 1467 10.64      
40 Bu 1457 1383 4.49      
41 Bu 1390 1319 10.96      
42 Bu 1312 1245 0.78      
43 Bu 1224 1162 0.20      
44 Bu 1068 1014 0.83      
45 Bu 922 875 2.08      
46 Bu 802 761 1.41      
47 Bu 432 410 0.16      
48 Bu 81i 76i 0.00      

Unscaled Zero Point Vibrational Energy (zpe) 37156.9 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 35269.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.27450 0.07553 0.06737

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is C2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 1.096
C2 0.000 0.000 -1.096
C3 0.000 1.104 0.000
C4 0.000 -1.104 0.000
C5 1.195 2.046 0.000
C6 -1.195 -2.046 0.000
H7 0.885 0.000 1.740
H8 -0.885 0.000 1.740
H9 0.885 0.000 -1.740
H10 -0.885 0.000 -1.740
H11 -0.924 1.692 0.000
H12 0.924 -1.692 0.000
H13 2.135 1.482 0.000
H14 -2.135 -1.482 0.000
H15 1.193 2.690 -0.886
H16 1.193 2.690 0.886
H17 -1.193 -2.690 -0.886
H18 -1.193 -2.690 0.886

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18
C12.19271.55581.55582.61092.61091.09451.09452.97132.97132.21732.21732.82072.82073.54832.95073.54832.9507
C22.19271.55581.55582.61092.61092.97132.97131.09451.09452.21732.21732.82072.82072.95073.54832.95073.5483
C31.55581.55582.20791.52203.36912.24282.24282.24282.24281.09482.94422.16813.35342.17382.17384.07494.0749
C41.55581.55582.20793.36911.52202.24282.24282.24282.24282.94421.09483.35342.16814.07494.07492.17382.1738
C52.61092.61091.52203.36914.73922.70373.39752.70373.39752.14853.74741.09564.85171.09521.09525.37815.3781
C62.61092.61093.36911.52204.73923.39752.70373.39752.70373.74742.14854.85171.09565.37815.37811.09521.0952
H71.09452.97132.24282.24282.70373.39751.77043.48013.90463.02682.42712.60503.78753.77182.83964.29573.5056
H81.09452.97132.24282.24283.39752.70371.77043.90463.48012.42713.02683.78752.60504.29573.50563.77182.8396
H92.97131.09452.24282.24282.70373.39753.48013.90461.77043.02682.42712.60503.78752.83963.77183.50564.2957
H102.97131.09452.24282.24283.39752.70373.90463.48011.77042.42713.02683.78752.60503.50564.29572.83963.7718
H112.21732.21731.09482.94422.14853.74743.02682.42713.02682.42713.85473.06573.39702.50282.50284.47874.4787
H122.21732.21732.94421.09483.74742.14852.42713.02682.42713.02683.85473.39703.06574.47874.47872.50282.5028
H132.82072.82072.16813.35341.09564.85172.60503.78752.60503.78753.06573.39705.19791.76921.76925.41045.4104
H142.82072.82073.35342.16814.85171.09563.78752.60503.78752.60503.39703.06575.19795.41045.41041.76921.7692
H153.54832.95072.17384.07491.09525.37813.77184.29572.83963.50562.50284.47871.76925.41041.77115.88646.1470
H162.95073.54832.17384.07491.09525.37812.83963.50563.77184.29572.50284.47871.76925.41041.77116.14705.8864
H173.54832.95074.07492.17385.37811.09524.29573.77183.50562.83964.47872.50285.41041.76925.88646.14701.7711
H182.95073.54834.07492.17385.37811.09523.50562.83964.29573.77184.47872.50285.41041.76926.14705.88641.7711

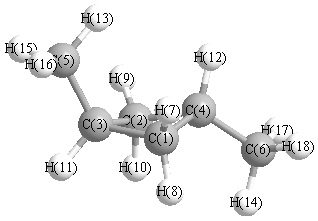
picture of (1r,3r)-1,3-dimethylcyclobutane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C3 C2 89.605 C1 C3 C5 116.051
C1 C3 H11 112.388 C1 C4 C2 89.605
C1 C4 C6 116.051 C1 C4 H12 112.388
C2 C3 C5 116.051 C2 C3 H11 112.388
C2 C4 C6 116.051 C2 C4 H12 112.388
C3 C1 C4 90.395 C3 C1 H7 114.483
C3 C1 H8 114.483 C3 C2 C4 90.395
C3 C2 H9 114.483 C3 C2 H10 114.483
C3 C5 H13 110.791 C3 C5 H15 111.271
C3 C5 H16 111.271 C4 C1 H7 114.483
C4 C1 H8 114.483 C4 C2 H9 114.483
C4 C2 H10 114.483 C4 C6 H14 110.791
C4 C6 H17 111.271 C4 C6 H18 111.271
C5 C3 H11 109.295 C6 C4 H12 109.295
H7 C1 H8 107.952 H9 C2 H10 107.952
H13 C5 H15 107.715 H13 C5 H16 107.715
H14 C6 H17 107.715 H14 C6 H18 107.715
H15 C5 H16 107.918 H17 C6 H18 107.918
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.308      
2 C -0.308      
3 C -0.104      
4 C -0.104      
5 C -0.461      
6 C -0.461      
7 H 0.145      
8 H 0.145      
9 H 0.145      
10 H 0.145      
11 H 0.141      
12 H 0.141      
13 H 0.144      
14 H 0.144      
15 H 0.149      
16 H 0.149      
17 H 0.149      
18 H 0.149      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 8.514 0.757 0.000
y 0.757 9.680 0.000
z 0.000 0.000 8.689


<r2> (average value of r2) Å2
<r2> 195.439
(<r2>)1/2 13.980