Jump to
S1C2
S1C3
Energy calculated at MP3/6-31G*
| | hartrees |
|---|
| Energy at 0K | -100.057158 |
| Energy at 298.15K | -100.056209 |
| HF Energy | -99.771176 |
| Nuclear repulsion energy | 27.472065 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at MP3/6-31G*
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| Li1 |
0.000 |
0.000 |
-2.089 |
| C2 |
0.000 |
0.000 |
-0.148 |
| N3 |
0.000 |
0.000 |
1.022 |
Atom - Atom Distances (Å)
| |
Li1 |
C2 |
N3 |
| Li1 | | 1.9411 | 3.1113 |
C2 | 1.9411 | | 1.1702 | N3 | 3.1113 | 1.1702 | |
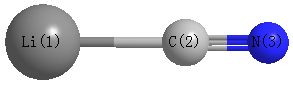 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Li1 |
C2 |
N3 |
180.000 |
|
Li1 |
N3 |
C2 |
0.000 |
| C2 |
Li1 |
N3 |
0.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Energy calculated at MP3/6-31G*
| | hartrees |
|---|
| Energy at 0K | -100.063513 |
| Energy at 298.15K | -100.062792 |
| HF Energy | -99.778901 |
| Nuclear repulsion energy | 29.066518 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at MP3/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| Li1 |
1.442 |
-0.600 |
0.000 |
| C2 |
-0.721 |
-0.367 |
0.000 |
| N3 |
0.000 |
0.572 |
0.000 |
Atom - Atom Distances (Å)
| |
Li1 |
C2 |
N3 |
| Li1 | | 2.1754 | 1.8579 |
C2 | 2.1754 | | 1.1835 | N3 | 1.8579 | 1.1835 | |
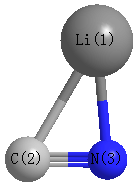 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Li1 |
C2 |
N3 |
58.622 |
|
Li1 |
N3 |
C2 |
88.433 |
| C2 |
Li1 |
N3 |
32.946 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at MP3/6-31G*
| | hartrees |
|---|
| Energy at 0K | -100.059101 |
| Energy at 298.15K | -100.057675 |
| HF Energy | -99.779493 |
| Nuclear repulsion energy | 28.174616 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at MP3/6-31G*
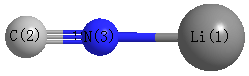
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| Li1 |
0.000 |
0.000 |
1.905 |
| C2 |
0.000 |
0.000 |
-1.076 |
| N3 |
0.000 |
0.000 |
0.106 |
Atom - Atom Distances (Å)
| |
Li1 |
C2 |
N3 |
| Li1 | | 2.9813 | 1.7993 |
C2 | 2.9813 | | 1.1820 | N3 | 1.7993 | 1.1820 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Li1 |
C2 |
N3 |
0.000 |
|
Li1 |
N3 |
C2 |
180.000 |
| C2 |
Li1 |
N3 |
0.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
