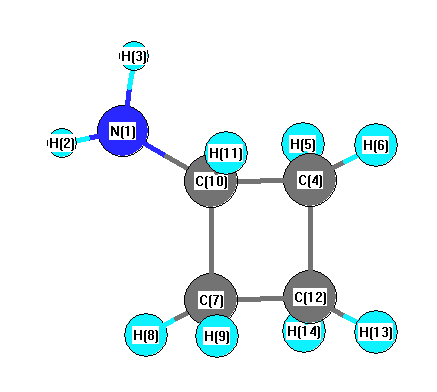
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3479 |
3328 |
2.68 |
|
|
|
| 2 |
A' |
3158 |
3022 |
50.41 |
|
|
|
| 3 |
A' |
3127 |
2991 |
23.09 |
|
|
|
| 4 |
A' |
3095 |
2961 |
36.91 |
|
|
|
| 5 |
A' |
3078 |
2945 |
19.19 |
|
|
|
| 6 |
A' |
2977 |
2848 |
104.38 |
|
|
|
| 7 |
A' |
1692 |
1619 |
22.78 |
|
|
|
| 8 |
A' |
1526 |
1460 |
4.14 |
|
|
|
| 9 |
A' |
1498 |
1433 |
2.26 |
|
|
|
| 10 |
A' |
1403 |
1342 |
25.60 |
|
|
|
| 11 |
A' |
1308 |
1251 |
4.82 |
|
|
|
| 12 |
A' |
1256 |
1202 |
1.47 |
|
|
|
| 13 |
A' |
1175 |
1124 |
8.62 |
|
|
|
| 14 |
A' |
1101 |
1053 |
8.25 |
|
|
|
| 15 |
A' |
984 |
942 |
8.28 |
|
|
|
| 16 |
A' |
901 |
862 |
5.12 |
|
|
|
| 17 |
A' |
885 |
846 |
101.51 |
|
|
|
| 18 |
A' |
831 |
795 |
35.55 |
|
|
|
| 19 |
A' |
671 |
642 |
1.04 |
|
|
|
| 20 |
A' |
404 |
387 |
5.39 |
|
|
|
| 21 |
A' |
162 |
155 |
1.63 |
|
|
|
| 22 |
A" |
3569 |
3414 |
0.11 |
|
|
|
| 23 |
A" |
3131 |
2996 |
9.63 |
|
|
|
| 24 |
A" |
3074 |
2941 |
72.76 |
|
|
|
| 25 |
A" |
1491 |
1426 |
1.57 |
|
|
|
| 26 |
A" |
1364 |
1305 |
0.23 |
|
|
|
| 27 |
A" |
1289 |
1233 |
0.74 |
|
|
|
| 28 |
A" |
1273 |
1217 |
0.13 |
|
|
|
| 29 |
A" |
1239 |
1185 |
0.52 |
|
|
|
| 30 |
A" |
1180 |
1129 |
0.82 |
|
|
|
| 31 |
A" |
1045 |
1000 |
0.25 |
|
|
|
| 32 |
A" |
964 |
922 |
0.69 |
|
|
|
| 33 |
A" |
936 |
896 |
2.05 |
|
|
|
| 34 |
A" |
776 |
743 |
1.05 |
|
|
|
| 35 |
A" |
392 |
375 |
12.09 |
|
|
|
| 36 |
A" |
288 |
275 |
34.76 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28360.5 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 27132.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.747 |
|
|
-0.887 |
| 2 |
H |
0.311 |
|
|
0.326 |
| 3 |
H |
0.311 |
|
|
0.326 |
| 4 |
C |
-0.313 |
|
|
-0.312 |
| 5 |
H |
0.161 |
|
|
0.093 |
| 6 |
H |
0.157 |
|
|
0.081 |
| 7 |
C |
-0.313 |
|
|
-0.312 |
| 8 |
H |
0.161 |
|
|
0.093 |
| 9 |
H |
0.157 |
|
|
0.081 |
| 10 |
C |
-0.040 |
|
|
0.482 |
| 11 |
H |
0.133 |
|
|
-0.084 |
| 12 |
C |
-0.316 |
|
|
0.080 |
| 13 |
H |
0.157 |
|
|
0.010 |
| 14 |
H |
0.181 |
|
|
0.023 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.238 |
1.310 |
0.000 |
1.331 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
0.237 |
1.305 |
0.000 |
1.326 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.088 |
-2.510 |
0.000 |
| y |
-2.510 |
-30.124 |
0.000 |
| z |
0.000 |
0.000 |
-30.937 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.557 |
-2.510 |
0.000 |
| y |
-2.510 |
1.389 |
0.000 |
| z |
0.000 |
0.000 |
0.169 |
|
| Polar |
|---|
| 3z2-r2 | 0.337 |
|---|
| x2-y2 | -1.964 |
|---|
| xy | -2.510 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
113.304 |
| (<r2>)1/2 |
10.644 |