return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H9N (Cyclobutylamine)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-212.290392
Energy at 298.15K-212.301597
HF Energy-212.065079
Nuclear repulsion energy187.301528
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3480 3303 2.31      
2 A' 3181 3019 47.34      
3 A' 3147 2988 26.51      
4 A' 3117 2959 37.14      
5 A' 3099 2942 17.89      
6 A' 3012 2859 96.15      
7 A' 1709 1622 22.18      
8 A' 1552 1473 3.30      
9 A' 1525 1448 1.91      
10 A' 1419 1347 25.36      
11 A' 1325 1258 3.61      
12 A' 1268 1204 1.08      
13 A' 1186 1126 8.83      
14 A' 1109 1053 8.74      
15 A' 987 937 13.26      
16 A' 912 866 28.87      
17 A' 897 851 73.97      
18 A' 846 803 28.51      
19 A' 682 648 0.99      
20 A' 410 389 4.92      
21 A' 175 166 1.72      
22 A" 3570 3388 0.20      
23 A" 3151 2991 10.49      
24 A" 3095 2937 67.75      
25 A" 1517 1440 0.92      
26 A" 1379 1309 0.34      
27 A" 1306 1240 0.42      
28 A" 1289 1224 0.20      
29 A" 1251 1187 0.33      
30 A" 1192 1131 0.90      
31 A" 1055 1001 0.29      
32 A" 961 912 0.49      
33 A" 939 891 2.01      
34 A" 787 747 0.85      
35 A" 395 375 12.53      
36 A" 291 277 33.68      

Unscaled Zero Point Vibrational Energy (zpe) 28607.4 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 27154.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.27749 0.15882 0.13251

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 -1.356 0.984 0.000
H2 -1.591 1.586 0.787
H3 -1.591 1.586 -0.787
C4 0.459 -0.267 -1.091
H5 -0.304 -0.361 -1.865
H6 1.455 -0.098 -1.499
C7 0.459 -0.267 1.091
H8 -0.304 -0.361 1.865
H9 1.455 -0.098 1.499
C10 0.093 0.761 0.000
H11 0.685 1.689 0.000
C12 0.459 -1.372 0.000
H13 1.336 -2.021 0.000
H14 -0.471 -1.937 0.000

Atom - Atom Distances (Å)
  N1 H2 H3 C4 H5 H6 C7 H8 H9 C10 H11 C12 H13 H14
N11.01821.01822.45982.52843.36432.45982.52843.36431.46612.15922.97464.03513.0521
H21.01821.57383.34143.53244.16402.78042.57083.55262.03392.41063.68454.71203.7795
H31.01821.57382.78042.57083.55263.34143.53244.16402.03392.41063.68454.71203.7795
C42.45983.34142.78041.09061.08972.18193.05392.77991.54322.25121.55282.24422.2002
H52.52843.53242.57081.09061.81603.05393.72913.80482.21202.94222.25432.98742.4468
H63.36434.16403.55261.08971.81602.77993.80482.99762.19972.45572.20532.44163.0557
C72.45982.78043.34142.18193.05392.77991.09061.08971.54322.25121.55282.24422.2002
H82.52842.57083.53243.05393.72913.80481.09061.81602.21202.94222.25432.98742.4468
H93.36433.55264.16402.77993.80482.99761.08971.81602.19972.45572.20532.44163.0557
C101.46612.03392.03391.54322.21202.19971.54322.21202.19971.10072.16443.04742.7558
H112.15922.41062.41062.25122.94222.45572.25122.94222.45571.10073.06943.76683.8052
C122.97463.68453.68451.55282.25432.20531.55282.25432.20532.16443.06941.09131.0876
H134.03514.71204.71202.24422.98742.44162.24422.98742.44163.04743.76681.09131.8091
H143.05213.77953.77952.20022.44683.05572.20022.44683.05572.75583.80521.08761.8091

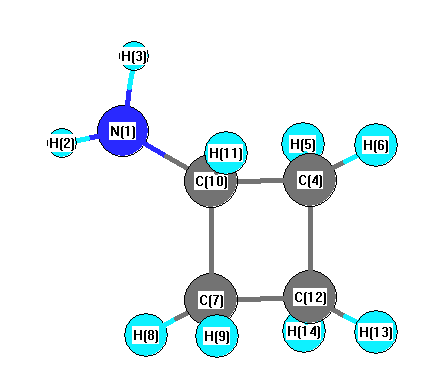
picture of Cyclobutylamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C10 C4 109.625 N1 C10 C7 109.625
N1 C10 H11 113.779 H2 N1 H3 101.215
H2 N1 C10 108.561 H3 N1 C10 108.561
C4 C10 C7 89.974 C4 C10 H11 115.733
C4 C12 C7 89.271 C4 C12 H13 115.040
C4 C12 H14 111.661 H5 C4 H6 112.798
H5 C4 C10 113.124 H5 C4 C12 115.936
H6 C4 C10 112.178 H6 C4 C12 111.948
C7 C10 H11 115.733 C7 C12 H13 115.040
C7 C12 H14 111.661 H8 C7 H9 112.798
H8 C7 C10 113.124 H8 C7 C12 115.936
H9 C7 C10 112.178 H9 C7 C12 111.948
C10 C4 C12 88.710 C10 C7 C12 88.710
H13 C12 H14 112.256
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 N -0.768      
2 H 0.309      
3 H 0.309      
4 C -0.287      
5 H 0.148      
6 H 0.143      
7 C -0.287      
8 H 0.148      
9 H 0.143      
10 C -0.005      
11 H 0.124      
12 C -0.288      
13 H 0.143      
14 H 0.168      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 7.071 -0.517 0.000
y -0.517 6.719 0.000
z 0.000 0.000 6.966


<r2> (average value of r2) Å2
<r2> 113.432
(<r2>)1/2 10.650