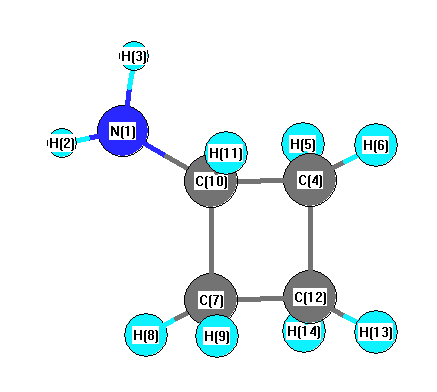
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3358 |
3303 |
6.41 |
|
|
|
| 2 |
A' |
3082 |
3031 |
47.66 |
|
|
|
| 3 |
A' |
3050 |
2999 |
23.41 |
|
|
|
| 4 |
A' |
3019 |
2969 |
37.26 |
|
|
|
| 5 |
A' |
3002 |
2952 |
15.14 |
|
|
|
| 6 |
A' |
2887 |
2839 |
108.24 |
|
|
|
| 7 |
A' |
1642 |
1615 |
17.65 |
|
|
|
| 8 |
A' |
1478 |
1453 |
4.28 |
|
|
|
| 9 |
A' |
1452 |
1428 |
2.13 |
|
|
|
| 10 |
A' |
1352 |
1330 |
24.54 |
|
|
|
| 11 |
A' |
1259 |
1238 |
4.63 |
|
|
|
| 12 |
A' |
1215 |
1195 |
0.74 |
|
|
|
| 13 |
A' |
1135 |
1116 |
6.47 |
|
|
|
| 14 |
A' |
1072 |
1054 |
8.66 |
|
|
|
| 15 |
A' |
961 |
945 |
10.60 |
|
|
|
| 16 |
A' |
877 |
862 |
8.74 |
|
|
|
| 17 |
A' |
853 |
839 |
92.33 |
|
|
|
| 18 |
A' |
811 |
797 |
28.84 |
|
|
|
| 19 |
A' |
646 |
636 |
0.99 |
|
|
|
| 20 |
A' |
393 |
386 |
5.20 |
|
|
|
| 21 |
A' |
153 |
151 |
1.61 |
|
|
|
| 22 |
A" |
3448 |
3392 |
1.20 |
|
|
|
| 23 |
A" |
3055 |
3004 |
8.33 |
|
|
|
| 24 |
A" |
2997 |
2948 |
72.68 |
|
|
|
| 25 |
A" |
1444 |
1420 |
1.41 |
|
|
|
| 26 |
A" |
1320 |
1299 |
0.17 |
|
|
|
| 27 |
A" |
1247 |
1226 |
0.54 |
|
|
|
| 28 |
A" |
1226 |
1206 |
0.07 |
|
|
|
| 29 |
A" |
1200 |
1180 |
0.37 |
|
|
|
| 30 |
A" |
1137 |
1119 |
1.15 |
|
|
|
| 31 |
A" |
1012 |
995 |
0.31 |
|
|
|
| 32 |
A" |
937 |
922 |
0.60 |
|
|
|
| 33 |
A" |
908 |
893 |
1.71 |
|
|
|
| 34 |
A" |
753 |
741 |
1.11 |
|
|
|
| 35 |
A" |
380 |
374 |
11.93 |
|
|
|
| 36 |
A" |
276 |
271 |
32.67 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 27517.9 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 27063.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.698 |
|
|
|
| 2 |
H |
0.295 |
|
|
|
| 3 |
H |
0.295 |
|
|
|
| 4 |
C |
-0.286 |
|
|
|
| 5 |
H |
0.147 |
|
|
|
| 6 |
H |
0.144 |
|
|
|
| 7 |
C |
-0.286 |
|
|
|
| 8 |
H |
0.147 |
|
|
|
| 9 |
H |
0.144 |
|
|
|
| 10 |
C |
-0.039 |
|
|
|
| 11 |
H |
0.117 |
|
|
|
| 12 |
C |
-0.291 |
|
|
|
| 13 |
H |
0.144 |
|
|
|
| 14 |
H |
0.168 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.202 |
1.293 |
0.000 |
1.309 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.250 |
-2.533 |
-0.016 |
| y |
-2.533 |
-30.368 |
-0.002 |
| z |
-0.016 |
-0.002 |
-31.201 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.466 |
-2.533 |
-0.016 |
| y |
-2.533 |
1.358 |
-0.002 |
| z |
-0.016 |
-0.002 |
0.108 |
|
| Polar |
|---|
| 3z2-r2 | 0.216 |
|---|
| x2-y2 | -1.883 |
|---|
| xy | -2.533 |
|---|
| xz | -0.016 |
|---|
| yz | -0.002 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.386 |
-0.571 |
-0.003 |
| y |
-0.571 |
7.034 |
-0.001 |
| z |
-0.003 |
-0.001 |
7.241 |
<r2> (average value of r
2) Å
2
| <r2> |
114.342 |
| (<r2>)1/2 |
10.693 |