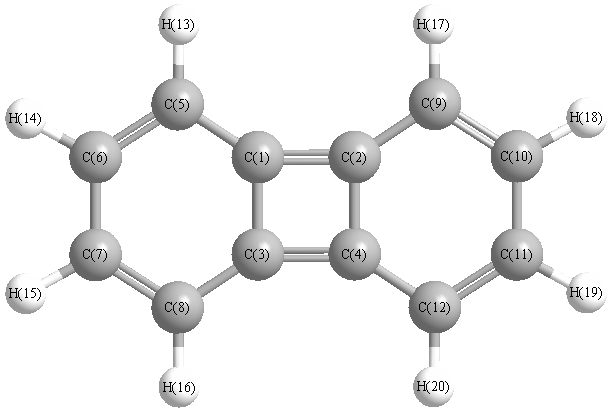
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3224 |
3084 |
0.00 |
|
|
|
| 2 |
Ag |
3208 |
3069 |
0.00 |
|
|
|
| 3 |
Ag |
1737 |
1662 |
0.00 |
|
|
|
| 4 |
Ag |
1538 |
1472 |
0.00 |
|
|
|
| 5 |
Ag |
1452 |
1389 |
0.00 |
|
|
|
| 6 |
Ag |
1203 |
1150 |
0.00 |
|
|
|
| 7 |
Ag |
1158 |
1107 |
0.00 |
|
|
|
| 8 |
Ag |
1029 |
985 |
0.00 |
|
|
|
| 9 |
Ag |
785 |
751 |
0.00 |
|
|
|
| 10 |
Ag |
403 |
386 |
0.00 |
|
|
|
| 11 |
Au |
974 |
932 |
0.00 |
|
|
|
| 12 |
Au |
900 |
861 |
0.00 |
|
|
|
| 13 |
Au |
815 |
780 |
0.00 |
|
|
|
| 14 |
Au |
585 |
560 |
0.00 |
|
|
|
| 15 |
Au |
148 |
142 |
0.00 |
|
|
|
| 16 |
B1g |
974 |
932 |
0.00 |
|
|
|
| 17 |
B1g |
885 |
846 |
0.00 |
|
|
|
| 18 |
B1g |
719 |
687 |
0.00 |
|
|
|
| 19 |
B1g |
463 |
443 |
0.00 |
|
|
|
| 20 |
B1u |
3223 |
3083 |
44.72 |
|
|
|
| 21 |
B1u |
3207 |
3068 |
19.82 |
|
|
|
| 22 |
B1u |
1661 |
1589 |
0.47 |
|
|
|
| 23 |
B1u |
1481 |
1416 |
34.58 |
|
|
|
| 24 |
B1u |
1337 |
1279 |
32.52 |
|
|
|
| 25 |
B1u |
1193 |
1142 |
8.19 |
|
|
|
| 26 |
B1u |
1061 |
1015 |
0.09 |
|
|
|
| 27 |
B1u |
1009 |
965 |
17.74 |
|
|
|
| 28 |
B1u |
625 |
598 |
4.50 |
|
|
|
| 29 |
B2g |
928 |
888 |
0.00 |
|
|
|
| 30 |
B2g |
761 |
728 |
0.00 |
|
|
|
| 31 |
B2g |
446 |
427 |
0.00 |
|
|
|
| 32 |
B2g |
324 |
310 |
0.00 |
|
|
|
| 33 |
B2u |
3218 |
3079 |
52.16 |
|
|
|
| 34 |
B2u |
3195 |
3056 |
0.62 |
|
|
|
| 35 |
B2u |
1658 |
1586 |
1.94 |
|
|
|
| 36 |
B2u |
1496 |
1431 |
5.77 |
|
|
|
| 37 |
B2u |
1306 |
1249 |
2.98 |
|
|
|
| 38 |
B2u |
1160 |
1109 |
11.33 |
|
|
|
| 39 |
B2u |
1077 |
1031 |
1.18 |
|
|
|
| 40 |
B2u |
737 |
705 |
0.43 |
|
|
|
| 41 |
B2u |
216 |
207 |
0.80 |
|
|
|
| 42 |
B3g |
3217 |
3078 |
0.00 |
|
|
|
| 43 |
B3g |
3195 |
3056 |
0.00 |
|
|
|
| 44 |
B3g |
1666 |
1594 |
0.00 |
|
|
|
| 45 |
B3g |
1497 |
1432 |
0.00 |
|
|
|
| 46 |
B3g |
1317 |
1260 |
0.00 |
|
|
|
| 47 |
B3g |
1124 |
1076 |
0.00 |
|
|
|
| 48 |
B3g |
997 |
953 |
0.00 |
|
|
|
| 49 |
B3g |
612 |
585 |
0.00 |
|
|
|
| 50 |
B3g |
566 |
542 |
0.00 |
|
|
|
| 51 |
B3u |
923 |
883 |
6.99 |
|
|
|
| 52 |
B3u |
749 |
717 |
121.41 |
|
|
|
| 53 |
B3u |
378 |
362 |
1.76 |
|
|
|
| 54 |
B3u |
108 |
103 |
2.32 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 34931.5 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 33419.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.093 |
|
|
|
| 2 |
C |
0.093 |
|
|
|
| 3 |
C |
0.093 |
|
|
|
| 4 |
C |
0.093 |
|
|
|
| 5 |
C |
-0.258 |
|
|
|
| 6 |
C |
-0.171 |
|
|
|
| 7 |
C |
-0.171 |
|
|
|
| 8 |
C |
-0.258 |
|
|
|
| 9 |
C |
-0.258 |
|
|
|
| 10 |
C |
-0.171 |
|
|
|
| 11 |
C |
-0.171 |
|
|
|
| 12 |
C |
-0.258 |
|
|
|
| 13 |
H |
0.172 |
|
|
|
| 14 |
H |
0.164 |
|
|
|
| 15 |
H |
0.164 |
|
|
|
| 16 |
H |
0.172 |
|
|
|
| 17 |
H |
0.172 |
|
|
|
| 18 |
H |
0.164 |
|
|
|
| 19 |
H |
0.164 |
|
|
|
| 20 |
H |
0.172 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-73.226 |
0.000 |
0.000 |
| y |
0.000 |
-59.020 |
0.000 |
| z |
0.000 |
0.000 |
-58.217 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-14.607 |
0.000 |
0.000 |
| y |
0.000 |
6.702 |
0.000 |
| z |
0.000 |
0.000 |
7.905 |
|
| Polar |
|---|
| 3z2-r2 | 15.810 |
|---|
| x2-y2 | -14.206 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.767 |
0.000 |
0.000 |
| y |
0.000 |
18.571 |
0.000 |
| z |
0.000 |
0.000 |
30.689 |
<r2> (average value of r
2) Å
2
| <r2> |
560.101 |
| (<r2>)1/2 |
23.666 |