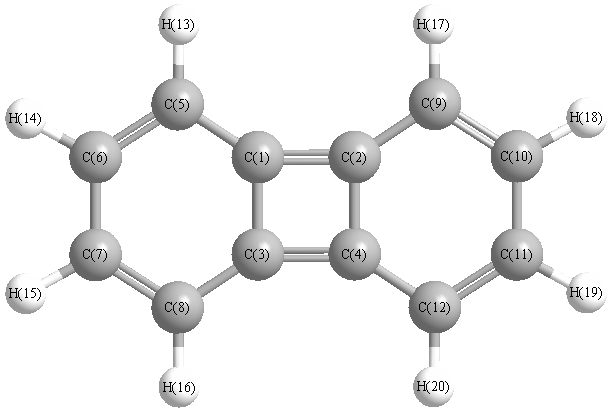
Vibrational Frequencies calculated at B2PLYP/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3237 |
3073 |
0.00 |
|
|
|
| 2 |
Ag |
3222 |
3058 |
0.00 |
|
|
|
| 3 |
Ag |
1729 |
1641 |
0.00 |
|
|
|
| 4 |
Ag |
1534 |
1456 |
0.00 |
|
|
|
| 5 |
Ag |
1458 |
1384 |
0.00 |
|
|
|
| 6 |
Ag |
1213 |
1151 |
0.00 |
|
|
|
| 7 |
Ag |
1157 |
1098 |
0.00 |
|
|
|
| 8 |
Ag |
1028 |
976 |
0.00 |
|
|
|
| 9 |
Ag |
785 |
745 |
0.00 |
|
|
|
| 10 |
Ag |
402 |
382 |
0.00 |
|
|
|
| 11 |
Au |
937 |
889 |
0.00 |
|
|
|
| 12 |
Au |
876 |
831 |
0.00 |
|
|
|
| 13 |
Au |
618 |
586 |
0.00 |
|
|
|
| 14 |
Au |
566 |
537 |
0.00 |
|
|
|
| 15 |
Au |
148 |
140 |
0.00 |
|
|
|
| 16 |
B1g |
938 |
890 |
0.00 |
|
|
|
| 17 |
B1g |
874 |
830 |
0.00 |
|
|
|
| 18 |
B1g |
655 |
621 |
0.00 |
|
|
|
| 19 |
B1g |
458 |
435 |
0.00 |
|
|
|
| 20 |
B1u |
3236 |
3072 |
50.24 |
|
|
|
| 21 |
B1u |
3221 |
3058 |
15.25 |
|
|
|
| 22 |
B1u |
1655 |
1571 |
0.09 |
|
|
|
| 23 |
B1u |
1485 |
1410 |
35.90 |
|
|
|
| 24 |
B1u |
1339 |
1271 |
32.31 |
|
|
|
| 25 |
B1u |
1203 |
1142 |
9.59 |
|
|
|
| 26 |
B1u |
1060 |
1006 |
0.02 |
|
|
|
| 27 |
B1u |
1005 |
954 |
14.95 |
|
|
|
| 28 |
B1u |
629 |
597 |
4.31 |
|
|
|
| 29 |
B2g |
898 |
853 |
0.00 |
|
|
|
| 30 |
B2g |
756 |
718 |
0.00 |
|
|
|
| 31 |
B2g |
417 |
396 |
0.00 |
|
|
|
| 32 |
B2g |
321 |
304 |
0.00 |
|
|
|
| 33 |
B2u |
3231 |
3067 |
52.67 |
|
|
|
| 34 |
B2u |
3210 |
3047 |
0.29 |
|
|
|
| 35 |
B2u |
1653 |
1569 |
1.50 |
|
|
|
| 36 |
B2u |
1502 |
1426 |
6.39 |
|
|
|
| 37 |
B2u |
1314 |
1247 |
2.05 |
|
|
|
| 38 |
B2u |
1165 |
1106 |
10.15 |
|
|
|
| 39 |
B2u |
1082 |
1027 |
1.53 |
|
|
|
| 40 |
B2u |
739 |
701 |
0.31 |
|
|
|
| 41 |
B2u |
215 |
204 |
0.72 |
|
|
|
| 42 |
B3g |
3230 |
3066 |
0.00 |
|
|
|
| 43 |
B3g |
3210 |
3046 |
0.00 |
|
|
|
| 44 |
B3g |
1660 |
1576 |
0.00 |
|
|
|
| 45 |
B3g |
1504 |
1428 |
0.00 |
|
|
|
| 46 |
B3g |
1328 |
1260 |
0.00 |
|
|
|
| 47 |
B3g |
1130 |
1073 |
0.00 |
|
|
|
| 48 |
B3g |
1003 |
952 |
0.00 |
|
|
|
| 49 |
B3g |
615 |
584 |
0.00 |
|
|
|
| 50 |
B3g |
567 |
539 |
0.00 |
|
|
|
| 51 |
B3u |
898 |
853 |
5.56 |
|
|
|
| 52 |
B3u |
749 |
711 |
114.11 |
|
|
|
| 53 |
B3u |
371 |
352 |
1.42 |
|
|
|
| 54 |
B3u |
104 |
99 |
2.15 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 34768.7 cm
-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 33002.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.078 |
|
|
|
| 2 |
C |
0.078 |
|
|
|
| 3 |
C |
0.078 |
|
|
|
| 4 |
C |
0.078 |
|
|
|
| 5 |
C |
-0.231 |
|
|
|
| 6 |
C |
-0.155 |
|
|
|
| 7 |
C |
-0.155 |
|
|
|
| 8 |
C |
-0.231 |
|
|
|
| 9 |
C |
-0.231 |
|
|
|
| 10 |
C |
-0.155 |
|
|
|
| 11 |
C |
-0.155 |
|
|
|
| 12 |
C |
-0.231 |
|
|
|
| 13 |
H |
0.159 |
|
|
|
| 14 |
H |
0.149 |
|
|
|
| 15 |
H |
0.149 |
|
|
|
| 16 |
H |
0.159 |
|
|
|
| 17 |
H |
0.159 |
|
|
|
| 18 |
H |
0.149 |
|
|
|
| 19 |
H |
0.149 |
|
|
|
| 20 |
H |
0.159 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-73.511 |
0.000 |
0.000 |
| y |
0.000 |
-60.146 |
0.000 |
| z |
0.000 |
0.000 |
-59.486 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-13.695 |
0.000 |
0.000 |
| y |
0.000 |
6.353 |
0.000 |
| z |
0.000 |
0.000 |
7.342 |
|
| Polar |
|---|
| 3z2-r2 | 14.685 |
|---|
| x2-y2 | -13.365 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.670 |
0.000 |
0.000 |
| y |
0.000 |
18.461 |
0.000 |
| z |
0.000 |
0.000 |
30.056 |
<r2> (average value of r
2) Å
2
| <r2> |
562.147 |
| (<r2>)1/2 |
23.710 |