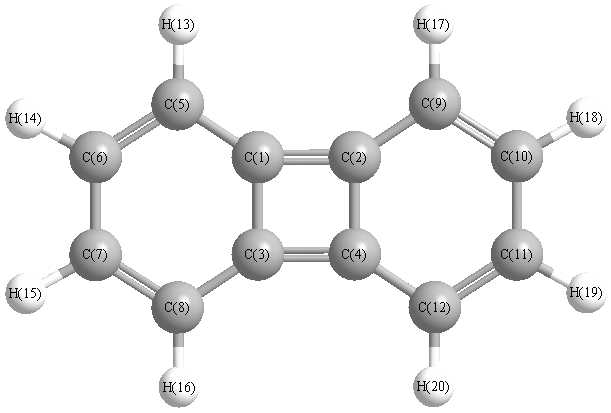
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3143 |
3091 |
0.00 |
|
|
|
| 2 |
Ag |
3127 |
3076 |
0.00 |
|
|
|
| 3 |
Ag |
1687 |
1659 |
0.00 |
|
|
|
| 4 |
Ag |
1499 |
1474 |
0.00 |
|
|
|
| 5 |
Ag |
1407 |
1384 |
0.00 |
|
|
|
| 6 |
Ag |
1172 |
1153 |
0.00 |
|
|
|
| 7 |
Ag |
1123 |
1104 |
0.00 |
|
|
|
| 8 |
Ag |
1003 |
986 |
0.00 |
|
|
|
| 9 |
Ag |
763 |
751 |
0.00 |
|
|
|
| 10 |
Ag |
392 |
386 |
0.00 |
|
|
|
| 11 |
Au |
928 |
913 |
0.00 |
|
|
|
| 12 |
Au |
862 |
848 |
0.00 |
|
|
|
| 13 |
Au |
790 |
777 |
0.00 |
|
|
|
| 14 |
Au |
565 |
556 |
0.00 |
|
|
|
| 15 |
Au |
143 |
141 |
0.00 |
|
|
|
| 16 |
B1g |
929 |
914 |
0.00 |
|
|
|
| 17 |
B1g |
847 |
833 |
0.00 |
|
|
|
| 18 |
B1g |
694 |
683 |
0.00 |
|
|
|
| 19 |
B1g |
448 |
441 |
0.00 |
|
|
|
| 20 |
B1u |
3142 |
3090 |
47.09 |
|
|
|
| 21 |
B1u |
3127 |
3075 |
16.32 |
|
|
|
| 22 |
B1u |
1603 |
1576 |
1.23 |
|
|
|
| 23 |
B1u |
1432 |
1409 |
26.82 |
|
|
|
| 24 |
B1u |
1325 |
1303 |
25.67 |
|
|
|
| 25 |
B1u |
1162 |
1143 |
4.99 |
|
|
|
| 26 |
B1u |
1028 |
1011 |
0.16 |
|
|
|
| 27 |
B1u |
985 |
969 |
13.81 |
|
|
|
| 28 |
B1u |
606 |
596 |
4.08 |
|
|
|
| 29 |
B2g |
883 |
868 |
0.00 |
|
|
|
| 30 |
B2g |
732 |
720 |
0.00 |
|
|
|
| 31 |
B2g |
435 |
428 |
0.00 |
|
|
|
| 32 |
B2g |
312 |
307 |
0.00 |
|
|
|
| 33 |
B2u |
3137 |
3085 |
55.19 |
|
|
|
| 34 |
B2u |
3115 |
3064 |
0.25 |
|
|
|
| 35 |
B2u |
1596 |
1570 |
2.02 |
|
|
|
| 36 |
B2u |
1450 |
1426 |
4.16 |
|
|
|
| 37 |
B2u |
1265 |
1244 |
3.15 |
|
|
|
| 38 |
B2u |
1127 |
1109 |
11.15 |
|
|
|
| 39 |
B2u |
1046 |
1028 |
1.43 |
|
|
|
| 40 |
B2u |
716 |
704 |
0.32 |
|
|
|
| 41 |
B2u |
210 |
206 |
0.76 |
|
|
|
| 42 |
B3g |
3136 |
3084 |
0.00 |
|
|
|
| 43 |
B3g |
3115 |
3063 |
0.00 |
|
|
|
| 44 |
B3g |
1607 |
1581 |
0.00 |
|
|
|
| 45 |
B3g |
1450 |
1426 |
0.00 |
|
|
|
| 46 |
B3g |
1274 |
1253 |
0.00 |
|
|
|
| 47 |
B3g |
1094 |
1076 |
0.00 |
|
|
|
| 48 |
B3g |
965 |
949 |
0.00 |
|
|
|
| 49 |
B3g |
593 |
583 |
0.00 |
|
|
|
| 50 |
B3g |
548 |
539 |
0.00 |
|
|
|
| 51 |
B3u |
879 |
864 |
5.84 |
|
|
|
| 52 |
B3u |
721 |
709 |
108.15 |
|
|
|
| 53 |
B3u |
363 |
357 |
1.44 |
|
|
|
| 54 |
B3u |
102 |
100 |
2.07 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 33899.1 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 33339.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.108 |
|
|
|
| 2 |
C |
0.108 |
|
|
|
| 3 |
C |
0.108 |
|
|
|
| 4 |
C |
0.108 |
|
|
|
| 5 |
C |
-0.246 |
|
|
|
| 6 |
C |
-0.146 |
|
|
|
| 7 |
C |
-0.146 |
|
|
|
| 8 |
C |
-0.246 |
|
|
|
| 9 |
C |
-0.246 |
|
|
|
| 10 |
C |
-0.146 |
|
|
|
| 11 |
C |
-0.146 |
|
|
|
| 12 |
C |
-0.246 |
|
|
|
| 13 |
H |
0.145 |
|
|
|
| 14 |
H |
0.140 |
|
|
|
| 15 |
H |
0.140 |
|
|
|
| 16 |
H |
0.145 |
|
|
|
| 17 |
H |
0.145 |
|
|
|
| 18 |
H |
0.140 |
|
|
|
| 19 |
H |
0.140 |
|
|
|
| 20 |
H |
0.145 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-73.120 |
0.000 |
0.000 |
| y |
0.000 |
-59.657 |
0.000 |
| z |
0.000 |
0.000 |
-58.670 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-13.957 |
0.000 |
0.000 |
| y |
0.000 |
6.238 |
0.000 |
| z |
0.000 |
0.000 |
7.718 |
|
| Polar |
|---|
| 3z2-r2 | 15.437 |
|---|
| x2-y2 | -13.463 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.818 |
0.000 |
0.000 |
| y |
0.000 |
19.040 |
0.000 |
| z |
0.000 |
0.000 |
32.314 |
<r2> (average value of r
2) Å
2
| <r2> |
566.217 |
| (<r2>)1/2 |
23.795 |