Jump to
S1C2
S2C1
S2C2
Energy calculated at MP3/6-31G*
| | hartrees |
|---|
| Energy at 0K | -131.023350 |
| Energy at 298.15K | -131.022869 |
| HF Energy | -130.679204 |
| Nuclear repulsion energy | 47.394129 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP3/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3396 |
3187 |
18.43 |
|
|
|
| 2 |
A' |
1985 |
1863 |
30.79 |
|
|
|
| 3 |
A' |
1145 |
1075 |
31.50 |
|
|
|
| 4 |
A' |
738 |
693 |
16.31 |
|
|
|
| 5 |
A' |
425 |
399 |
3.55 |
|
|
|
| 6 |
A" |
483 |
454 |
4.82 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4086.1 cm
-1
Scaled (by 0.9386) Zero Point Vibrational Energy (zpe) 3835.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP3/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.099 |
-1.241 |
0.000 |
| C2 |
0.000 |
0.107 |
0.000 |
| N3 |
0.009 |
1.274 |
0.000 |
| H4 |
0.535 |
-2.110 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
| C1 | | 1.3519 | 2.5174 | 1.0756 |
C2 | 1.3519 | | 1.1668 | 2.2803 | N3 | 2.5174 | 1.1668 | | 3.4240 | H4 | 1.0756 | 2.2803 | 3.4240 | |
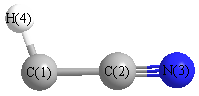 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
176.211 |
|
C2 |
C1 |
H4 |
139.600 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S2C1
S2C2
Energy calculated at MP3/6-31G*
| | hartrees |
|---|
| Energy at 0K | -131.019907 |
| Energy at 298.15K | |
| HF Energy | -130.677611 |
| Nuclear repulsion energy | 47.565924 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP3/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Σ |
3506 |
3290 |
70.27 |
|
|
|
| 2 |
Σ |
1585 |
1487 |
62.85 |
|
|
|
| 3 |
Σ |
1276 |
1198 |
40.57 |
|
|
|
| 4 |
Π |
473 |
444 |
0.26 |
|
|
|
| 4 |
Π |
473 |
444 |
0.26 |
|
|
|
| 5 |
Π |
500i |
470i |
59.02 |
|
|
|
| 5 |
Π |
500i |
470i |
59.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 3155.7 cm
-1
Scaled (by 0.9386) Zero Point Vibrational Energy (zpe) 2962.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP3/6-31G*
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-1.208 |
| C2 |
0.000 |
0.000 |
0.089 |
| N3 |
0.000 |
0.000 |
1.284 |
| H4 |
0.000 |
0.000 |
-2.275 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
| C1 | | 1.2972 | 2.4920 | 1.0669 |
C2 | 1.2972 | | 1.1948 | 2.3641 | N3 | 2.4920 | 1.1948 | | 3.5589 | H4 | 1.0669 | 2.3641 | 3.5589 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
180.000 |
|
C2 |
C1 |
H4 |
180.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at MP3/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.139 |
|
|
|
| 2 |
C |
0.236 |
|
|
|
| 3 |
N |
-0.397 |
|
|
|
| 4 |
H |
0.300 |
|
|
|
Electric dipole moments
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
35.883 |
| (<r2>)1/2 |
5.990 |
Jump to
S1C1
S1C2
S2C2
Energy calculated at MP3/6-31G*
| | hartrees |
|---|
| Energy at 0K | -131.005336 |
| Energy at 298.15K | -131.004738 |
| HF Energy | -130.616321 |
| Nuclear repulsion energy | 46.733829 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP3/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3093 |
2904 |
19.65 |
|
|
|
| 2 |
A' |
2266 |
2127 |
21.48 |
|
|
|
| 3 |
A' |
1077 |
1011 |
81.93 |
|
|
|
| 4 |
A' |
995 |
934 |
1.79 |
|
|
|
| 5 |
A' |
407 |
382 |
18.37 |
|
|
|
| 6 |
A" |
314 |
295 |
9.25 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4076.5 cm
-1
Scaled (by 0.9386) Zero Point Vibrational Energy (zpe) 3826.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP3/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.067 |
-1.302 |
0.000 |
| C2 |
0.000 |
0.103 |
0.000 |
| N3 |
-0.218 |
1.256 |
0.000 |
| H4 |
1.129 |
-1.597 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
| C1 | | 1.4059 | 2.5733 | 1.1028 |
C2 | 1.4059 | | 1.1736 | 2.0404 | N3 | 2.5733 | 1.1736 | | 3.1549 | H4 | 1.1028 | 2.0404 | 3.1549 | |
More geometry information
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
S2C1
Energy calculated at MP3/6-31G*
| | hartrees |
|---|
| Energy at 0K | -131.005336 |
| Energy at 298.15K | -131.004738 |
| HF Energy | -130.616321 |
| Nuclear repulsion energy | 46.733829 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP3/6-31G*
Geometric Data calculated at MP3/6-31G*
Point Group is Cs
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
