Jump to
S1C2
S2C1
S2C2
Energy calculated at PBEPBEultrafine/6-31G*
| | hartrees |
|---|
| Energy at 0K | -131.260309 |
| Energy at 298.15K | -131.259608 |
| HF Energy | -131.260309 |
| Nuclear repulsion energy | 46.896621 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3339 |
3284 |
31.96 |
|
|
|
| 2 |
A' |
1823 |
1793 |
23.57 |
|
|
|
| 3 |
A' |
1223 |
1203 |
3.98 |
|
|
|
| 4 |
A' |
478 |
470 |
34.02 |
|
|
|
| 5 |
A' |
389 |
382 |
20.62 |
|
|
|
| 6 |
A" |
439 |
432 |
2.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 3845.0 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 3781.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBEultrafine/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.060 |
-1.231 |
0.000 |
| C2 |
0.000 |
0.084 |
0.000 |
| N3 |
-0.139 |
1.292 |
0.000 |
| H4 |
0.612 |
-2.158 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
| C1 | | 1.3157 | 2.5303 | 1.0795 |
C2 | 1.3157 | | 1.2161 | 2.3237 | N3 | 2.5303 | 1.2161 | | 3.5305 | H4 | 1.0795 | 2.3237 | 3.5305 | |
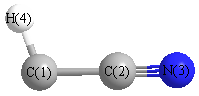 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
176.053 |
|
C2 |
C1 |
H4 |
151.791 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S2C1
S2C2
Energy calculated at PBEPBEultrafine/6-31G*
| | hartrees |
|---|
| Energy at 0K | -131.259611 |
| Energy at 298.15K | |
| HF Energy | -131.259611 |
| Nuclear repulsion energy | 46.945920 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Σ |
3400 |
3344 |
62.31 |
|
|
|
| 2 |
Σ |
1784 |
1755 |
24.05 |
|
|
|
| 3 |
Σ |
1254 |
1234 |
3.08 |
|
|
|
| 4 |
Π |
441 |
434 |
0.87 |
|
|
|
| 4 |
Π |
441 |
434 |
0.87 |
|
|
|
| 5 |
Π |
311i |
306i |
50.14 |
|
|
|
| 5 |
Π |
311i |
306i |
50.14 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 3349.4 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 3294.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBEultrafine/6-31G*
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-1.220 |
| C2 |
0.000 |
0.000 |
0.081 |
| N3 |
0.000 |
0.000 |
1.304 |
| H4 |
0.000 |
0.000 |
-2.294 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
| C1 | | 1.3005 | 2.5237 | 1.0745 |
C2 | 1.3005 | | 1.2232 | 2.3750 | N3 | 2.5237 | 1.2232 | | 3.5982 | H4 | 1.0745 | 2.3750 | 3.5982 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
180.000 |
|
C2 |
C1 |
H4 |
180.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.076 |
|
|
|
| 2 |
C |
0.303 |
|
|
|
| 3 |
N |
-0.467 |
|
|
|
| 4 |
H |
0.240 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-3.209 |
3.209 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-16.898 |
0.000 |
0.000 |
| y |
0.000 |
-16.898 |
0.000 |
| z |
0.000 |
0.000 |
-15.074 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.912 |
0.000 |
0.000 |
| y |
0.000 |
-0.912 |
0.000 |
| z |
0.000 |
0.000 |
1.824 |
|
| Polar |
|---|
| 3z2-r2 | 3.648 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
1.644 |
0.000 |
0.000 |
| y |
0.000 |
1.644 |
0.000 |
| z |
0.000 |
0.000 |
6.289 |
<r2> (average value of r
2) Å
2
| <r2> |
36.306 |
| (<r2>)1/2 |
6.025 |
Jump to
S1C1
S1C2
S2C2
Energy calculated at PBEPBEultrafine/6-31G*
| | hartrees |
|---|
| Energy at 0K | -131.232976 |
| Energy at 298.15K | -131.232369 |
| HF Energy | -131.232976 |
| Nuclear repulsion energy | 46.555787 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3007 |
2957 |
15.10 |
|
|
|
| 2 |
A' |
2029 |
1996 |
13.88 |
|
|
|
| 3 |
A' |
1099 |
1081 |
19.71 |
|
|
|
| 4 |
A' |
894 |
879 |
66.86 |
|
|
|
| 5 |
A' |
435 |
428 |
19.51 |
|
|
|
| 6 |
A" |
301 |
297 |
7.32 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 3882.7 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 3818.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBEultrafine/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.334 |
-1.242 |
0.000 |
| C2 |
0.000 |
0.090 |
0.000 |
| N3 |
-0.491 |
1.188 |
0.000 |
| H4 |
1.432 |
-1.404 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
| C1 | | 1.3740 | 2.5664 | 1.1099 |
C2 | 1.3740 | | 1.2022 | 2.0694 | N3 | 2.5664 | 1.2022 | | 3.2268 | H4 | 1.1099 | 2.0694 | 3.2268 | |
More geometry information
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.148 |
|
|
|
| 2 |
C |
0.336 |
|
|
|
| 3 |
N |
-0.390 |
|
|
|
| 4 |
H |
0.202 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.923 |
-0.967 |
0.000 |
2.152 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-15.983 |
-1.382 |
0.000 |
| y |
-1.382 |
-21.096 |
0.000 |
| z |
0.000 |
0.000 |
-15.553 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.341 |
-1.382 |
0.000 |
| y |
-1.382 |
-5.328 |
0.000 |
| z |
0.000 |
0.000 |
2.987 |
|
| Polar |
|---|
| 3z2-r2 | 5.973 |
|---|
| x2-y2 | 5.113 |
|---|
| xy | -1.382 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.918 |
-1.514 |
0.000 |
| y |
-1.514 |
5.833 |
0.000 |
| z |
0.000 |
0.000 |
2.146 |
<r2> (average value of r
2) Å
2
| <r2> |
36.521 |
| (<r2>)1/2 |
6.043 |
Jump to
S1C1
S1C2
S2C1
Energy calculated at PBEPBEultrafine/6-31G*
| | hartrees |
|---|
| Energy at 0K | -131.232976 |
| Energy at 298.15K | -131.232369 |
| HF Energy | -131.232976 |
| Nuclear repulsion energy | 46.555787 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
Geometric Data calculated at PBEPBEultrafine/6-31G*
Point Group is Cs
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
