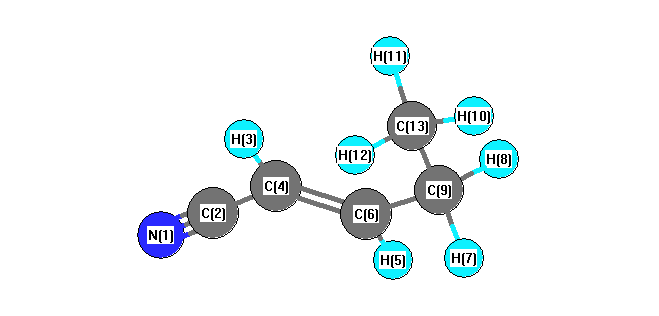
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3211 |
3072 |
2.08 |
|
|
|
| 2 |
A' |
3185 |
3047 |
7.42 |
|
|
|
| 3 |
A' |
3147 |
3010 |
22.42 |
|
|
|
| 4 |
A' |
3064 |
2931 |
17.16 |
|
|
|
| 5 |
A' |
3031 |
2900 |
16.16 |
|
|
|
| 6 |
A' |
2353 |
2251 |
28.47 |
|
|
|
| 7 |
A' |
1720 |
1645 |
28.48 |
|
|
|
| 8 |
A' |
1532 |
1466 |
5.83 |
|
|
|
| 9 |
A' |
1488 |
1424 |
12.18 |
|
|
|
| 10 |
A' |
1442 |
1379 |
6.89 |
|
|
|
| 11 |
A' |
1396 |
1336 |
8.13 |
|
|
|
| 12 |
A' |
1345 |
1287 |
1.33 |
|
|
|
| 13 |
A' |
1307 |
1250 |
2.25 |
|
|
|
| 14 |
A' |
1125 |
1076 |
3.08 |
|
|
|
| 15 |
A' |
1081 |
1035 |
4.55 |
|
|
|
| 16 |
A' |
1000 |
956 |
1.90 |
|
|
|
| 17 |
A' |
910 |
871 |
4.30 |
|
|
|
| 18 |
A' |
580 |
555 |
0.62 |
|
|
|
| 19 |
A' |
511 |
488 |
0.07 |
|
|
|
| 20 |
A' |
259 |
248 |
1.37 |
|
|
|
| 21 |
A' |
147 |
140 |
4.08 |
|
|
|
| 22 |
A" |
3139 |
3003 |
24.90 |
|
|
|
| 23 |
A" |
3057 |
2925 |
10.16 |
|
|
|
| 24 |
A" |
1527 |
1461 |
8.79 |
|
|
|
| 25 |
A" |
1300 |
1244 |
0.10 |
|
|
|
| 26 |
A" |
1110 |
1062 |
1.32 |
|
|
|
| 27 |
A" |
1003 |
960 |
30.23 |
|
|
|
| 28 |
A" |
854 |
817 |
9.68 |
|
|
|
| 29 |
A" |
740 |
708 |
0.49 |
|
|
|
| 30 |
A" |
496 |
474 |
6.70 |
|
|
|
| 31 |
A" |
281 |
269 |
0.01 |
|
|
|
| 32 |
A" |
180 |
172 |
0.02 |
|
|
|
| 33 |
A" |
124 |
119 |
3.75 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23821.6 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 22790.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.492 |
|
|
-0.455 |
| 2 |
C |
0.322 |
|
|
0.411 |
| 3 |
H |
0.208 |
|
|
0.196 |
| 4 |
C |
-0.181 |
|
|
-0.338 |
| 5 |
H |
0.191 |
|
|
0.122 |
| 6 |
C |
-0.092 |
|
|
-0.078 |
| 7 |
H |
0.193 |
|
|
0.025 |
| 8 |
H |
0.193 |
|
|
0.025 |
| 9 |
C |
-0.362 |
|
|
0.131 |
| 10 |
H |
0.183 |
|
|
0.081 |
| 11 |
H |
0.182 |
|
|
0.083 |
| 12 |
H |
0.182 |
|
|
0.083 |
| 13 |
C |
-0.527 |
|
|
-0.287 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.147 |
-4.374 |
0.000 |
4.873 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
2.159 |
-4.347 |
0.000 |
4.853 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.398 |
6.961 |
0.000 |
| y |
6.961 |
-44.556 |
0.000 |
| z |
0.000 |
0.000 |
-36.657 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.209 |
6.961 |
0.000 |
| y |
6.961 |
-7.528 |
0.000 |
| z |
0.000 |
0.000 |
4.320 |
|
| Polar |
|---|
| 3z2-r2 | 8.639 |
|---|
| x2-y2 | 7.158 |
|---|
| xy | 6.961 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
226.662 |
| (<r2>)1/2 |
15.055 |