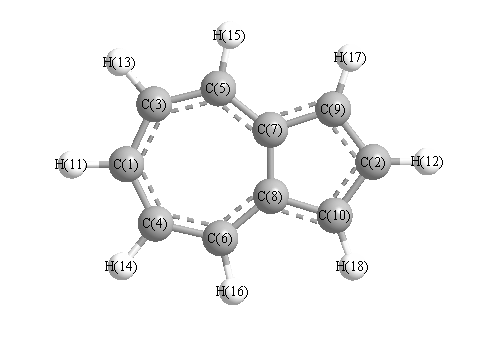
Vibrational Frequencies calculated at B2PLYP/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3265 |
3100 |
14.41 |
|
|
|
| 2 |
A1 |
3240 |
3076 |
6.98 |
|
|
|
| 3 |
A1 |
3213 |
3050 |
30.11 |
|
|
|
| 4 |
A1 |
3185 |
3023 |
7.01 |
|
|
|
| 5 |
A1 |
3174 |
3012 |
0.39 |
|
|
|
| 6 |
A1 |
1658 |
1574 |
50.43 |
|
|
|
| 7 |
A1 |
1606 |
1524 |
5.22 |
|
|
|
| 8 |
A1 |
1513 |
1437 |
18.39 |
|
|
|
| 9 |
A1 |
1449 |
1375 |
78.13 |
|
|
|
| 10 |
A1 |
1321 |
1253 |
0.13 |
|
|
|
| 11 |
A1 |
1263 |
1199 |
0.52 |
|
|
|
| 12 |
A1 |
1099 |
1043 |
5.14 |
|
|
|
| 13 |
A1 |
973 |
923 |
1.14 |
|
|
|
| 14 |
A1 |
920 |
874 |
2.41 |
|
|
|
| 15 |
A1 |
837 |
795 |
4.29 |
|
|
|
| 16 |
A1 |
684 |
650 |
0.96 |
|
|
|
| 17 |
A1 |
413 |
392 |
1.22 |
|
|
|
| 18 |
A2 |
974 |
924 |
0.00 |
|
|
|
| 19 |
A2 |
869 |
824 |
0.00 |
|
|
|
| 20 |
A2 |
772 |
733 |
0.00 |
|
|
|
| 21 |
A2 |
695 |
660 |
0.00 |
|
|
|
| 22 |
A2 |
425 |
403 |
0.00 |
|
|
|
| 23 |
A2 |
161 |
153 |
0.00 |
|
|
|
| 24 |
B1 |
978 |
928 |
0.06 |
|
|
|
| 25 |
B1 |
967 |
917 |
5.07 |
|
|
|
| 26 |
B1 |
908 |
862 |
0.72 |
|
|
|
| 27 |
B1 |
773 |
734 |
104.90 |
|
|
|
| 28 |
B1 |
744 |
706 |
0.75 |
|
|
|
| 29 |
B1 |
596 |
566 |
1.08 |
|
|
|
| 30 |
B1 |
561 |
532 |
3.84 |
|
|
|
| 31 |
B1 |
314 |
298 |
6.68 |
|
|
|
| 32 |
B1 |
171 |
162 |
1.31 |
|
|
|
| 33 |
B2 |
3256 |
3091 |
13.67 |
|
|
|
| 34 |
B2 |
3204 |
3042 |
34.96 |
|
|
|
| 35 |
B2 |
3175 |
3013 |
16.63 |
|
|
|
| 36 |
B2 |
1670 |
1585 |
1.64 |
|
|
|
| 37 |
B2 |
1567 |
1487 |
9.77 |
|
|
|
| 38 |
B2 |
1514 |
1437 |
6.55 |
|
|
|
| 39 |
B2 |
1454 |
1380 |
0.15 |
|
|
|
| 40 |
B2 |
1371 |
1301 |
0.00 |
|
|
|
| 41 |
B2 |
1344 |
1276 |
3.59 |
|
|
|
| 42 |
B2 |
1269 |
1205 |
8.34 |
|
|
|
| 43 |
B2 |
1209 |
1148 |
0.67 |
|
|
|
| 44 |
B2 |
1078 |
1023 |
0.16 |
|
|
|
| 45 |
B2 |
1063 |
1009 |
14.82 |
|
|
|
| 46 |
B2 |
760 |
721 |
0.48 |
|
|
|
| 47 |
B2 |
500 |
475 |
1.73 |
|
|
|
| 48 |
B2 |
336 |
319 |
0.73 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 32243.8 cm
-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 30605.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.124 |
|
|
|
| 2 |
C |
-0.160 |
|
|
|
| 3 |
C |
-0.161 |
|
|
|
| 4 |
C |
-0.161 |
|
|
|
| 5 |
C |
-0.172 |
|
|
|
| 6 |
C |
-0.172 |
|
|
|
| 7 |
C |
0.083 |
|
|
|
| 8 |
C |
0.083 |
|
|
|
| 9 |
C |
-0.206 |
|
|
|
| 10 |
C |
-0.206 |
|
|
|
| 11 |
H |
0.153 |
|
|
|
| 12 |
H |
0.148 |
|
|
|
| 13 |
H |
0.149 |
|
|
|
| 14 |
H |
0.149 |
|
|
|
| 15 |
H |
0.156 |
|
|
|
| 16 |
H |
0.156 |
|
|
|
| 17 |
H |
0.143 |
|
|
|
| 18 |
H |
0.143 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-1.034 |
1.034 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-62.761 |
0.000 |
0.000 |
| y |
0.000 |
-50.720 |
0.000 |
| z |
0.000 |
0.000 |
-51.424 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-11.689 |
0.000 |
0.000 |
| y |
0.000 |
6.373 |
0.000 |
| z |
0.000 |
0.000 |
5.316 |
|
| Polar |
|---|
| 3z2-r2 | 10.632 |
|---|
| x2-y2 | -12.041 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.830 |
0.000 |
0.000 |
| y |
0.000 |
17.164 |
0.000 |
| z |
0.000 |
0.000 |
25.962 |
<r2> (average value of r
2) Å
2
| <r2> |
366.317 |
| (<r2>)1/2 |
19.139 |