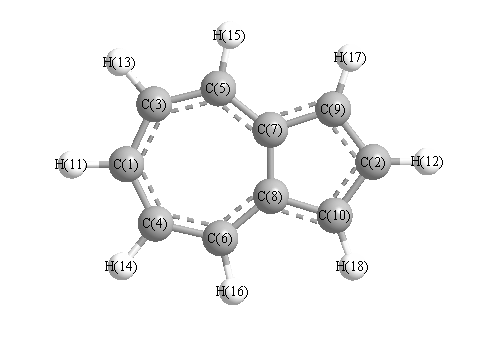
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3271 |
3111 |
13.82 |
|
|
|
| 2 |
A1 |
3245 |
3086 |
6.63 |
|
|
|
| 3 |
A1 |
3215 |
3058 |
25.35 |
|
|
|
| 4 |
A1 |
3187 |
3031 |
7.32 |
|
|
|
| 5 |
A1 |
3176 |
3021 |
0.27 |
|
|
|
| 6 |
A1 |
1680 |
1597 |
60.10 |
|
|
|
| 7 |
A1 |
1613 |
1534 |
7.93 |
|
|
|
| 8 |
A1 |
1518 |
1443 |
32.14 |
|
|
|
| 9 |
A1 |
1453 |
1381 |
75.43 |
|
|
|
| 10 |
A1 |
1327 |
1262 |
0.03 |
|
|
|
| 11 |
A1 |
1258 |
1197 |
0.30 |
|
|
|
| 12 |
A1 |
1098 |
1044 |
6.79 |
|
|
|
| 13 |
A1 |
981 |
932 |
1.39 |
|
|
|
| 14 |
A1 |
915 |
870 |
2.19 |
|
|
|
| 15 |
A1 |
836 |
795 |
5.03 |
|
|
|
| 16 |
A1 |
692 |
658 |
0.76 |
|
|
|
| 17 |
A1 |
411 |
391 |
1.24 |
|
|
|
| 18 |
A2 |
1001 |
952 |
0.00 |
|
|
|
| 19 |
A2 |
885 |
841 |
0.00 |
|
|
|
| 20 |
A2 |
799 |
760 |
0.00 |
|
|
|
| 21 |
A2 |
731 |
695 |
0.00 |
|
|
|
| 22 |
A2 |
432 |
411 |
0.00 |
|
|
|
| 23 |
A2 |
165 |
156 |
0.00 |
|
|
|
| 24 |
B1 |
1013 |
963 |
0.05 |
|
|
|
| 25 |
B1 |
984 |
936 |
6.51 |
|
|
|
| 26 |
B1 |
942 |
896 |
0.09 |
|
|
|
| 27 |
B1 |
789 |
751 |
106.76 |
|
|
|
| 28 |
B1 |
748 |
711 |
3.63 |
|
|
|
| 29 |
B1 |
614 |
584 |
1.75 |
|
|
|
| 30 |
B1 |
575 |
547 |
6.75 |
|
|
|
| 31 |
B1 |
321 |
305 |
7.79 |
|
|
|
| 32 |
B1 |
171 |
162 |
1.45 |
|
|
|
| 33 |
B2 |
3261 |
3102 |
12.38 |
|
|
|
| 34 |
B2 |
3207 |
3049 |
32.27 |
|
|
|
| 35 |
B2 |
3178 |
3022 |
15.01 |
|
|
|
| 36 |
B2 |
1681 |
1599 |
3.39 |
|
|
|
| 37 |
B2 |
1578 |
1501 |
9.49 |
|
|
|
| 38 |
B2 |
1510 |
1436 |
6.82 |
|
|
|
| 39 |
B2 |
1451 |
1380 |
0.89 |
|
|
|
| 40 |
B2 |
1354 |
1288 |
0.57 |
|
|
|
| 41 |
B2 |
1335 |
1269 |
3.85 |
|
|
|
| 42 |
B2 |
1258 |
1196 |
7.39 |
|
|
|
| 43 |
B2 |
1205 |
1146 |
0.34 |
|
|
|
| 44 |
B2 |
1078 |
1025 |
0.02 |
|
|
|
| 45 |
B2 |
1051 |
1000 |
13.49 |
|
|
|
| 46 |
B2 |
754 |
717 |
0.35 |
|
|
|
| 47 |
B2 |
500 |
475 |
1.68 |
|
|
|
| 48 |
B2 |
334 |
318 |
0.70 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 32387.8 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 30800.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.139 |
|
|
|
| 2 |
C |
-0.180 |
|
|
|
| 3 |
C |
-0.183 |
|
|
|
| 4 |
C |
-0.183 |
|
|
|
| 5 |
C |
-0.190 |
|
|
|
| 6 |
C |
-0.190 |
|
|
|
| 7 |
C |
0.088 |
|
|
|
| 8 |
C |
0.088 |
|
|
|
| 9 |
C |
-0.224 |
|
|
|
| 10 |
C |
-0.224 |
|
|
|
| 11 |
H |
0.171 |
|
|
|
| 12 |
H |
0.165 |
|
|
|
| 13 |
H |
0.167 |
|
|
|
| 14 |
H |
0.167 |
|
|
|
| 15 |
H |
0.174 |
|
|
|
| 16 |
H |
0.174 |
|
|
|
| 17 |
H |
0.159 |
|
|
|
| 18 |
H |
0.159 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-1.089 |
1.089 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-62.698 |
0.000 |
0.000 |
| y |
0.000 |
-49.693 |
0.000 |
| z |
0.000 |
0.000 |
-50.465 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-12.620 |
0.000 |
0.000 |
| y |
0.000 |
6.889 |
0.000 |
| z |
0.000 |
0.000 |
5.731 |
|
| Polar |
|---|
| 3z2-r2 | 11.462 |
|---|
| x2-y2 | -13.006 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.910 |
0.000 |
0.000 |
| y |
0.000 |
17.162 |
0.000 |
| z |
0.000 |
0.000 |
25.844 |
<r2> (average value of r
2) Å
2
| <r2> |
364.321 |
| (<r2>)1/2 |
19.087 |