Jump to
S1C2
Energy calculated at B2PLYP/6-31G*
| | hartrees |
|---|
| Energy at 0K | -344.920391 |
| Energy at 298.15K | -344.937572 |
| HF Energy | -344.554270 |
| Nuclear repulsion energy | 424.419166 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3111 |
2953 |
0.00 |
|
|
|
| 2 |
A1 |
1291 |
1226 |
0.00 |
|
|
|
| 3 |
A1 |
1044 |
991 |
0.00 |
|
|
|
| 4 |
A1 |
81 |
77 |
0.00 |
|
|
|
| 5 |
A1' |
3086 |
2929 |
0.00 |
|
|
|
| 6 |
A1' |
1547 |
1469 |
0.00 |
|
|
|
| 7 |
A1' |
1393 |
1322 |
0.00 |
|
|
|
| 8 |
A1' |
988 |
938 |
0.00 |
|
|
|
| 9 |
A1' |
822 |
780 |
0.00 |
|
|
|
| 10 |
A1' |
613 |
581 |
0.00 |
|
|
|
| 11 |
A2 |
3073 |
2917 |
111.91 |
|
|
|
| 12 |
A2 |
1539 |
1461 |
2.66 |
|
|
|
| 13 |
A2 |
1414 |
1343 |
7.87 |
|
|
|
| 14 |
A2 |
1006 |
955 |
21.10 |
|
|
|
| 15 |
A2 |
776 |
737 |
63.27 |
|
|
|
| 16 |
A2' |
3131 |
2972 |
0.00 |
|
|
|
| 17 |
A2' |
1218 |
1156 |
0.00 |
|
|
|
| 18 |
A2' |
827 |
785 |
0.00 |
|
|
|
| 19 |
E |
3114 |
2956 |
0.00 |
|
|
|
| 19 |
E |
3114 |
2956 |
0.00 |
|
|
|
| 20 |
E |
3070 |
2915 |
0.00 |
|
|
|
| 20 |
E |
3070 |
2914 |
0.00 |
|
|
|
| 21 |
E |
1526 |
1449 |
0.00 |
|
|
|
| 21 |
E |
1526 |
1449 |
0.00 |
|
|
|
| 22 |
E |
1372 |
1303 |
0.00 |
|
|
|
| 22 |
E |
1372 |
1303 |
0.00 |
|
|
|
| 23 |
E |
1354 |
1285 |
0.00 |
|
|
|
| 23 |
E |
1354 |
1285 |
0.00 |
|
|
|
| 24 |
E |
1230 |
1167 |
0.00 |
|
|
|
| 24 |
E |
1230 |
1167 |
0.00 |
|
|
|
| 25 |
E |
1059 |
1005 |
0.00 |
|
|
|
| 25 |
E |
1059 |
1005 |
0.00 |
|
|
|
| 26 |
E |
592 |
562 |
0.00 |
|
|
|
| 26 |
E |
592 |
562 |
0.00 |
|
|
|
| 27 |
E |
339 |
322 |
0.00 |
|
|
|
| 27 |
E |
339 |
322 |
0.00 |
|
|
|
| 28 |
E' |
3139 |
2979 |
91.11 |
|
|
|
| 28 |
E' |
3139 |
2979 |
91.16 |
|
|
|
| 29 |
E' |
3079 |
2922 |
102.10 |
|
|
|
| 29 |
E' |
3079 |
2922 |
102.02 |
|
|
|
| 30 |
E' |
1538 |
1460 |
7.47 |
|
|
|
| 30 |
E' |
1538 |
1460 |
7.48 |
|
|
|
| 31 |
E' |
1378 |
1308 |
8.26 |
|
|
|
| 31 |
E' |
1378 |
1308 |
8.26 |
|
|
|
| 32 |
E' |
1360 |
1291 |
0.08 |
|
|
|
| 32 |
E' |
1360 |
1291 |
0.08 |
|
|
|
| 33 |
E' |
1103 |
1047 |
39.89 |
|
|
|
| 33 |
E' |
1103 |
1047 |
39.89 |
|
|
|
| 34 |
E' |
917 |
870 |
4.36 |
|
|
|
| 34 |
E' |
917 |
870 |
4.36 |
|
|
|
| 35 |
E' |
847 |
804 |
4.24 |
|
|
|
| 35 |
E' |
847 |
804 |
4.24 |
|
|
|
| 36 |
E' |
431 |
409 |
0.00 |
|
|
|
| 36 |
E' |
431 |
409 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 40927.8 cm
-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 38848.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G*
Point Group is D3h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
0.000 |
0.000 |
1.290 |
| N2 |
0.000 |
0.000 |
-1.290 |
| C3 |
0.000 |
1.381 |
0.780 |
| C4 |
1.196 |
-0.690 |
0.780 |
| C5 |
-1.196 |
-0.690 |
0.780 |
| C6 |
0.000 |
1.381 |
-0.780 |
| C7 |
-1.196 |
-0.690 |
-0.780 |
| C8 |
1.196 |
-0.690 |
-0.780 |
| H9 |
0.883 |
1.888 |
1.183 |
| H10 |
-0.883 |
1.888 |
1.183 |
| H11 |
1.194 |
-1.709 |
1.183 |
| H12 |
2.077 |
-0.180 |
1.183 |
| H13 |
-2.077 |
-0.180 |
1.183 |
| H14 |
-1.194 |
-1.709 |
1.183 |
| H15 |
-0.883 |
1.888 |
-1.183 |
| H16 |
0.883 |
1.888 |
-1.183 |
| H17 |
-1.194 |
-1.709 |
-1.183 |
| H18 |
-2.077 |
-0.180 |
-1.183 |
| H19 |
2.077 |
-0.180 |
-1.183 |
| H20 |
1.194 |
-1.709 |
-1.183 |
Atom - Atom Distances (Å)
| |
N1 |
N2 |
C3 |
C4 |
C5 |
C6 |
C7 |
C8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
H19 |
H20 |
| N1 | | 2.5791 | 1.4718 | 1.4718 | 1.4718 | 2.4881 | 2.4881 | 2.4881 | 2.0873 | 2.0873 | 2.0873 | 2.0873 | 2.0873 | 2.0873 | 3.2338 | 3.2338 | 3.2338 | 3.2338 | 3.2338 | 3.2338 |
N2 | 2.5791 | | 2.4881 | 2.4881 | 2.4881 | 1.4718 | 1.4718 | 1.4718 | 3.2338 | 3.2338 | 3.2338 | 3.2338 | 3.2338 | 3.2338 | 2.0873 | 2.0873 | 2.0873 | 2.0873 | 2.0873 | 2.0873 | C3 | 1.4718 | 2.4881 | | 2.3917 | 2.3917 | 1.5605 | 2.8558 | 2.8558 | 1.0950 | 1.0950 | 3.3367 | 2.6288 | 2.6288 | 3.3367 | 2.2113 | 2.2113 | 3.8503 | 3.2560 | 3.2560 | 3.8503 | C4 | 1.4718 | 2.4881 | 2.3917 | | 2.3917 | 2.8558 | 2.8558 | 1.5605 | 2.6288 | 3.3367 | 1.0950 | 1.0950 | 3.3367 | 2.6288 | 3.8503 | 3.2560 | 3.2560 | 3.8503 | 2.2113 | 2.2113 | C5 | 1.4718 | 2.4881 | 2.3917 | 2.3917 | | 2.8558 | 1.5605 | 2.8558 | 3.3367 | 2.6288 | 2.6288 | 3.3367 | 1.0950 | 1.0950 | 3.2560 | 3.8503 | 2.2113 | 2.2113 | 3.8503 | 3.2560 | C6 | 2.4881 | 1.4718 | 1.5605 | 2.8558 | 2.8558 | | 2.3917 | 2.3917 | 2.2113 | 2.2113 | 3.8503 | 3.2560 | 3.2560 | 3.8503 | 1.0950 | 1.0950 | 3.3367 | 2.6288 | 2.6288 | 3.3367 | C7 | 2.4881 | 1.4718 | 2.8558 | 2.8558 | 1.5605 | 2.3917 | | 2.3917 | 3.8503 | 3.2560 | 3.2560 | 3.8503 | 2.2113 | 2.2113 | 2.6288 | 3.3367 | 1.0950 | 1.0950 | 3.3367 | 2.6288 | C8 | 2.4881 | 1.4718 | 2.8558 | 1.5605 | 2.8558 | 2.3917 | 2.3917 | | 3.2560 | 3.8503 | 2.2113 | 2.2113 | 3.8503 | 3.2560 | 3.3367 | 2.6288 | 2.6288 | 3.3367 | 1.0950 | 1.0950 | H9 | 2.0873 | 3.2338 | 1.0950 | 2.6288 | 3.3367 | 2.2113 | 3.8503 | 3.2560 | | 1.7658 | 3.6106 | 2.3879 | 3.6106 | 4.1537 | 2.9517 | 2.3653 | 4.7799 | 4.3164 | 3.3611 | 4.3164 | H10 | 2.0873 | 3.2338 | 1.0950 | 3.3367 | 2.6288 | 2.2113 | 3.2560 | 3.8503 | 1.7658 | | 4.1537 | 3.6106 | 2.3879 | 3.6106 | 2.3653 | 2.9517 | 4.3164 | 3.3611 | 4.3164 | 4.7799 | H11 | 2.0873 | 3.2338 | 3.3367 | 1.0950 | 2.6288 | 3.8503 | 3.2560 | 2.2113 | 3.6106 | 4.1537 | | 1.7658 | 3.6106 | 2.3879 | 4.7799 | 4.3164 | 3.3611 | 4.3164 | 2.9517 | 2.3653 | H12 | 2.0873 | 3.2338 | 2.6288 | 1.0950 | 3.3367 | 3.2560 | 3.8503 | 2.2113 | 2.3879 | 3.6106 | 1.7658 | | 4.1537 | 3.6106 | 4.3164 | 3.3611 | 4.3164 | 4.7799 | 2.3653 | 2.9517 | H13 | 2.0873 | 3.2338 | 2.6288 | 3.3367 | 1.0950 | 3.2560 | 2.2113 | 3.8503 | 3.6106 | 2.3879 | 3.6106 | 4.1537 | | 1.7658 | 3.3611 | 4.3164 | 2.9517 | 2.3653 | 4.7799 | 4.3164 | H14 | 2.0873 | 3.2338 | 3.3367 | 2.6288 | 1.0950 | 3.8503 | 2.2113 | 3.2560 | 4.1537 | 3.6106 | 2.3879 | 3.6106 | 1.7658 | | 4.3164 | 4.7799 | 2.3653 | 2.9517 | 4.3164 | 3.3611 | H15 | 3.2338 | 2.0873 | 2.2113 | 3.8503 | 3.2560 | 1.0950 | 2.6288 | 3.3367 | 2.9517 | 2.3653 | 4.7799 | 4.3164 | 3.3611 | 4.3164 | | 1.7658 | 3.6106 | 2.3879 | 3.6106 | 4.1537 | H16 | 3.2338 | 2.0873 | 2.2113 | 3.2560 | 3.8503 | 1.0950 | 3.3367 | 2.6288 | 2.3653 | 2.9517 | 4.3164 | 3.3611 | 4.3164 | 4.7799 | 1.7658 | | 4.1537 | 3.6106 | 2.3879 | 3.6106 | H17 | 3.2338 | 2.0873 | 3.8503 | 3.2560 | 2.2113 | 3.3367 | 1.0950 | 2.6288 | 4.7799 | 4.3164 | 3.3611 | 4.3164 | 2.9517 | 2.3653 | 3.6106 | 4.1537 | | 1.7658 | 3.6106 | 2.3879 | H18 | 3.2338 | 2.0873 | 3.2560 | 3.8503 | 2.2113 | 2.6288 | 1.0950 | 3.3367 | 4.3164 | 3.3611 | 4.3164 | 4.7799 | 2.3653 | 2.9517 | 2.3879 | 3.6106 | 1.7658 | | 4.1537 | 3.6106 | H19 | 3.2338 | 2.0873 | 3.2560 | 2.2113 | 3.8503 | 2.6288 | 3.3367 | 1.0950 | 3.3611 | 4.3164 | 2.9517 | 2.3653 | 4.7799 | 4.3164 | 3.6106 | 2.3879 | 3.6106 | 4.1537 | | 1.7658 | H20 | 3.2338 | 2.0873 | 3.8503 | 2.2113 | 3.2560 | 3.3367 | 2.6288 | 1.0950 | 4.3164 | 4.7799 | 2.3653 | 2.9517 | 4.3164 | 3.3611 | 4.1537 | 3.6106 | 2.3879 | 3.6106 | 1.7658 | |
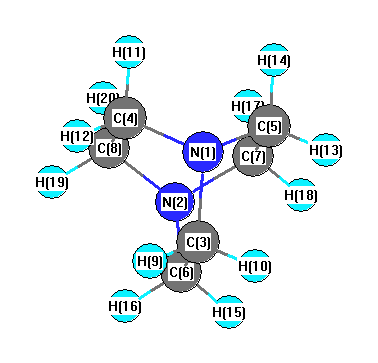 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
C3 |
C6 |
110.245 |
|
N1 |
C3 |
H9 |
107.921 |
| N1 |
C3 |
H10 |
107.921 |
|
N1 |
C4 |
C8 |
110.245 |
| N1 |
C4 |
H11 |
107.921 |
|
N1 |
C4 |
H12 |
107.921 |
| N1 |
C5 |
C7 |
110.245 |
|
N1 |
C5 |
H13 |
107.921 |
| N1 |
C5 |
H14 |
107.921 |
|
N2 |
C6 |
C3 |
110.245 |
| N2 |
C6 |
H15 |
107.921 |
|
N2 |
C6 |
H16 |
107.921 |
| N2 |
C7 |
C5 |
110.245 |
|
N2 |
C7 |
H17 |
107.921 |
| N2 |
C7 |
H18 |
107.921 |
|
N2 |
C8 |
C4 |
110.245 |
| N2 |
C8 |
H19 |
107.921 |
|
N2 |
C8 |
H20 |
107.921 |
| C3 |
N1 |
C4 |
108.686 |
|
C3 |
N1 |
C5 |
108.686 |
| C3 |
C6 |
H15 |
111.561 |
|
C3 |
C6 |
H16 |
111.561 |
| C4 |
N1 |
C5 |
108.686 |
|
C4 |
C8 |
H19 |
111.561 |
| C4 |
C8 |
H20 |
111.561 |
|
C5 |
C6 |
H15 |
101.499 |
| C5 |
C6 |
H16 |
151.001 |
|
C6 |
N2 |
C7 |
108.686 |
| C6 |
N2 |
C8 |
108.686 |
|
C6 |
C3 |
H9 |
111.561 |
| C6 |
C3 |
H10 |
111.561 |
|
C7 |
N2 |
C8 |
108.686 |
| C7 |
C5 |
H13 |
111.561 |
|
C7 |
C5 |
H14 |
111.561 |
| C8 |
C4 |
H11 |
111.561 |
|
C8 |
C4 |
H12 |
111.561 |
| H9 |
C3 |
H10 |
107.471 |
|
H11 |
C4 |
H12 |
107.471 |
| H13 |
C5 |
H14 |
107.471 |
|
H15 |
C6 |
H16 |
107.471 |
| H17 |
C7 |
H18 |
107.471 |
|
H19 |
C8 |
H20 |
107.471 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.440 |
|
|
|
| 2 |
N |
-0.440 |
|
|
|
| 3 |
C |
-0.163 |
|
|
|
| 4 |
C |
-0.163 |
|
|
|
| 5 |
C |
-0.163 |
|
|
|
| 6 |
C |
-0.163 |
|
|
|
| 7 |
C |
-0.163 |
|
|
|
| 8 |
C |
-0.163 |
|
|
|
| 9 |
H |
0.155 |
|
|
|
| 10 |
H |
0.155 |
|
|
|
| 11 |
H |
0.155 |
|
|
|
| 12 |
H |
0.155 |
|
|
|
| 13 |
H |
0.155 |
|
|
|
| 14 |
H |
0.155 |
|
|
|
| 15 |
H |
0.155 |
|
|
|
| 16 |
H |
0.155 |
|
|
|
| 17 |
H |
0.155 |
|
|
|
| 18 |
H |
0.155 |
|
|
|
| 19 |
H |
0.155 |
|
|
|
| 20 |
H |
0.155 |
|
|
|
Electric dipole moments
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.780 |
0.000 |
0.000 |
| y |
0.000 |
10.780 |
0.000 |
| z |
0.000 |
0.000 |
9.222 |
<r2> (average value of r
2) Å
2
| <r2> |
214.280 |
| (<r2>)1/2 |
14.638 |
Jump to
S1C1
Energy calculated at B2PLYP/6-31G*
| | hartrees |
|---|
| Energy at 0K | -344.920386 |
| Energy at 298.15K | -344.937553 |
| HF Energy | -344.554266 |
| Nuclear repulsion energy | 424.416145 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3111 |
2953 |
0.00 |
|
|
|
| 2 |
A1 |
3086 |
2929 |
0.00 |
|
|
|
| 3 |
A1 |
1548 |
1469 |
0.00 |
|
|
|
| 4 |
A1 |
1393 |
1322 |
0.00 |
|
|
|
| 5 |
A1 |
1291 |
1226 |
0.00 |
|
|
|
| 6 |
A1 |
1044 |
991 |
0.00 |
|
|
|
| 7 |
A1 |
988 |
938 |
0.00 |
|
|
|
| 8 |
A1 |
822 |
780 |
0.00 |
|
|
|
| 9 |
A1 |
613 |
582 |
0.00 |
|
|
|
| 10 |
A1 |
76 |
72 |
0.00 |
|
|
|
| 11 |
A2 |
3131 |
2972 |
0.00 |
|
|
|
| 12 |
A2 |
3073 |
2917 |
111.93 |
|
|
|
| 13 |
A2 |
1539 |
1461 |
2.65 |
|
|
|
| 14 |
A2 |
1414 |
1342 |
7.89 |
|
|
|
| 15 |
A2 |
1217 |
1155 |
0.00 |
|
|
|
| 16 |
A2 |
1006 |
955 |
21.17 |
|
|
|
| 17 |
A2 |
826 |
784 |
0.00 |
|
|
|
| 18 |
A2 |
776 |
736 |
63.19 |
|
|
|
| 19 |
E |
3139 |
2979 |
91.03 |
|
|
|
| 19 |
E |
3139 |
2979 |
91.08 |
|
|
|
| 20 |
E |
3114 |
2956 |
0.01 |
|
|
|
| 20 |
E |
3114 |
2956 |
0.01 |
|
|
|
| 21 |
E |
3079 |
2922 |
102.16 |
|
|
|
| 21 |
E |
3079 |
2922 |
102.08 |
|
|
|
| 22 |
E |
3071 |
2915 |
0.00 |
|
|
|
| 22 |
E |
3071 |
2915 |
0.00 |
|
|
|
| 23 |
E |
1538 |
1460 |
7.48 |
|
|
|
| 23 |
E |
1538 |
1460 |
7.49 |
|
|
|
| 24 |
E |
1526 |
1448 |
0.00 |
|
|
|
| 24 |
E |
1526 |
1448 |
0.00 |
|
|
|
| 25 |
E |
1378 |
1308 |
8.26 |
|
|
|
| 25 |
E |
1378 |
1308 |
8.25 |
|
|
|
| 26 |
E |
1372 |
1303 |
0.00 |
|
|
|
| 26 |
E |
1372 |
1303 |
0.00 |
|
|
|
| 27 |
E |
1360 |
1291 |
0.08 |
|
|
|
| 27 |
E |
1360 |
1291 |
0.08 |
|
|
|
| 28 |
E |
1354 |
1285 |
0.00 |
|
|
|
| 28 |
E |
1354 |
1285 |
0.00 |
|
|
|
| 29 |
E |
1230 |
1167 |
0.00 |
|
|
|
| 29 |
E |
1230 |
1167 |
0.00 |
|
|
|
| 30 |
E |
1103 |
1047 |
39.87 |
|
|
|
| 30 |
E |
1103 |
1047 |
39.87 |
|
|
|
| 31 |
E |
1059 |
1005 |
0.00 |
|
|
|
| 31 |
E |
1059 |
1005 |
0.00 |
|
|
|
| 32 |
E |
917 |
870 |
4.34 |
|
|
|
| 32 |
E |
917 |
870 |
4.34 |
|
|
|
| 33 |
E |
847 |
804 |
4.27 |
|
|
|
| 33 |
E |
847 |
804 |
4.27 |
|
|
|
| 34 |
E |
592 |
562 |
0.00 |
|
|
|
| 34 |
E |
592 |
562 |
0.00 |
|
|
|
| 35 |
E |
431 |
409 |
0.00 |
|
|
|
| 35 |
E |
431 |
409 |
0.00 |
|
|
|
| 36 |
E |
338 |
321 |
0.00 |
|
|
|
| 36 |
E |
338 |
321 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 40921.2 cm
-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 38842.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G*
Point Group is D3
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
0.000 |
0.000 |
1.290 |
| N2 |
0.000 |
0.000 |
-1.290 |
| C3 |
-0.001 |
1.381 |
0.780 |
| C4 |
1.196 |
-0.690 |
0.780 |
| C5 |
-1.195 |
-0.691 |
0.780 |
| C6 |
0.001 |
1.381 |
-0.780 |
| C7 |
-1.196 |
-0.690 |
-0.780 |
| C8 |
1.195 |
-0.691 |
-0.780 |
| H9 |
0.881 |
1.890 |
1.184 |
| H10 |
-0.885 |
1.887 |
1.182 |
| H11 |
1.196 |
-1.708 |
1.184 |
| H12 |
2.077 |
-0.177 |
1.182 |
| H13 |
-2.077 |
-0.182 |
1.184 |
| H14 |
-1.192 |
-1.710 |
1.182 |
| H15 |
-0.881 |
1.890 |
-1.184 |
| H16 |
0.885 |
1.887 |
-1.182 |
| H17 |
-1.196 |
-1.708 |
-1.184 |
| H18 |
-2.077 |
-0.177 |
-1.182 |
| H19 |
2.077 |
-0.182 |
-1.184 |
| H20 |
1.192 |
-1.710 |
-1.182 |
Atom - Atom Distances (Å)
| |
N1 |
N2 |
C3 |
C4 |
C5 |
C6 |
C7 |
C8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
H19 |
H20 |
| N1 | | 2.5790 | 1.4718 | 1.4718 | 1.4718 | 2.4881 | 2.4881 | 2.4881 | 2.0875 | 2.0873 | 2.0875 | 2.0873 | 2.0875 | 2.0873 | 3.2347 | 3.2329 | 3.2347 | 3.2329 | 3.2347 | 3.2329 |
N2 | 2.5790 | | 2.4881 | 2.4881 | 2.4881 | 1.4718 | 1.4718 | 1.4718 | 3.2347 | 3.2329 | 3.2347 | 3.2329 | 3.2347 | 3.2329 | 2.0875 | 2.0873 | 2.0875 | 2.0873 | 2.0875 | 2.0873 | C3 | 1.4718 | 2.4881 | | 2.3918 | 2.3918 | 1.5604 | 2.8551 | 2.8565 | 1.0950 | 1.0950 | 3.3368 | 2.6281 | 2.6296 | 3.3367 | 2.2114 | 2.2113 | 3.8502 | 3.2538 | 3.2582 | 3.8504 | C4 | 1.4718 | 2.4881 | 2.3918 | | 2.3918 | 2.8551 | 2.8565 | 1.5604 | 2.6296 | 3.3367 | 1.0950 | 1.0950 | 3.3368 | 2.6281 | 3.8503 | 3.2538 | 3.2582 | 3.8504 | 2.2114 | 2.2113 | C5 | 1.4718 | 2.4881 | 2.3918 | 2.3918 | | 2.8565 | 1.5604 | 2.8551 | 3.3368 | 2.6281 | 2.6296 | 3.3367 | 1.0950 | 1.0950 | 3.2582 | 3.8504 | 2.2114 | 2.2113 | 3.8503 | 3.2538 | C6 | 2.4881 | 1.4718 | 1.5604 | 2.8551 | 2.8565 | | 2.3918 | 2.3918 | 2.2114 | 2.2113 | 3.8502 | 3.2538 | 3.2582 | 3.8504 | 1.0950 | 1.0950 | 3.3368 | 2.6281 | 2.6296 | 3.3367 | C7 | 2.4881 | 1.4718 | 2.8551 | 2.8565 | 1.5604 | 2.3918 | | 2.3918 | 3.8503 | 3.2538 | 3.2582 | 3.8504 | 2.2114 | 2.2113 | 2.6296 | 3.3367 | 1.0950 | 1.0950 | 3.3368 | 2.6281 | C8 | 2.4881 | 1.4718 | 2.8565 | 1.5604 | 2.8551 | 2.3918 | 2.3918 | | 3.2582 | 3.8504 | 2.2114 | 2.2113 | 3.8503 | 3.2538 | 3.3368 | 2.6281 | 2.6296 | 3.3367 | 1.0950 | 1.0950 | H9 | 2.0875 | 3.2347 | 1.0950 | 2.6296 | 3.3368 | 2.2114 | 3.8503 | 3.2582 | | 1.7657 | 3.6110 | 2.3881 | 3.6110 | 4.1538 | 2.9510 | 2.3653 | 4.7810 | 4.3146 | 3.3653 | 4.3183 | H10 | 2.0873 | 3.2329 | 1.0950 | 3.3367 | 2.6281 | 2.2113 | 3.2538 | 3.8504 | 1.7657 | | 4.1538 | 3.6105 | 2.3881 | 3.6105 | 2.3653 | 2.9523 | 4.3146 | 3.3571 | 4.3183 | 4.7791 | H11 | 2.0875 | 3.2347 | 3.3368 | 1.0950 | 2.6296 | 3.8502 | 3.2582 | 2.2114 | 3.6110 | 4.1538 | | 1.7657 | 3.6110 | 2.3881 | 4.7810 | 4.3146 | 3.3653 | 4.3183 | 2.9510 | 2.3653 | H12 | 2.0873 | 3.2329 | 2.6281 | 1.0950 | 3.3367 | 3.2538 | 3.8504 | 2.2113 | 2.3881 | 3.6105 | 1.7657 | | 4.1538 | 3.6105 | 4.3146 | 3.3571 | 4.3183 | 4.7791 | 2.3653 | 2.9523 | H13 | 2.0875 | 3.2347 | 2.6296 | 3.3368 | 1.0950 | 3.2582 | 2.2114 | 3.8503 | 3.6110 | 2.3881 | 3.6110 | 4.1538 | | 1.7657 | 3.3653 | 4.3183 | 2.9510 | 2.3653 | 4.7810 | 4.3146 | H14 | 2.0873 | 3.2329 | 3.3367 | 2.6281 | 1.0950 | 3.8504 | 2.2113 | 3.2538 | 4.1538 | 3.6105 | 2.3881 | 3.6105 | 1.7657 | | 4.3183 | 4.7791 | 2.3653 | 2.9523 | 4.3146 | 3.3571 | H15 | 3.2347 | 2.0875 | 2.2114 | 3.8503 | 3.2582 | 1.0950 | 2.6296 | 3.3368 | 2.9510 | 2.3653 | 4.7810 | 4.3146 | 3.3653 | 4.3183 | | 1.7657 | 3.6110 | 2.3881 | 3.6110 | 4.1538 | H16 | 3.2329 | 2.0873 | 2.2113 | 3.2538 | 3.8504 | 1.0950 | 3.3367 | 2.6281 | 2.3653 | 2.9523 | 4.3146 | 3.3571 | 4.3183 | 4.7791 | 1.7657 | | 4.1538 | 3.6105 | 2.3881 | 3.6105 | H17 | 3.2347 | 2.0875 | 3.8502 | 3.2582 | 2.2114 | 3.3368 | 1.0950 | 2.6296 | 4.7810 | 4.3146 | 3.3653 | 4.3183 | 2.9510 | 2.3653 | 3.6110 | 4.1538 | | 1.7657 | 3.6110 | 2.3881 | H18 | 3.2329 | 2.0873 | 3.2538 | 3.8504 | 2.2113 | 2.6281 | 1.0950 | 3.3367 | 4.3146 | 3.3571 | 4.3183 | 4.7791 | 2.3653 | 2.9523 | 2.3881 | 3.6105 | 1.7657 | | 4.1538 | 3.6105 | H19 | 3.2347 | 2.0875 | 3.2582 | 2.2114 | 3.8503 | 2.6296 | 3.3368 | 1.0950 | 3.3653 | 4.3183 | 2.9510 | 2.3653 | 4.7810 | 4.3146 | 3.6110 | 2.3881 | 3.6110 | 4.1538 | | 1.7657 | H20 | 3.2329 | 2.0873 | 3.8504 | 2.2113 | 3.2538 | 3.3367 | 2.6281 | 1.0950 | 4.3183 | 4.7791 | 2.3653 | 2.9523 | 4.3146 | 3.3571 | 4.1538 | 3.6105 | 2.3881 | 3.6105 | 1.7657 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
C3 |
C6 |
110.245 |
|
N1 |
C3 |
H9 |
107.932 |
| N1 |
C3 |
H10 |
107.914 |
|
N1 |
C4 |
C8 |
110.245 |
| N1 |
C4 |
H11 |
107.932 |
|
N1 |
C4 |
H12 |
107.914 |
| N1 |
C5 |
C7 |
110.245 |
|
N1 |
C5 |
H13 |
107.932 |
| N1 |
C5 |
H14 |
107.914 |
|
N2 |
C6 |
C3 |
110.245 |
| N2 |
C6 |
H15 |
107.932 |
|
N2 |
C6 |
H16 |
107.914 |
| N2 |
C7 |
C5 |
110.245 |
|
N2 |
C7 |
H17 |
107.932 |
| N2 |
C7 |
H18 |
107.914 |
|
N2 |
C8 |
C4 |
110.245 |
| N2 |
C8 |
H19 |
107.932 |
|
N2 |
C8 |
H20 |
107.914 |
| C3 |
N1 |
C4 |
108.686 |
|
C3 |
N1 |
C5 |
108.687 |
| C3 |
C6 |
H15 |
111.567 |
|
C3 |
C6 |
H16 |
111.559 |
| C4 |
N1 |
C5 |
108.687 |
|
C4 |
C8 |
H19 |
111.567 |
| C4 |
C8 |
H20 |
111.559 |
|
C5 |
C6 |
H15 |
101.596 |
| C5 |
C6 |
H16 |
150.912 |
|
C6 |
N2 |
C7 |
108.686 |
| C6 |
N2 |
C8 |
108.687 |
|
C6 |
C3 |
H9 |
111.567 |
| C6 |
C3 |
H10 |
111.559 |
|
C7 |
N2 |
C8 |
108.687 |
| C7 |
C5 |
H13 |
111.567 |
|
C7 |
C5 |
H14 |
111.559 |
| C8 |
C4 |
H11 |
111.567 |
|
C8 |
C4 |
H12 |
111.559 |
| H9 |
C3 |
H10 |
107.463 |
|
H11 |
C4 |
H12 |
107.463 |
| H13 |
C5 |
H14 |
107.463 |
|
H15 |
C6 |
H16 |
107.463 |
| H17 |
C7 |
H18 |
107.463 |
|
H19 |
C8 |
H20 |
107.463 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.393 |
|
|
|
| 2 |
N |
-0.393 |
|
|
|
| 3 |
C |
-0.171 |
|
|
|
| 4 |
C |
-0.171 |
|
|
|
| 5 |
C |
-0.171 |
|
|
|
| 6 |
C |
-0.171 |
|
|
|
| 7 |
C |
-0.171 |
|
|
|
| 8 |
C |
-0.171 |
|
|
|
| 9 |
H |
0.151 |
|
|
|
| 10 |
H |
0.151 |
|
|
|
| 11 |
H |
0.151 |
|
|
|
| 12 |
H |
0.151 |
|
|
|
| 13 |
H |
0.151 |
|
|
|
| 14 |
H |
0.151 |
|
|
|
| 15 |
H |
0.151 |
|
|
|
| 16 |
H |
0.151 |
|
|
|
| 17 |
H |
0.151 |
|
|
|
| 18 |
H |
0.151 |
|
|
|
| 19 |
H |
0.151 |
|
|
|
| 20 |
H |
0.151 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-46.879 |
0.000 |
0.000 |
| y |
0.000 |
-46.879 |
0.000 |
| z |
0.000 |
0.000 |
-57.882 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.502 |
0.000 |
0.000 |
| y |
0.000 |
5.502 |
0.000 |
| z |
0.000 |
0.000 |
-11.003 |
|
| Polar |
|---|
| 3z2-r2 | -22.007 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.780 |
0.000 |
0.000 |
| y |
0.000 |
10.780 |
0.000 |
| z |
0.000 |
0.000 |
9.222 |
<r2> (average value of r
2) Å
2
| <r2> |
214.347 |
| (<r2>)1/2 |
14.641 |
