Jump to
S1C2
Energy calculated at HSEh1PBE/6-31G*
| | hartrees |
|---|
| Energy at 0K | -344.955206 |
| Energy at 298.15K | -344.972417 |
| Nuclear repulsion energy | 426.035339 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1' |
3076 |
2925 |
0.00 |
|
|
|
| 2 |
A1' |
1523 |
1448 |
0.00 |
|
|
|
| 3 |
A1' |
1385 |
1317 |
0.00 |
|
|
|
| 4 |
A1' |
998 |
949 |
0.00 |
|
|
|
| 5 |
A1' |
839 |
798 |
0.00 |
|
|
|
| 6 |
A1' |
613 |
583 |
0.00 |
|
|
|
| 7 |
A1" |
3103 |
2951 |
0.00 |
|
|
|
| 8 |
A1" |
1279 |
1217 |
0.00 |
|
|
|
| 9 |
A1" |
1036 |
985 |
0.00 |
|
|
|
| 10 |
A1" |
91 |
86 |
0.00 |
|
|
|
| 11 |
A2' |
3123 |
2970 |
0.00 |
|
|
|
| 12 |
A2' |
1213 |
1153 |
0.00 |
|
|
|
| 13 |
A2' |
818 |
778 |
0.00 |
|
|
|
| 14 |
A2" |
3062 |
2912 |
115.61 |
|
|
|
| 15 |
A2" |
1515 |
1440 |
5.25 |
|
|
|
| 16 |
A2" |
1403 |
1334 |
5.93 |
|
|
|
| 17 |
A2" |
1011 |
961 |
19.27 |
|
|
|
| 18 |
A2" |
792 |
753 |
66.26 |
|
|
|
| 19 |
E' |
3130 |
2976 |
87.25 |
|
|
|
| 19 |
E' |
3130 |
2976 |
87.19 |
|
|
|
| 20 |
E' |
3068 |
2918 |
108.08 |
|
|
|
| 20 |
E' |
3068 |
2918 |
108.09 |
|
|
|
| 21 |
E' |
1514 |
1440 |
10.20 |
|
|
|
| 21 |
E' |
1514 |
1440 |
10.20 |
|
|
|
| 22 |
E' |
1378 |
1311 |
10.73 |
|
|
|
| 22 |
E' |
1378 |
1311 |
10.74 |
|
|
|
| 23 |
E' |
1350 |
1284 |
0.65 |
|
|
|
| 23 |
E' |
1350 |
1284 |
0.65 |
|
|
|
| 24 |
E' |
1122 |
1067 |
40.09 |
|
|
|
| 24 |
E' |
1122 |
1067 |
40.10 |
|
|
|
| 25 |
E' |
925 |
880 |
3.78 |
|
|
|
| 25 |
E' |
925 |
880 |
3.78 |
|
|
|
| 26 |
E' |
837 |
796 |
4.07 |
|
|
|
| 26 |
E' |
837 |
796 |
4.08 |
|
|
|
| 27 |
E' |
429 |
408 |
0.00 |
|
|
|
| 27 |
E' |
429 |
408 |
0.00 |
|
|
|
| 28 |
E" |
3105 |
2953 |
0.00 |
|
|
|
| 28 |
E" |
3105 |
2953 |
0.00 |
|
|
|
| 29 |
E" |
3060 |
2911 |
0.00 |
|
|
|
| 29 |
E" |
3060 |
2911 |
0.00 |
|
|
|
| 30 |
E" |
1501 |
1428 |
0.00 |
|
|
|
| 30 |
E" |
1501 |
1428 |
0.00 |
|
|
|
| 31 |
E" |
1370 |
1303 |
0.00 |
|
|
|
| 31 |
E" |
1370 |
1303 |
0.00 |
|
|
|
| 32 |
E" |
1344 |
1278 |
0.00 |
|
|
|
| 32 |
E" |
1343 |
1278 |
0.00 |
|
|
|
| 33 |
E" |
1223 |
1163 |
0.00 |
|
|
|
| 33 |
E" |
1223 |
1163 |
0.00 |
|
|
|
| 34 |
E" |
1071 |
1019 |
0.00 |
|
|
|
| 34 |
E" |
1071 |
1019 |
0.00 |
|
|
|
| 35 |
E" |
594 |
565 |
0.00 |
|
|
|
| 35 |
E" |
594 |
565 |
0.00 |
|
|
|
| 36 |
E" |
343 |
326 |
0.00 |
|
|
|
| 36 |
E" |
343 |
326 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 40802.8 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 38803.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G*
Point Group is D3h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
0.000 |
0.000 |
1.281 |
| N2 |
0.000 |
0.000 |
-1.281 |
| C3 |
0.000 |
1.374 |
0.778 |
| C4 |
1.190 |
-0.687 |
0.778 |
| C5 |
-1.190 |
-0.687 |
0.778 |
| C6 |
0.000 |
1.374 |
-0.778 |
| C7 |
-1.190 |
-0.687 |
-0.778 |
| C8 |
1.190 |
-0.687 |
-0.778 |
| H9 |
0.883 |
1.883 |
1.182 |
| H10 |
-0.883 |
1.883 |
1.182 |
| H11 |
1.189 |
-1.707 |
1.182 |
| H12 |
2.072 |
-0.176 |
1.182 |
| H13 |
-2.072 |
-0.176 |
1.182 |
| H14 |
-1.189 |
-1.707 |
1.182 |
| H15 |
-0.883 |
1.883 |
-1.182 |
| H16 |
0.883 |
1.883 |
-1.182 |
| H17 |
-1.189 |
-1.707 |
-1.182 |
| H18 |
-2.072 |
-0.176 |
-1.182 |
| H19 |
2.072 |
-0.176 |
-1.182 |
| H20 |
1.189 |
-1.707 |
-1.182 |
Atom - Atom Distances (Å)
| |
N1 |
N2 |
C3 |
C4 |
C5 |
C6 |
C7 |
C8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
H19 |
H20 |
| N1 | | 2.5624 | 1.4631 | 1.4631 | 1.4631 | 2.4755 | 2.4755 | 2.4755 | 2.0823 | 2.0823 | 2.0823 | 2.0823 | 2.0823 | 2.0823 | 3.2240 | 3.2240 | 3.2240 | 3.2240 | 3.2240 | 3.2240 |
N2 | 2.5624 | | 2.4755 | 2.4755 | 2.4755 | 1.4631 | 1.4631 | 1.4631 | 3.2240 | 3.2240 | 3.2240 | 3.2240 | 3.2240 | 3.2240 | 2.0823 | 2.0823 | 2.0823 | 2.0823 | 2.0823 | 2.0823 | C3 | 1.4631 | 2.4755 | | 2.3795 | 2.3795 | 1.5561 | 2.8432 | 2.8432 | 1.0968 | 1.0968 | 3.3266 | 2.6195 | 2.6195 | 3.3266 | 2.2096 | 2.2096 | 3.8400 | 3.2466 | 3.2466 | 3.8400 | C4 | 1.4631 | 2.4755 | 2.3795 | | 2.3795 | 2.8432 | 2.8432 | 1.5561 | 2.6195 | 3.3266 | 1.0968 | 1.0968 | 3.3266 | 2.6195 | 3.8400 | 3.2466 | 3.2466 | 3.8400 | 2.2096 | 2.2096 | C5 | 1.4631 | 2.4755 | 2.3795 | 2.3795 | | 2.8432 | 1.5561 | 2.8432 | 3.3266 | 2.6195 | 2.6195 | 3.3266 | 1.0968 | 1.0968 | 3.2466 | 3.8400 | 2.2096 | 2.2096 | 3.8400 | 3.2466 | C6 | 2.4755 | 1.4631 | 1.5561 | 2.8432 | 2.8432 | | 2.3795 | 2.3795 | 2.2096 | 2.2096 | 3.8400 | 3.2466 | 3.2466 | 3.8400 | 1.0968 | 1.0968 | 3.3266 | 2.6195 | 2.6195 | 3.3266 | C7 | 2.4755 | 1.4631 | 2.8432 | 2.8432 | 1.5561 | 2.3795 | | 2.3795 | 3.8400 | 3.2466 | 3.2466 | 3.8400 | 2.2096 | 2.2096 | 2.6195 | 3.3266 | 1.0968 | 1.0968 | 3.3266 | 2.6195 | C8 | 2.4755 | 1.4631 | 2.8432 | 1.5561 | 2.8432 | 2.3795 | 2.3795 | | 3.2466 | 3.8400 | 2.2096 | 2.2096 | 3.8400 | 3.2466 | 3.3266 | 2.6195 | 2.6195 | 3.3266 | 1.0968 | 1.0968 | H9 | 2.0823 | 3.2240 | 1.0968 | 2.6195 | 3.3266 | 2.2096 | 3.8400 | 3.2466 | | 1.7670 | 3.6026 | 2.3780 | 3.6026 | 4.1449 | 2.9516 | 2.3643 | 4.7718 | 4.3091 | 3.3533 | 4.3091 | H10 | 2.0823 | 3.2240 | 1.0968 | 3.3266 | 2.6195 | 2.2096 | 3.2466 | 3.8400 | 1.7670 | | 4.1449 | 3.6026 | 2.3780 | 3.6026 | 2.3643 | 2.9516 | 4.3091 | 3.3533 | 4.3091 | 4.7718 | H11 | 2.0823 | 3.2240 | 3.3266 | 1.0968 | 2.6195 | 3.8400 | 3.2466 | 2.2096 | 3.6026 | 4.1449 | | 1.7670 | 3.6026 | 2.3780 | 4.7718 | 4.3091 | 3.3533 | 4.3091 | 2.9516 | 2.3643 | H12 | 2.0823 | 3.2240 | 2.6195 | 1.0968 | 3.3266 | 3.2466 | 3.8400 | 2.2096 | 2.3780 | 3.6026 | 1.7670 | | 4.1449 | 3.6026 | 4.3091 | 3.3533 | 4.3091 | 4.7718 | 2.3643 | 2.9516 | H13 | 2.0823 | 3.2240 | 2.6195 | 3.3266 | 1.0968 | 3.2466 | 2.2096 | 3.8400 | 3.6026 | 2.3780 | 3.6026 | 4.1449 | | 1.7670 | 3.3533 | 4.3091 | 2.9516 | 2.3643 | 4.7718 | 4.3091 | H14 | 2.0823 | 3.2240 | 3.3266 | 2.6195 | 1.0968 | 3.8400 | 2.2096 | 3.2466 | 4.1449 | 3.6026 | 2.3780 | 3.6026 | 1.7670 | | 4.3091 | 4.7718 | 2.3643 | 2.9516 | 4.3091 | 3.3533 | H15 | 3.2240 | 2.0823 | 2.2096 | 3.8400 | 3.2466 | 1.0968 | 2.6195 | 3.3266 | 2.9516 | 2.3643 | 4.7718 | 4.3091 | 3.3533 | 4.3091 | | 1.7670 | 3.6026 | 2.3780 | 3.6026 | 4.1449 | H16 | 3.2240 | 2.0823 | 2.2096 | 3.2466 | 3.8400 | 1.0968 | 3.3266 | 2.6195 | 2.3643 | 2.9516 | 4.3091 | 3.3533 | 4.3091 | 4.7718 | 1.7670 | | 4.1449 | 3.6026 | 2.3780 | 3.6026 | H17 | 3.2240 | 2.0823 | 3.8400 | 3.2466 | 2.2096 | 3.3266 | 1.0968 | 2.6195 | 4.7718 | 4.3091 | 3.3533 | 4.3091 | 2.9516 | 2.3643 | 3.6026 | 4.1449 | | 1.7670 | 3.6026 | 2.3780 | H18 | 3.2240 | 2.0823 | 3.2466 | 3.8400 | 2.2096 | 2.6195 | 1.0968 | 3.3266 | 4.3091 | 3.3533 | 4.3091 | 4.7718 | 2.3643 | 2.9516 | 2.3780 | 3.6026 | 1.7670 | | 4.1449 | 3.6026 | H19 | 3.2240 | 2.0823 | 3.2466 | 2.2096 | 3.8400 | 2.6195 | 3.3266 | 1.0968 | 3.3533 | 4.3091 | 2.9516 | 2.3643 | 4.7718 | 4.3091 | 3.6026 | 2.3780 | 3.6026 | 4.1449 | | 1.7670 | H20 | 3.2240 | 2.0823 | 3.8400 | 2.2096 | 3.2466 | 3.3266 | 2.6195 | 1.0968 | 4.3091 | 4.7718 | 2.3643 | 2.9516 | 4.3091 | 3.3533 | 4.1449 | 3.6026 | 2.3780 | 3.6026 | 1.7670 | |
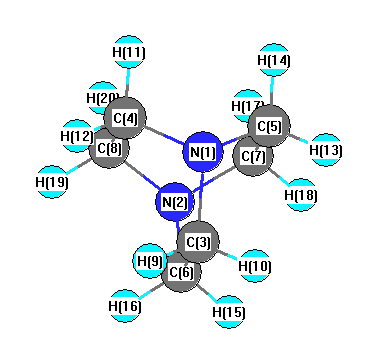 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
C3 |
C6 |
110.114 |
|
N1 |
C3 |
H9 |
108.013 |
| N1 |
C3 |
H10 |
108.013 |
|
N1 |
C4 |
C8 |
110.114 |
| N1 |
C4 |
H11 |
108.013 |
|
N1 |
C4 |
H12 |
108.013 |
| N1 |
C5 |
C7 |
110.114 |
|
N1 |
C5 |
H13 |
108.013 |
| N1 |
C5 |
H14 |
108.013 |
|
N2 |
C6 |
C3 |
110.114 |
| N2 |
C6 |
H15 |
108.013 |
|
N2 |
C6 |
H16 |
108.013 |
| N2 |
C7 |
C5 |
110.114 |
|
N2 |
C7 |
H17 |
108.013 |
| N2 |
C7 |
H18 |
108.013 |
|
N2 |
C8 |
C4 |
110.114 |
| N2 |
C8 |
H19 |
108.013 |
|
N2 |
C8 |
H20 |
108.013 |
| C3 |
N1 |
C4 |
108.821 |
|
C3 |
N1 |
C5 |
108.821 |
| C3 |
C6 |
H15 |
111.616 |
|
C3 |
C6 |
H16 |
111.616 |
| C4 |
N1 |
C5 |
108.821 |
|
C4 |
C8 |
H19 |
111.616 |
| C4 |
C8 |
H20 |
111.616 |
|
C5 |
C6 |
H15 |
101.597 |
| C5 |
C6 |
H16 |
151.063 |
|
C6 |
N2 |
C7 |
108.821 |
| C6 |
N2 |
C8 |
108.821 |
|
C6 |
C3 |
H9 |
111.616 |
| C6 |
C3 |
H10 |
111.616 |
|
C7 |
N2 |
C8 |
108.821 |
| C7 |
C5 |
H13 |
111.616 |
|
C7 |
C5 |
H14 |
111.616 |
| C8 |
C4 |
H11 |
111.616 |
|
C8 |
C4 |
H12 |
111.616 |
| H9 |
C3 |
H10 |
107.313 |
|
H11 |
C4 |
H12 |
107.313 |
| H13 |
C5 |
H14 |
107.313 |
|
H15 |
C6 |
H16 |
107.313 |
| H17 |
C7 |
H18 |
107.313 |
|
H19 |
C8 |
H20 |
107.313 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.377 |
|
|
|
| 2 |
N |
-0.377 |
|
|
|
| 3 |
C |
-0.214 |
|
|
|
| 4 |
C |
-0.214 |
|
|
|
| 5 |
C |
-0.214 |
|
|
|
| 6 |
C |
-0.214 |
|
|
|
| 7 |
C |
-0.214 |
|
|
|
| 8 |
C |
-0.214 |
|
|
|
| 9 |
H |
0.170 |
|
|
|
| 10 |
H |
0.170 |
|
|
|
| 11 |
H |
0.170 |
|
|
|
| 12 |
H |
0.170 |
|
|
|
| 13 |
H |
0.170 |
|
|
|
| 14 |
H |
0.170 |
|
|
|
| 15 |
H |
0.170 |
|
|
|
| 16 |
H |
0.170 |
|
|
|
| 17 |
H |
0.170 |
|
|
|
| 18 |
H |
0.170 |
|
|
|
| 19 |
H |
0.170 |
|
|
|
| 20 |
H |
0.170 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-46.385 |
0.000 |
0.000 |
| y |
0.000 |
-46.385 |
0.000 |
| z |
0.000 |
0.000 |
-57.214 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.414 |
0.000 |
0.000 |
| y |
0.000 |
5.414 |
0.000 |
| z |
0.000 |
0.000 |
-10.829 |
|
| Polar |
|---|
| 3z2-r2 | -21.658 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.854 |
0.000 |
0.000 |
| y |
0.000 |
10.855 |
0.000 |
| z |
0.000 |
0.000 |
9.317 |
<r2> (average value of r
2) Å
2
| <r2> |
212.631 |
| (<r2>)1/2 |
14.582 |
Jump to
S1C1
Energy calculated at HSEh1PBE/6-31G*
| | hartrees |
|---|
| Energy at 0K | -344.955195 |
| Energy at 298.15K | -344.972383 |
| HF Energy | -344.955195 |
| Nuclear repulsion energy | 426.015242 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3103 |
2951 |
0.00 |
|
|
|
| 2 |
A1 |
3076 |
2925 |
0.00 |
|
|
|
| 3 |
A1 |
1524 |
1449 |
0.00 |
|
|
|
| 4 |
A1 |
1385 |
1317 |
0.00 |
|
|
|
| 5 |
A1 |
1279 |
1217 |
0.00 |
|
|
|
| 6 |
A1 |
1035 |
985 |
0.00 |
|
|
|
| 7 |
A1 |
998 |
949 |
0.00 |
|
|
|
| 8 |
A1 |
839 |
798 |
0.00 |
|
|
|
| 9 |
A1 |
613 |
583 |
0.00 |
|
|
|
| 10 |
A1 |
83 |
78 |
0.00 |
|
|
|
| 11 |
A2 |
3123 |
2970 |
0.00 |
|
|
|
| 12 |
A2 |
3062 |
2912 |
115.54 |
|
|
|
| 13 |
A2 |
1515 |
1441 |
5.23 |
|
|
|
| 14 |
A2 |
1403 |
1334 |
5.95 |
|
|
|
| 15 |
A2 |
1212 |
1152 |
0.00 |
|
|
|
| 16 |
A2 |
1011 |
961 |
19.36 |
|
|
|
| 17 |
A2 |
816 |
776 |
0.01 |
|
|
|
| 18 |
A2 |
792 |
753 |
66.13 |
|
|
|
| 19 |
E |
3130 |
2976 |
87.02 |
|
|
|
| 19 |
E |
3130 |
2976 |
87.07 |
|
|
|
| 20 |
E |
3105 |
2953 |
0.01 |
|
|
|
| 20 |
E |
3105 |
2953 |
0.01 |
|
|
|
| 21 |
E |
3069 |
2918 |
108.18 |
|
|
|
| 21 |
E |
3069 |
2918 |
108.19 |
|
|
|
| 22 |
E |
3061 |
2911 |
0.00 |
|
|
|
| 22 |
E |
3061 |
2911 |
0.00 |
|
|
|
| 23 |
E |
1513 |
1439 |
10.21 |
|
|
|
| 23 |
E |
1513 |
1439 |
10.21 |
|
|
|
| 24 |
E |
1501 |
1427 |
0.00 |
|
|
|
| 24 |
E |
1501 |
1427 |
0.00 |
|
|
|
| 25 |
E |
1378 |
1310 |
10.71 |
|
|
|
| 25 |
E |
1378 |
1310 |
10.71 |
|
|
|
| 26 |
E |
1370 |
1303 |
0.00 |
|
|
|
| 26 |
E |
1370 |
1303 |
0.00 |
|
|
|
| 27 |
E |
1350 |
1283 |
0.62 |
|
|
|
| 27 |
E |
1350 |
1283 |
0.62 |
|
|
|
| 28 |
E |
1343 |
1277 |
0.00 |
|
|
|
| 28 |
E |
1343 |
1277 |
0.00 |
|
|
|
| 29 |
E |
1223 |
1163 |
0.00 |
|
|
|
| 29 |
E |
1223 |
1163 |
0.00 |
|
|
|
| 30 |
E |
1121 |
1066 |
40.05 |
|
|
|
| 30 |
E |
1121 |
1066 |
40.07 |
|
|
|
| 31 |
E |
1071 |
1018 |
0.00 |
|
|
|
| 31 |
E |
1071 |
1018 |
0.00 |
|
|
|
| 32 |
E |
925 |
879 |
3.75 |
|
|
|
| 32 |
E |
925 |
879 |
3.75 |
|
|
|
| 33 |
E |
838 |
797 |
4.12 |
|
|
|
| 33 |
E |
838 |
797 |
4.12 |
|
|
|
| 34 |
E |
594 |
565 |
0.00 |
|
|
|
| 34 |
E |
594 |
565 |
0.00 |
|
|
|
| 35 |
E |
428 |
407 |
0.00 |
|
|
|
| 35 |
E |
428 |
407 |
0.00 |
|
|
|
| 36 |
E |
341 |
324 |
0.00 |
|
|
|
| 36 |
E |
341 |
324 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 40790.7 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 38792.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G*
Point Group is D3
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
0.000 |
0.000 |
1.282 |
| N2 |
0.000 |
0.000 |
-1.282 |
| C3 |
-0.000 |
1.374 |
0.778 |
| C4 |
1.190 |
-0.687 |
0.778 |
| C5 |
-1.190 |
-0.687 |
0.778 |
| C6 |
0.000 |
1.374 |
-0.778 |
| C7 |
-1.190 |
-0.687 |
-0.778 |
| C8 |
1.190 |
-0.687 |
-0.778 |
| H9 |
0.882 |
1.884 |
1.182 |
| H10 |
-0.885 |
1.883 |
1.181 |
| H11 |
1.190 |
-1.706 |
1.182 |
| H12 |
2.073 |
-0.175 |
1.181 |
| H13 |
-2.073 |
-0.178 |
1.182 |
| H14 |
-1.188 |
-1.707 |
1.181 |
| H15 |
-0.882 |
1.884 |
-1.182 |
| H16 |
0.885 |
1.883 |
-1.181 |
| H17 |
-1.190 |
-1.706 |
-1.182 |
| H18 |
-2.073 |
-0.175 |
-1.181 |
| H19 |
2.073 |
-0.178 |
-1.182 |
| H20 |
1.188 |
-1.707 |
-1.181 |
Atom - Atom Distances (Å)
| |
N1 |
N2 |
C3 |
C4 |
C5 |
C6 |
C7 |
C8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
H19 |
H20 |
| N1 | | 2.5632 | 1.4631 | 1.4631 | 1.4631 | 2.4758 | 2.4758 | 2.4758 | 2.0827 | 2.0824 | 2.0827 | 2.0824 | 2.0827 | 2.0824 | 3.2248 | 3.2238 | 3.2248 | 3.2238 | 3.2248 | 3.2238 |
N2 | 2.5632 | | 2.4758 | 2.4758 | 2.4758 | 1.4631 | 1.4631 | 1.4631 | 3.2248 | 3.2238 | 3.2248 | 3.2238 | 3.2248 | 3.2238 | 2.0827 | 2.0824 | 2.0827 | 2.0824 | 2.0827 | 2.0824 | C3 | 1.4631 | 2.4758 | | 2.3794 | 2.3794 | 1.5563 | 2.8428 | 2.8436 | 1.0968 | 1.0968 | 3.3267 | 2.6190 | 2.6202 | 3.3265 | 2.2095 | 2.2095 | 3.8400 | 3.2455 | 3.2480 | 3.8399 | C4 | 1.4631 | 2.4758 | 2.3794 | | 2.3794 | 2.8428 | 2.8435 | 1.5563 | 2.6202 | 3.3265 | 1.0968 | 1.0968 | 3.3267 | 2.6190 | 3.8400 | 3.2455 | 3.2480 | 3.8399 | 2.2095 | 2.2095 | C5 | 1.4631 | 2.4758 | 2.3794 | 2.3794 | | 2.8436 | 1.5563 | 2.8428 | 3.3267 | 2.6190 | 2.6202 | 3.3265 | 1.0968 | 1.0968 | 3.2480 | 3.8399 | 2.2095 | 2.2095 | 3.8400 | 3.2455 | C6 | 2.4758 | 1.4631 | 1.5563 | 2.8428 | 2.8436 | | 2.3794 | 2.3794 | 2.2095 | 2.2095 | 3.8400 | 3.2455 | 3.2480 | 3.8399 | 1.0968 | 1.0968 | 3.3267 | 2.6190 | 2.6202 | 3.3265 | C7 | 2.4758 | 1.4631 | 2.8428 | 2.8435 | 1.5563 | 2.3794 | | 2.3794 | 3.8400 | 3.2455 | 3.2480 | 3.8399 | 2.2095 | 2.2095 | 2.6202 | 3.3265 | 1.0968 | 1.0968 | 3.3267 | 2.6190 | C8 | 2.4758 | 1.4631 | 2.8436 | 1.5563 | 2.8428 | 2.3794 | 2.3794 | | 3.2480 | 3.8399 | 2.2095 | 2.2095 | 3.8400 | 3.2455 | 3.3267 | 2.6190 | 2.6202 | 3.3265 | 1.0968 | 1.0968 | H9 | 2.0827 | 3.2248 | 1.0968 | 2.6202 | 3.3267 | 2.2095 | 3.8400 | 3.2480 | | 1.7668 | 3.6032 | 2.3784 | 3.6032 | 4.1453 | 2.9508 | 2.3639 | 4.7726 | 4.3081 | 3.3558 | 4.3103 | H10 | 2.0824 | 3.2238 | 1.0968 | 3.3265 | 2.6190 | 2.2095 | 3.2455 | 3.8399 | 1.7668 | | 4.1453 | 3.6026 | 2.3784 | 3.6026 | 2.3639 | 2.9517 | 4.3081 | 3.3509 | 4.3103 | 4.7713 | H11 | 2.0827 | 3.2248 | 3.3267 | 1.0968 | 2.6202 | 3.8400 | 3.2480 | 2.2095 | 3.6032 | 4.1453 | | 1.7668 | 3.6032 | 2.3784 | 4.7726 | 4.3081 | 3.3558 | 4.3103 | 2.9508 | 2.3639 | H12 | 2.0824 | 3.2238 | 2.6190 | 1.0968 | 3.3265 | 3.2455 | 3.8399 | 2.2095 | 2.3784 | 3.6026 | 1.7668 | | 4.1453 | 3.6026 | 4.3081 | 3.3509 | 4.3103 | 4.7713 | 2.3639 | 2.9517 | H13 | 2.0827 | 3.2248 | 2.6202 | 3.3267 | 1.0968 | 3.2480 | 2.2095 | 3.8400 | 3.6032 | 2.3784 | 3.6032 | 4.1453 | | 1.7668 | 3.3558 | 4.3103 | 2.9508 | 2.3639 | 4.7726 | 4.3081 | H14 | 2.0824 | 3.2238 | 3.3265 | 2.6190 | 1.0968 | 3.8399 | 2.2095 | 3.2455 | 4.1453 | 3.6026 | 2.3784 | 3.6026 | 1.7668 | | 4.3103 | 4.7713 | 2.3639 | 2.9517 | 4.3081 | 3.3509 | H15 | 3.2248 | 2.0827 | 2.2095 | 3.8400 | 3.2480 | 1.0968 | 2.6202 | 3.3267 | 2.9508 | 2.3639 | 4.7726 | 4.3081 | 3.3558 | 4.3103 | | 1.7668 | 3.6032 | 2.3784 | 3.6032 | 4.1453 | H16 | 3.2238 | 2.0824 | 2.2095 | 3.2455 | 3.8399 | 1.0968 | 3.3265 | 2.6190 | 2.3639 | 2.9517 | 4.3081 | 3.3509 | 4.3103 | 4.7713 | 1.7668 | | 4.1453 | 3.6026 | 2.3784 | 3.6026 | H17 | 3.2248 | 2.0827 | 3.8400 | 3.2480 | 2.2095 | 3.3267 | 1.0968 | 2.6202 | 4.7726 | 4.3081 | 3.3558 | 4.3103 | 2.9508 | 2.3639 | 3.6032 | 4.1453 | | 1.7668 | 3.6032 | 2.3784 | H18 | 3.2238 | 2.0824 | 3.2455 | 3.8399 | 2.2095 | 2.6190 | 1.0968 | 3.3265 | 4.3081 | 3.3509 | 4.3103 | 4.7713 | 2.3639 | 2.9517 | 2.3784 | 3.6026 | 1.7668 | | 4.1453 | 3.6026 | H19 | 3.2248 | 2.0827 | 3.2480 | 2.2095 | 3.8400 | 2.6202 | 3.3267 | 1.0968 | 3.3558 | 4.3103 | 2.9508 | 2.3639 | 4.7726 | 4.3081 | 3.6032 | 2.3784 | 3.6032 | 4.1453 | | 1.7668 | H20 | 3.2238 | 2.0824 | 3.8399 | 2.2095 | 3.2455 | 3.3265 | 2.6190 | 1.0968 | 4.3103 | 4.7713 | 2.3639 | 2.9517 | 4.3081 | 3.3509 | 4.1453 | 3.6026 | 2.3784 | 3.6026 | 1.7668 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
C3 |
C6 |
110.127 |
|
N1 |
C3 |
H9 |
108.038 |
| N1 |
C3 |
H10 |
108.011 |
|
N1 |
C4 |
C8 |
110.127 |
| N1 |
C4 |
H11 |
108.038 |
|
N1 |
C4 |
H12 |
108.011 |
| N1 |
C5 |
C7 |
110.127 |
|
N1 |
C5 |
H13 |
108.038 |
| N1 |
C5 |
H14 |
108.011 |
|
N2 |
C6 |
C3 |
110.127 |
| N2 |
C6 |
H15 |
108.038 |
|
N2 |
C6 |
H16 |
108.011 |
| N2 |
C7 |
C5 |
110.127 |
|
N2 |
C7 |
H17 |
108.038 |
| N2 |
C7 |
H18 |
108.011 |
|
N2 |
C8 |
C4 |
110.127 |
| N2 |
C8 |
H19 |
108.038 |
|
N2 |
C8 |
H20 |
108.011 |
| C3 |
N1 |
C4 |
108.808 |
|
C3 |
N1 |
C5 |
108.808 |
| C3 |
C6 |
H15 |
111.606 |
|
C3 |
C6 |
H16 |
111.600 |
| C4 |
N1 |
C5 |
108.808 |
|
C4 |
C8 |
H19 |
111.606 |
| C4 |
C8 |
H20 |
111.600 |
|
C5 |
C6 |
H15 |
101.661 |
| C5 |
C6 |
H16 |
151.010 |
|
C6 |
N2 |
C7 |
108.808 |
| C6 |
N2 |
C8 |
108.808 |
|
C6 |
C3 |
H9 |
111.606 |
| C6 |
C3 |
H10 |
111.600 |
|
C7 |
N2 |
C8 |
108.808 |
| C7 |
C5 |
H13 |
111.606 |
|
C7 |
C5 |
H14 |
111.600 |
| C8 |
C4 |
H11 |
111.606 |
|
C8 |
C4 |
H12 |
111.600 |
| H9 |
C3 |
H10 |
107.302 |
|
H11 |
C4 |
H12 |
107.302 |
| H13 |
C5 |
H14 |
107.302 |
|
H15 |
C6 |
H16 |
107.302 |
| H17 |
C7 |
H18 |
107.302 |
|
H19 |
C8 |
H20 |
107.302 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.377 |
|
|
|
| 2 |
N |
-0.377 |
|
|
|
| 3 |
C |
-0.214 |
|
|
|
| 4 |
C |
-0.214 |
|
|
|
| 5 |
C |
-0.214 |
|
|
|
| 6 |
C |
-0.214 |
|
|
|
| 7 |
C |
-0.214 |
|
|
|
| 8 |
C |
-0.214 |
|
|
|
| 9 |
H |
0.170 |
|
|
|
| 10 |
H |
0.170 |
|
|
|
| 11 |
H |
0.170 |
|
|
|
| 12 |
H |
0.170 |
|
|
|
| 13 |
H |
0.170 |
|
|
|
| 14 |
H |
0.170 |
|
|
|
| 15 |
H |
0.170 |
|
|
|
| 16 |
H |
0.170 |
|
|
|
| 17 |
H |
0.170 |
|
|
|
| 18 |
H |
0.170 |
|
|
|
| 19 |
H |
0.170 |
|
|
|
| 20 |
H |
0.170 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-46.383 |
0.000 |
0.000 |
| y |
0.000 |
-46.383 |
0.000 |
| z |
0.000 |
0.000 |
-57.220 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.419 |
0.000 |
0.000 |
| y |
0.000 |
5.419 |
0.000 |
| z |
0.000 |
0.000 |
-10.837 |
|
| Polar |
|---|
| 3z2-r2 | -21.674 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.854 |
0.000 |
0.000 |
| y |
0.000 |
10.855 |
0.000 |
| z |
0.000 |
0.000 |
9.317 |
<r2> (average value of r
2) Å
2
| <r2> |
212.648 |
| (<r2>)1/2 |
14.582 |
