return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

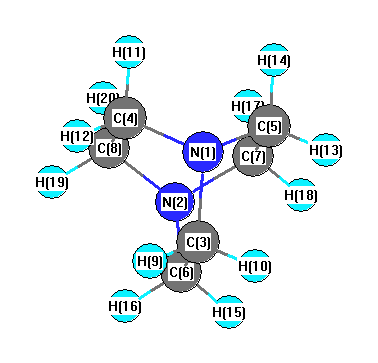
All results from a given calculation for C6H12N2 (Triethylenediamine)

using model chemistry: MP3/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes D3H 1A1'
Energy calculated at MP3/6-31G*
 hartrees
Energy at 0K-344.212573
Energy at 298.15K 
Nuclear repulsion energy424.585627
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP3/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1' 3112 2921 0.00      
2 A1' 1572 1476 0.00      
3 A1' 1415 1328 0.00      
4 A1' 1022 959 0.00      
5 A1' 842 790 0.00      
6 A1' 623 585 0.00      
7 A1" 3136 2944 0.00      
8 A1" 1301 1221 0.00      
9 A1" 1046 982 0.00      
10 A1" 27i 25i 0.00      
11 A2' 3155 2962 0.00      
12 A2' 1225 1149 0.00      
13 A2' 832 780 0.00      
14 A2" 3100 2909 93.68      
15 A2" 1558 1463 4.40      
16 A2" 1425 1337 4.53      
17 A2" 1021 958 21.88      
18 A2" 797 748 66.31      
19 E' 3162 2968 75.88      
20 E' 3105 2914 85.79      
21 E' 1558 1462 9.49      
22 E' 1396 1310 9.67      
23 E' 1374 1290 1.86      
24 E' 1141 1071 35.85      
25 E' 942 884 5.69      
25 E' 942 884 5.69      
26 E' 858 805 4.14      
26 E' 858 805 4.14      
27 E' 440 413 0.05      
27 E' 440 413 0.05      
28 E" 3139 2947 0.00      
29 E" 3096 2906 0.00      
30 E" 1544 1449 0.00      
31 E" 1390 1304 0.00      
32 E" 1361 1277 0.00      
33 E" 1246 1170 0.00      
34 E" 1081 1015 0.00      
35 E" 599 562 0.00      
35 E" 599 562 0.00      
36 E" 334 314 0.00      
36 E" 334 314 0.00      

Unscaled Zero Point Vibrational Energy (zpe) 41341.4 cm-1
Scaled (by 0.9386) Zero Point Vibrational Energy (zpe) 38803.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP3/6-31G*
ABC
0.08890 0.08344 0.08344

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP3/6-31G*

Point Group is D3h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 0.000 0.000 1.291
N2 0.000 0.000 -1.291
C3 0.000 1.380 0.779
C4 1.195 -0.690 0.779
C5 -1.195 -0.690 0.779
C6 0.000 1.380 -0.779
C7 -1.195 -0.690 -0.779
C8 1.195 -0.690 -0.779
H9 0.883 1.889 1.182
H10 -0.883 1.889 1.182
H11 1.194 -1.709 1.182
H12 2.077 -0.180 1.182
H13 -2.077 -0.180 1.182
H14 -1.194 -1.709 1.182
H15 -0.883 1.889 -1.182
H16 0.883 1.889 -1.182
H17 -1.194 -1.709 -1.182
H18 -2.077 -0.180 -1.182
H19 2.077 -0.180 -1.182
H20 1.194 -1.709 -1.182

Atom - Atom Distances (Å)
  N1 N2 C3 C4 C5 C6 C7 C8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18 H19 H20
N12.58141.47171.47171.47172.48722.48722.48722.08752.08752.08752.08752.08752.08753.23453.23453.23453.23453.23453.2345
N22.58142.48722.48722.48721.47171.47171.47173.23453.23453.23453.23453.23453.23452.08752.08752.08752.08752.08752.0875
C31.47172.48722.38992.38991.55742.85262.85261.09581.09583.33582.62862.62863.33582.20992.20993.84853.25463.25463.8485
C41.47172.48722.38992.38992.85262.85261.55742.62863.33581.09581.09583.33582.62863.84853.25463.25463.84852.20992.2099
C51.47172.48722.38992.38992.85261.55742.85263.33582.62862.62863.33581.09581.09583.25463.84852.20992.20993.84853.2546
C62.48721.47171.55742.85262.85262.38992.38992.20992.20993.84853.25463.25463.84851.09581.09583.33582.62862.62863.3358
C72.48721.47172.85262.85261.55742.38992.38993.84853.25463.25463.84852.20992.20992.62863.33581.09581.09583.33582.6286
C82.48721.47172.85261.55742.85262.38992.38993.25463.84852.20992.20993.84853.25463.33582.62862.62863.33581.09581.0958
H92.08753.23451.09582.62863.33582.20993.84853.25461.76513.61072.38863.61074.15372.95092.36484.77974.31623.36124.3162
H102.08753.23451.09583.33582.62862.20993.25463.84851.76514.15373.61072.38863.61072.36482.95094.31623.36124.31624.7797
H112.08753.23453.33581.09582.62863.84853.25462.20993.61074.15371.76513.61072.38864.77974.31623.36124.31622.95092.3648
H122.08753.23452.62861.09583.33583.25463.84852.20992.38863.61071.76514.15373.61074.31623.36124.31624.77972.36482.9509
H132.08753.23452.62863.33581.09583.25462.20993.84853.61072.38863.61074.15371.76513.36124.31622.95092.36484.77974.3162
H142.08753.23453.33582.62861.09583.84852.20993.25464.15373.61072.38863.61071.76514.31624.77972.36482.95094.31623.3612
H153.23452.08752.20993.84853.25461.09582.62863.33582.95092.36484.77974.31623.36124.31621.76513.61072.38863.61074.1537
H163.23452.08752.20993.25463.84851.09583.33582.62862.36482.95094.31623.36124.31624.77971.76514.15373.61072.38863.6107
H173.23452.08753.84853.25462.20993.33581.09582.62864.77974.31623.36124.31622.95092.36483.61074.15371.76513.61072.3886
H183.23452.08753.25463.84852.20992.62861.09583.33584.31623.36124.31624.77972.36482.95092.38863.61071.76514.15373.6107
H193.23452.08753.25462.20993.84852.62863.33581.09583.36124.31622.95092.36484.77974.31623.61072.38863.61074.15371.7651
H203.23452.08753.84852.20993.25463.33582.62861.09584.31624.77972.36482.95094.31623.36124.15373.61072.38863.61071.7651

picture of Triethylenediamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C3 C6 110.357 N1 C3 H9 107.889
N1 C3 H10 107.889 N1 C4 C8 110.357
N1 C4 H11 107.889 N1 C4 H12 107.889
N1 C5 C7 110.357 N1 C5 H13 107.889
N1 C5 H14 107.889 N2 C6 C3 110.357
N2 C6 H15 107.889 N2 C6 H16 107.889
N2 C7 C5 110.357 N2 C7 H17 107.889
N2 C7 H18 107.889 N2 C8 C4 110.357
N2 C8 H19 107.889 N2 C8 H20 107.889
C3 N1 C4 108.571 C3 N1 C5 108.571
C3 C6 H15 111.618 C3 C6 H16 111.618
C4 N1 C5 108.571 C4 C8 H19 111.618
C4 C8 H20 111.618 C5 C6 H15 101.576
C5 C6 H16 151.096 C6 N2 C7 108.571
C6 N2 C8 108.571 C6 C3 H9 111.618
C6 C3 H10 111.618 C7 N2 C8 108.571
C7 C5 H13 111.618 C7 C5 H14 111.618
C8 C4 H11 111.618 C8 C4 H12 111.618
H9 C3 H10 107.297 H11 C4 H12 107.297
H13 C5 H14 107.297 H15 C6 H16 107.297
H17 C7 H18 107.297 H19 C8 H20 107.297
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability