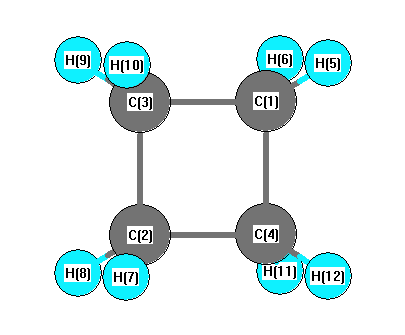
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3037 |
2987 |
0.00 |
|
|
|
| 2 |
A1 |
3004 |
2954 |
0.00 |
|
|
|
| 3 |
A1 |
1500 |
1475 |
0.00 |
|
|
|
| 4 |
A1 |
1149 |
1130 |
0.00 |
|
|
|
| 5 |
A1 |
1015 |
998 |
0.00 |
|
|
|
| 6 |
A1 |
199 |
196 |
0.00 |
|
|
|
| 7 |
A2 |
1219 |
1199 |
0.00 |
|
|
|
| 8 |
A2 |
942 |
927 |
0.00 |
|
|
|
| 9 |
B1 |
1234 |
1214 |
0.00 |
|
|
|
| 10 |
B1 |
1140 |
1121 |
0.00 |
|
|
|
| 11 |
B1 |
941 |
925 |
0.00 |
|
|
|
| 12 |
B2 |
3067 |
3016 |
95.93 |
|
|
|
| 13 |
B2 |
2997 |
2948 |
9.64 |
|
|
|
| 14 |
B2 |
1469 |
1445 |
4.00 |
|
|
|
| 15 |
B2 |
873 |
858 |
0.02 |
|
|
|
| 16 |
B2 |
614 |
604 |
3.75 |
|
|
|
| 17 |
E |
3051 |
3000 |
20.31 |
|
|
|
| 17 |
E |
3051 |
3000 |
20.31 |
|
|
|
| 18 |
E |
2997 |
2948 |
78.20 |
|
|
|
| 18 |
E |
2997 |
2948 |
78.20 |
|
|
|
| 19 |
E |
1460 |
1436 |
0.27 |
|
|
|
| 19 |
E |
1460 |
1436 |
0.27 |
|
|
|
| 20 |
E |
1261 |
1240 |
0.89 |
|
|
|
| 20 |
E |
1261 |
1240 |
0.89 |
|
|
|
| 21 |
E |
1226 |
1205 |
0.51 |
|
|
|
| 21 |
E |
1226 |
1205 |
0.51 |
|
|
|
| 22 |
E |
911 |
896 |
2.45 |
|
|
|
| 22 |
E |
911 |
896 |
2.45 |
|
|
|
| 23 |
E |
741 |
729 |
0.70 |
|
|
|
| 23 |
E |
741 |
729 |
0.70 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23845.6 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 23452.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.298 |
|
|
|
| 2 |
C |
-0.298 |
|
|
|
| 3 |
C |
-0.298 |
|
|
|
| 4 |
C |
-0.298 |
|
|
|
| 5 |
H |
0.151 |
|
|
|
| 6 |
H |
0.147 |
|
|
|
| 7 |
H |
0.151 |
|
|
|
| 8 |
H |
0.147 |
|
|
|
| 9 |
H |
0.151 |
|
|
|
| 10 |
H |
0.147 |
|
|
|
| 11 |
H |
0.151 |
|
|
|
| 12 |
H |
0.147 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-26.854 |
0.000 |
0.000 |
| y |
0.000 |
-26.854 |
0.000 |
| z |
0.000 |
0.000 |
-25.527 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.663 |
0.000 |
0.000 |
| y |
0.000 |
-0.663 |
0.000 |
| z |
0.000 |
0.000 |
1.327 |
|
| Polar |
|---|
| 3z2-r2 | 2.653 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.283 |
0.000 |
0.000 |
| y |
0.000 |
6.283 |
0.000 |
| z |
0.000 |
0.000 |
5.877 |
<r2> (average value of r
2) Å
2
| <r2> |
75.345 |
| (<r2>)1/2 |
8.680 |