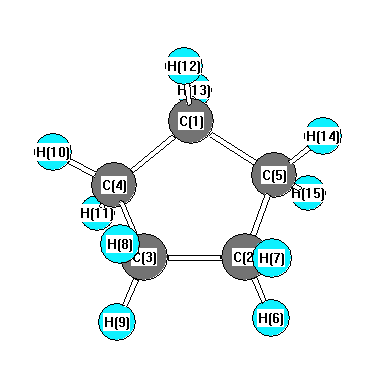
Vibrational Frequencies calculated at B2PLYP/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3149 |
2989 |
105.99 |
|
|
|
| 2 |
A |
3135 |
2975 |
21.04 |
|
|
|
| 3 |
A |
3134 |
2975 |
26.91 |
|
|
|
| 4 |
A |
3128 |
2970 |
11.64 |
|
|
|
| 5 |
A |
3119 |
2961 |
29.74 |
|
|
|
| 6 |
A |
3097 |
2940 |
47.13 |
|
|
|
| 7 |
A |
3082 |
2926 |
48.71 |
|
|
|
| 8 |
A |
3081 |
2924 |
3.49 |
|
|
|
| 9 |
A |
3075 |
2918 |
36.92 |
|
|
|
| 10 |
A |
3071 |
2915 |
21.21 |
|
|
|
| 11 |
A |
1574 |
1494 |
0.06 |
|
|
|
| 12 |
A |
1547 |
1469 |
2.78 |
|
|
|
| 13 |
A |
1547 |
1469 |
1.14 |
|
|
|
| 14 |
A |
1541 |
1462 |
3.50 |
|
|
|
| 15 |
A |
1535 |
1457 |
0.10 |
|
|
|
| 16 |
A |
1388 |
1317 |
0.37 |
|
|
|
| 17 |
A |
1385 |
1315 |
0.46 |
|
|
|
| 18 |
A |
1379 |
1309 |
0.06 |
|
|
|
| 19 |
A |
1358 |
1289 |
0.54 |
|
|
|
| 20 |
A |
1334 |
1267 |
0.00 |
|
|
|
| 21 |
A |
1311 |
1245 |
0.96 |
|
|
|
| 22 |
A |
1268 |
1203 |
0.19 |
|
|
|
| 23 |
A |
1258 |
1194 |
0.62 |
|
|
|
| 24 |
A |
1234 |
1171 |
0.35 |
|
|
|
| 25 |
A |
1209 |
1148 |
0.01 |
|
|
|
| 26 |
A |
1077 |
1022 |
0.02 |
|
|
|
| 27 |
A |
1062 |
1008 |
0.00 |
|
|
|
| 28 |
A |
1024 |
972 |
0.46 |
|
|
|
| 29 |
A |
989 |
939 |
1.11 |
|
|
|
| 30 |
A |
930 |
883 |
1.49 |
|
|
|
| 31 |
A |
924 |
877 |
1.43 |
|
|
|
| 32 |
A |
903 |
858 |
0.38 |
|
|
|
| 33 |
A |
890 |
844 |
0.49 |
|
|
|
| 34 |
A |
841 |
798 |
0.23 |
|
|
|
| 35 |
A |
786 |
746 |
0.85 |
|
|
|
| 36 |
A |
623 |
591 |
0.16 |
|
|
|
| 37 |
A |
557 |
528 |
1.05 |
|
|
|
| 38 |
A |
273 |
259 |
0.00 |
|
|
|
| 39 |
A |
27 |
26 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31422.1 cm
-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 29825.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.288 |
|
|
|
| 2 |
C |
-0.293 |
|
|
|
| 3 |
C |
-0.294 |
|
|
|
| 4 |
C |
-0.293 |
|
|
|
| 5 |
C |
-0.288 |
|
|
|
| 6 |
H |
0.145 |
|
|
|
| 7 |
H |
0.147 |
|
|
|
| 8 |
H |
0.148 |
|
|
|
| 9 |
H |
0.148 |
|
|
|
| 10 |
H |
0.146 |
|
|
|
| 11 |
H |
0.147 |
|
|
|
| 12 |
H |
0.146 |
|
|
|
| 13 |
H |
0.142 |
|
|
|
| 14 |
H |
0.142 |
|
|
|
| 15 |
H |
0.146 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.003 |
-0.000 |
0.000 |
0.003 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.662 |
-0.002 |
-0.000 |
| y |
-0.002 |
-33.644 |
0.017 |
| z |
-0.000 |
0.017 |
-32.543 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.568 |
-0.002 |
-0.000 |
| y |
-0.002 |
-0.542 |
0.017 |
| z |
-0.000 |
0.017 |
1.110 |
|
| Polar |
|---|
| 3z2-r2 | 2.220 |
|---|
| x2-y2 | -0.018 |
|---|
| xy | -0.002 |
|---|
| xz | -0.000 |
|---|
| yz | 0.017 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.576 |
-0.000 |
0.000 |
| y |
-0.000 |
7.585 |
-0.008 |
| z |
0.000 |
-0.008 |
6.948 |
<r2> (average value of r
2) Å
2
| <r2> |
115.672 |
| (<r2>)1/2 |
10.755 |