return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H4N2 (1H-Imidazole)

using model chemistry: QCISD/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at QCISD/6-31G*
 hartrees
Energy at 0K-225.542611
Energy at 298.15K-225.548522
HF Energy-224.811758
Nuclear repulsion energy162.842662
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3684 3508 63.29      
2 A' 3309 3152 2.36      
3 A' 3283 3127 1.75      
4 A' 3280 3124 6.54      
5 A' 1607 1531 13.26      
6 A' 1555 1481 19.54      
7 A' 1489 1418 24.93      
8 A' 1407 1340 9.49      
9 A' 1311 1249 0.56      
10 A' 1182 1126 3.76      
11 A' 1175 1119 5.23      
12 A' 1128 1074 13.21      
13 A' 1101 1048 38.89      
14 A' 951 906 1.85      
15 A' 914 871 7.14      
16 A" 839 799 5.17      
17 A" 820 781 35.07      
18 A" 717 682 28.75      
19 A" 680 648 7.05      
20 A" 636 606 18.25      
21 A" 477 454 110.01      

Unscaled Zero Point Vibrational Energy (zpe) 15772.5 cm-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 15020.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G*
ABC
0.32428 0.31187 0.15898

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
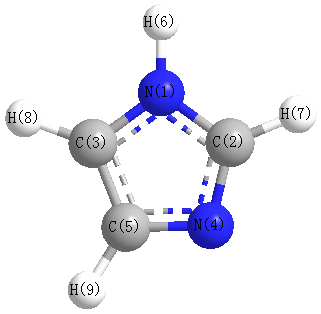
N1 0.000 1.107 0.000
C2 -1.089 0.282 0.000
C3 1.122 0.299 0.000
N4 -0.748 -0.988 0.000
C5 0.638 -0.983 0.000
H6 -0.011 2.118 0.000
H7 -2.104 0.663 0.000
H8 2.125 0.704 0.000
H9 1.200 -1.908 0.000

Atom - Atom Distances (Å)
  N1 C2 C3 N4 C5 H6 H7 H8 H9
N11.36641.38272.22452.18501.01062.15052.16253.2456
C21.36642.21121.31542.14102.12891.08383.24143.1689
C31.38272.21122.26941.36972.14273.24641.08152.2085
N42.22451.31542.26941.38573.19192.13673.33342.1545
C52.18502.14101.36971.38573.16753.19802.24811.0831
H61.01062.12892.14273.19193.16752.54932.56094.2043
H72.15051.08383.24642.13673.19802.54934.22884.1870
H82.16253.24141.08153.33342.24812.56094.22882.7709
H93.24563.16892.20852.15451.08314.20434.18702.7709

picture of 1H-Imidazole state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C2 N4 112.075 N1 C2 H7 122.311
N1 C3 C5 105.094 N1 C3 H8 122.222
C2 N1 C3 107.092 C2 N1 H6 126.526
C2 N4 C5 104.837 C3 N1 H6 126.383
C3 C5 N4 110.901 C3 C5 H9 128.039
N4 C2 H7 125.614 N4 C5 H9 121.059
C5 C3 H8 132.684
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability