return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H4N2 (Pyridazine)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-264.010798
Energy at 298.15K 
HF Energy-263.734028
Nuclear repulsion energy207.446648
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3250 3085 6.66      
2 A1 3229 3065 1.83      
3 A1 1631 1548 6.36      
4 A1 1497 1421 1.57      
5 A1 1225 1163 0.06      
6 A1 1191 1130 0.03      
7 A1 1104 1048 9.73      
8 A1 1008 957 8.13      
9 A1 682 647 2.80      
10 A2 994 943 0.00      
11 A2 945 897 0.00      
12 A2 751 712 0.00      
13 A2 372 353 0.00      
14 B1 971 922 0.00      
15 B1 766 727 34.32      
16 B1 376 357 9.07      
17 B2 3239 3074 25.11      
18 B2 3223 3059 4.81      
19 B2 1626 1544 4.64      
20 B2 1458 1384 17.07      
21 B2 1327 1260 2.36      
22 B2 1098 1042 1.79      
23 B2 1061 1007 1.36      
24 B2 632 600 0.01      

Unscaled Zero Point Vibrational Energy (zpe) 16827.4 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 15972.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.692 1.181
C2 0.000 -0.692 1.181
C3 0.000 -1.324 -0.066
C4 0.000 1.324 -0.066
H5 0.000 1.271 2.098
H6 0.000 -1.271 2.098
H7 0.000 -2.406 -0.151
H8 0.000 2.406 -0.151
N9 0.000 0.670 -1.234
N10 0.000 -0.670 -1.234

Atom - Atom Distances (Å)
  C1 C2 C3 C4 H5 H6 H7 H8 N9 N10
C11.38402.37041.39821.08452.16673.37192.17042.41572.7733
C21.38401.39822.37042.16671.08452.17043.37192.77332.4157
C32.37041.39822.64713.37892.16511.08553.73022.31091.3383
C41.39822.37042.64712.16513.37893.73021.08551.33832.3109
H51.08452.16673.37892.16512.54204.31002.51903.38633.8569
H62.16671.08452.16513.37892.54202.51904.31003.85693.3863
H73.37192.17041.08553.73024.31002.51904.81143.26142.0459
H82.17043.37193.73021.08552.51904.31004.81142.04593.2614
N92.41572.77332.31091.33833.38633.85693.26142.04591.3408
N102.77332.41571.33832.31093.85693.38632.04593.26141.3408

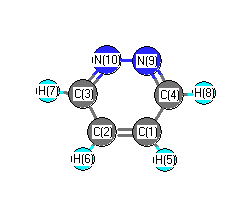
picture of Pyridazine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 116.851 C1 C2 H6 122.267
C1 C4 H8 121.313 C1 C4 N9 123.938
C2 C1 C4 116.851 C2 C1 H5 122.267
C2 C3 H7 121.313 C2 C3 N10 123.938
C3 C2 H6 120.882 C3 N10 N9 119.211
C4 C1 H5 120.882 C4 N9 N10 119.211
H7 C3 N10 114.749 H8 C4 N9 114.749
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.152      
2 C -0.152      
3 C 0.035      
4 C 0.035      
5 H 0.189      
6 H 0.189      
7 H 0.187      
8 H 0.187      
9 N -0.259      
10 N -0.259      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 2.977 0.000 0.000
y 0.000 9.210 0.000
z 0.000 0.000 8.354


<r2> (average value of r2) Å2
<r2> 116.183
(<r2>)1/2 10.779