return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H4N2 (1,3-Diazine)

using model chemistry: MP3/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP3/6-31G*
 hartrees
Energy at 0K-263.524503
Energy at 298.15K-263.530410
HF Energy-262.691346
Nuclear repulsion energy208.793709
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP3/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3276 3075 5.42      
2 A1 3254 3055 18.55      
3 A1 3235 3036 8.32      
4 A1 1681 1578 30.21      
5 A1 1482 1391 57.56      
6 A1 1194 1121 2.57      
7 A1 1107 1039 0.95      
8 A1 1039 975 5.09      
9 A1 705 661 4.44      
10 A2 1001 940 0.00      
11 A2 416 390 0.00      
12 B1 1017 955 0.07      
13 B1 954 895 0.19      
14 B1 825 775 7.73      
15 B1 735 690 40.00      
16 B1 365 343 3.18      
17 B2 3238 3040 22.12      
18 B2 1688 1585 63.18      
19 B2 1547 1452 13.29      
20 B2 1434 1346 0.19      
21 B2 1289 1210 5.43      
22 B2 1177 1105 5.76      
23 B2 1130 1060 2.38      
24 B2 642 603 11.64      

Unscaled Zero Point Vibrational Energy (zpe) 17215.8 cm-1
Scaled (by 0.9386) Zero Point Vibrational Energy (zpe) 16158.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP3/6-31G*
ABC
0.20964 0.20260 0.10303

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP3/6-31G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 1.354
C2 0.000 0.000 -1.309
C3 0.000 1.184 0.623
C4 0.000 -1.184 0.623
N5 0.000 1.199 -0.716
N6 0.000 -1.199 -0.716
H7 0.000 0.000 2.438
H8 0.000 0.000 -2.395
H9 0.000 2.153 1.116
H10 0.000 -2.153 1.116

Atom - Atom Distances (Å)
  C1 C2 C3 C4 N5 N6 H7 H8 H9 H10
C12.66231.39111.39112.39152.39151.08433.74892.16622.1662
C22.66232.26532.26531.33731.33733.74661.08653.24233.2423
C31.39112.26532.36731.33862.73262.16703.24181.08753.3729
C41.39112.26532.36732.73261.33862.16703.24183.37291.0875
N52.39151.33731.33862.73262.39733.37382.06332.06513.8194
N62.39151.33732.73261.33862.39733.37382.06333.81942.0651
H71.08433.74662.16702.16703.37383.37384.83312.52682.5268
H83.74891.08653.24183.24182.06332.06334.83314.11844.1184
H92.16623.24231.08753.37292.06513.81942.52684.11844.3062
H102.16623.24233.37291.08753.81942.06512.52684.11844.3062

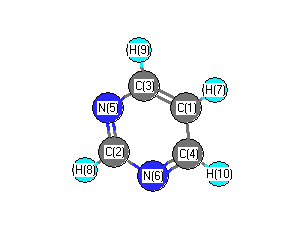
picture of 1,3-Diazine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C3 N5 122.337 C1 C3 H9 121.361
C1 C4 N6 122.337 C1 C4 H10 121.361
C2 N5 C3 115.677 C2 N6 C4 115.677
C3 C1 C4 116.611 C3 C1 H7 121.695
C4 C1 H7 121.695 N5 C2 N6 127.363
N5 C2 H8 116.318 N5 C3 H9 116.303
N6 C2 H8 116.318 N6 C4 H10 116.303
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability