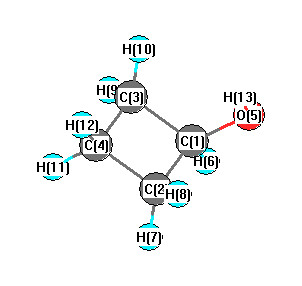
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3612 |
3553 |
3.59 |
|
|
|
| 2 |
A' |
3072 |
3021 |
53.49 |
|
|
|
| 3 |
A' |
3053 |
3003 |
21.90 |
|
|
|
| 4 |
A' |
3014 |
2965 |
24.12 |
|
|
|
| 5 |
A' |
3012 |
2963 |
26.39 |
|
|
|
| 6 |
A' |
2986 |
2937 |
17.53 |
|
|
|
| 7 |
A' |
1488 |
1464 |
6.22 |
|
|
|
| 8 |
A' |
1466 |
1442 |
6.38 |
|
|
|
| 9 |
A' |
1410 |
1386 |
65.69 |
|
|
|
| 10 |
A' |
1315 |
1293 |
0.99 |
|
|
|
| 11 |
A' |
1232 |
1212 |
22.00 |
|
|
|
| 12 |
A' |
1192 |
1172 |
0.06 |
|
|
|
| 13 |
A' |
1123 |
1105 |
114.25 |
|
|
|
| 14 |
A' |
1061 |
1044 |
45.12 |
|
|
|
| 15 |
A' |
971 |
955 |
11.79 |
|
|
|
| 16 |
A' |
889 |
874 |
0.79 |
|
|
|
| 17 |
A' |
741 |
729 |
3.03 |
|
|
|
| 18 |
A' |
591 |
581 |
2.49 |
|
|
|
| 19 |
A' |
443 |
436 |
4.32 |
|
|
|
| 20 |
A' |
180 |
177 |
1.71 |
|
|
|
| 21 |
A" |
3053 |
3003 |
38.85 |
|
|
|
| 22 |
A" |
2982 |
2932 |
38.27 |
|
|
|
| 23 |
A" |
1450 |
1426 |
1.00 |
|
|
|
| 24 |
A" |
1262 |
1241 |
0.71 |
|
|
|
| 25 |
A" |
1220 |
1200 |
0.71 |
|
|
|
| 26 |
A" |
1212 |
1192 |
0.00 |
|
|
|
| 27 |
A" |
1155 |
1136 |
0.00 |
|
|
|
| 28 |
A" |
1016 |
999 |
8.23 |
|
|
|
| 29 |
A" |
922 |
907 |
5.12 |
|
|
|
| 30 |
A" |
907 |
892 |
2.10 |
|
|
|
| 31 |
A" |
770 |
757 |
1.20 |
|
|
|
| 32 |
A" |
397 |
391 |
59.63 |
|
|
|
| 33 |
A" |
308 |
303 |
51.14 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24752.7 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 24344.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.104 |
|
|
|
| 2 |
C |
-0.324 |
|
|
|
| 3 |
C |
-0.324 |
|
|
|
| 4 |
C |
-0.311 |
|
|
|
| 5 |
O |
-0.577 |
|
|
|
| 6 |
H |
0.151 |
|
|
|
| 7 |
H |
0.156 |
|
|
|
| 8 |
H |
0.141 |
|
|
|
| 9 |
H |
0.156 |
|
|
|
| 10 |
H |
0.141 |
|
|
|
| 11 |
H |
0.159 |
|
|
|
| 12 |
H |
0.154 |
|
|
|
| 13 |
H |
0.375 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.385 |
-1.485 |
0.000 |
1.534 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-27.437 |
1.015 |
0.026 |
| y |
1.015 |
-36.017 |
-0.012 |
| z |
0.026 |
-0.012 |
-31.890 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
6.516 |
1.015 |
0.026 |
| y |
1.015 |
-6.353 |
-0.012 |
| z |
0.026 |
-0.012 |
-0.163 |
|
| Polar |
|---|
| 3z2-r2 | -0.326 |
|---|
| x2-y2 | 8.579 |
|---|
| xy | 1.015 |
|---|
| xz | 0.026 |
|---|
| yz | -0.012 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.776 |
-0.370 |
0.004 |
| y |
-0.370 |
6.750 |
-0.001 |
| z |
0.004 |
-0.001 |
6.544 |
<r2> (average value of r
2) Å
2
| <r2> |
116.669 |
| (<r2>)1/2 |
10.801 |