return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H6 (1,2,4,5-Hexatetraene)

using model chemistry: QCISD/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2H 1Ag
Energy calculated at QCISD/6-31G*
 hartrees
Energy at 0K-231.373022
Energy at 298.15K-231.376942
HF Energy-230.571092
Nuclear repulsion energy175.595626
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 Ag 3179 3027 0.00      
2 Ag 3164 3013 0.00      
3 Ag 2060 1962 0.00      
4 Ag 1541 1468 0.00      
5 Ag 1440 1371 0.00      
6 Ag 1197 1140 0.00      
7 Ag 1061 1011 0.00      
8 Ag 891 849 0.00      
9 Ag 538 512 0.00      
10 Ag 230 219 0.00      
11 Au 3239 3084 10.99      
12 Au 1047 997 0.32      
13 Au 921 877 32.95      
14 Au 487 463 4.24      
15 Au 340 324 9.26      
16 Au 75 71 0.23      
17 Bg 3239 3084 0.00      
18 Bg 1049 999 0.00      
19 Bg 901 858 0.00      
20 Bg 673 641 0.00      
21 Bg 318 303 0.00      
22 Bu 3185 3033 8.87      
23 Bu 3164 3013 18.25      
24 Bu 2069 1970 43.60      
25 Bu 1503 1431 0.32      
26 Bu 1306 1244 9.98      
27 Bu 1120 1066 1.00      
28 Bu 892 849 101.50      
29 Bu 537 511 19.31      
30 Bu 121 116 0.56      

Unscaled Zero Point Vibrational Energy (zpe) 20741.9 cm-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 19752.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G*
ABC
1.05712 0.04507 0.04401

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G*

Point Group is C2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.331 0.659 0.000
C2 0.331 -0.659 0.000
C3 0.331 1.801 0.000
C4 -0.331 -1.801 0.000
C5 0.996 2.935 0.000
C6 -0.996 -2.935 0.000
H7 -1.422 0.669 0.000
H8 1.422 -0.669 0.000
H9 1.283 3.428 0.928
H10 1.283 3.428 -0.928
H11 -1.283 -3.428 0.928
H12 -1.283 -3.428 -0.928

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12
C11.47431.32072.45962.63513.65461.09102.19963.33713.33714.29704.2970
C21.47432.45961.32073.65462.63512.19961.09104.29704.29703.33713.3371
C31.32072.45963.66251.31444.91862.08702.70062.10082.10085.55045.5504
C42.45961.32073.66254.91861.31442.70062.08705.55045.55042.10082.1008
C52.63513.65461.31444.91866.19893.31363.62971.08931.08936.82206.8220
C63.65462.63514.91861.31446.19893.62973.31366.82206.82201.08931.0893
H71.09102.19962.08702.70063.31363.62973.14383.97353.97354.20324.2032
H82.19961.09102.70062.08703.62973.31363.14384.20324.20323.97353.9735
H93.33714.29702.10085.55041.08936.82203.97354.20321.85637.31997.5516
H103.33714.29702.10085.55041.08936.82203.97354.20321.85637.55167.3199
H114.29703.33715.55042.10086.82201.08934.20323.97357.31997.55161.8563
H124.29703.33715.55042.10086.82201.08934.20323.97357.55167.31991.8563

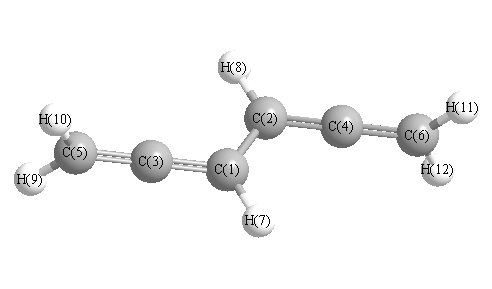
picture of 1,2,4,5-Hexatetraene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 123.187 C1 C2 H8 117.276
C1 C3 C5 179.743 C2 C1 C3 123.187
C2 C1 H7 117.276 C2 C4 C6 179.743
C3 C1 H7 119.537 C3 C5 H9 121.568
C3 C5 H10 121.568 C4 C2 H8 119.537
C4 C6 H11 121.568 C4 C6 H12 121.568
H9 C5 H10 116.864 H11 C6 H12 116.864
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability