return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H5NO (3-Methylisoxazole)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-285.060707
Energy at 298.15K-285.067133
HF Energy-284.772958
Nuclear repulsion energy222.800379
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3324 3156 0.58      
2 A 3304 3136 0.09      
3 A 3189 3027 6.04      
4 A 3155 2995 13.03      
5 A 3091 2934 14.74      
6 A 1628 1545 22.03      
7 A 1540 1462 11.16      
8 A 1532 1454 7.65      
9 A 1505 1428 8.09      
10 A 1469 1394 29.72      
11 A 1449 1375 7.28      
12 A 1312 1246 0.73      
13 A 1172 1112 16.70      
14 A 1100 1044 9.17      
15 A 1090 1034 2.38      
16 A 1049 995 1.81      
17 A 995 944 1.49      
18 A 938 891 1.80      
19 A 899 853 25.82      
20 A 851 808 0.85      
21 A 784 745 39.66      
22 A 663 630 6.01      
23 A 657 624 0.83      
24 A 610 579 4.62      
25 A 337 320 6.31      
26 A 271 258 4.44      
27 A 126 119 0.03      

Unscaled Zero Point Vibrational Energy (zpe) 19019.4 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 18053.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.30470 0.11707 0.08593

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 2.087 0.012 -0.001
H2 2.465 0.203 1.005
H3 2.475 -0.949 -0.341
H4 2.457 0.796 -0.664
C5 -1.446 0.607 -0.001
O6 -1.370 -0.744 0.002
N7 -0.080 -1.090 -0.002
C8 0.622 0.006 0.000
C9 -0.199 1.094 0.003
H10 0.078 2.097 -0.005
H11 -2.350 1.122 -0.005

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 O6 N7 C8 C9 H10 H11
C11.09111.09151.09113.58253.53882.43141.46472.52882.89494.5741
H21.09111.77171.77114.05854.07593.02722.10812.98253.21005.0056
H31.09151.77171.77504.23223.86582.58112.11213.38283.89075.2620
H41.09111.77111.77503.96294.17833.22972.10502.75422.79034.8632
C53.58254.05854.23223.96291.35332.17902.15391.33862.13071.0408
O63.53884.07593.86584.17831.35331.33602.12872.17943.18822.1075
N72.43143.02722.58113.22972.17901.33601.30172.18783.19103.1701
C81.46472.10812.11212.10502.15392.12871.30171.36352.16073.1755
C92.52882.98253.38282.75421.33862.17942.18781.36351.04002.1520
H102.89493.21003.89072.79032.13073.18823.19102.16071.04002.6169
H114.57415.00565.26204.86321.04082.10753.17013.17552.15202.6169

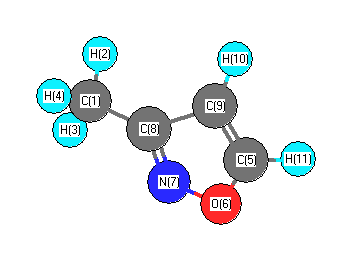
picture of 3-Methylisoxazole state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C8 N7 122.909 C1 C8 C9 126.767
H2 C1 H3 108.537 H2 C1 H4 108.504
H2 C1 C8 110.290 H3 C1 H4 108.831
H3 C1 C8 110.589 H4 C1 C8 110.036
C5 O6 N7 108.243 C5 C9 C8 105.705
C5 C9 H10 126.752 O6 C5 C9 108.118
O6 C5 H11 122.821 O6 N7 C8 107.608
N7 C8 C9 110.324 C8 C9 H10 127.538
C9 C5 H11 129.060
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.528      
2 H 0.179      
3 H 0.198      
4 H 0.179      
5 C 0.175      
6 O -0.382      
7 N -0.208      
8 C 0.313      
9 C -0.308      
10 H 0.183      
11 H 0.201      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 8.991 0.109 -0.000
y 0.109 7.120 0.000
z -0.000 0.000 3.837


<r2> (average value of r2) Å2
<r2> 137.138
(<r2>)1/2 11.711