return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H5NO (3-Methylisoxazole)

using model chemistry: MP3/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP3/6-31G*
 hartrees
Energy at 0K-284.503246
Energy at 298.15K-284.509723
HF Energy-283.629018
Nuclear repulsion energy223.319506
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP3/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3345 3140 0.32      
2 A 3323 3119 0.02      
3 A 3211 3014 5.26      
4 A 3182 2986 11.56      
5 A 3112 2921 12.15      
6 A 1693 1589 20.04      
7 A 1569 1473 9.27      
8 A 1548 1453 4.70      
9 A 1540 1446 8.59      
10 A 1508 1415 47.09      
11 A 1466 1376 1.32      
12 A 1336 1254 1.06      
13 A 1210 1136 17.93      
14 A 1120 1051 10.03      
15 A 1099 1031 1.84      
16 A 1069 1003 2.87      
17 A 1006 944 19.16      
18 A 1004 943 8.67      
19 A 939 882 10.91      
20 A 854 801 3.68      
21 A 779 731 41.08      
22 A 677 635 6.44      
23 A 653 613 0.44      
24 A 611 573 6.97      
25 A 343 322 6.45      
26 A 273 257 4.43      
27 A 123 115 0.08      

Unscaled Zero Point Vibrational Energy (zpe) 19296.5 cm-1
Scaled (by 0.9386) Zero Point Vibrational Energy (zpe) 18111.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP3/6-31G*
ABC
0.30562 0.11755 0.08627

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP3/6-31G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
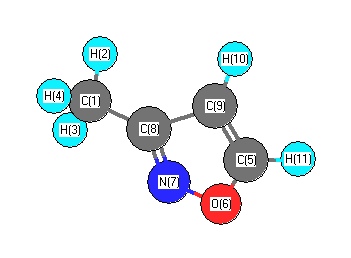
C1 -2.131 0.026 -0.000
H2 -2.520 0.539 -0.885
H3 -2.497 -1.002 -0.000
H4 -2.520 0.539 0.885
C5 1.483 0.575 0.000
O6 1.374 -0.769 -0.000
N7 0.027 -1.106 0.000
C8 -0.635 0.027 0.000
C9 0.257 1.147 -0.000
H10 0.009 2.198 0.000
H11 2.492 0.963 -0.000

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 O6 N7 C8 C9 H10 H11
C11.09421.09181.09423.65603.59402.43721.49652.63873.04944.7167
H21.09421.77761.76994.10004.20163.15852.14452.97753.15155.1065
H31.09181.77761.77754.28203.87842.52692.12813.49444.06535.3621
H41.09421.76991.77754.10004.20163.15842.14462.97773.15155.1066
C53.65604.10004.28204.10001.34812.22362.18791.35312.19261.0801
O63.59404.20163.87844.20161.34811.38802.16032.21763.26552.0606
N72.43723.15852.52693.15842.22361.38801.31172.26473.30373.2171
C81.49652.14452.12812.14462.18792.16031.31171.43242.26483.2634
C92.63872.97753.49442.97771.35312.21762.26471.43241.07962.2419
H103.04943.15154.06533.15152.19263.26553.30372.26481.07962.7728
H114.71675.10655.36215.10661.08012.06063.21713.26342.24192.7728

picture of 3-Methylisoxazole state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C8 N7 120.287 C1 C8 C9 128.549
H2 C1 H3 108.812 H2 C1 H4 107.951
H2 C1 C8 110.791 H3 C1 H4 108.810
H3 C1 C8 109.627 H4 C1 C8 110.792
C5 O6 N7 108.710 C5 C9 C8 103.495
C5 C9 H10 128.314 O6 C5 C9 110.368
O6 C5 H11 115.686 O6 N7 C8 106.263
N7 C8 C9 111.164 C8 C9 H10 128.191
C9 C5 H11 133.945
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability