Jump to
S1C2
Energy calculated at wB97X-D/6-31G*
| | hartrees |
|---|
| Energy at 0K | -697.502345 |
| Energy at 298.15K | -697.503594 |
| HF Energy | -697.502345 |
| Nuclear repulsion energy | |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at wB97X-D/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
873 |
828 |
95.87 |
|
|
|
| 2 |
A' |
620 |
588 |
12.07 |
|
|
|
| 3 |
A' |
362 |
343 |
14.54 |
|
|
|
| 4 |
A' |
180 |
171 |
7.34 |
|
|
|
| 5 |
A" |
773 |
733 |
406.68 |
|
|
|
| 6 |
A" |
455 |
432 |
2.35 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 1631.4 cm
-1
Scaled (by 0.9485) Zero Point Vibrational Energy (zpe) 1547.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at wB97X-D/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
-0.328 |
-0.197 |
0.000 |
| F2 |
1.238 |
0.054 |
0.000 |
| F3 |
-0.328 |
0.148 |
1.626 |
| F4 |
-0.328 |
0.148 |
-1.626 |
Atom - Atom Distances (Å)
| |
S1 |
F2 |
F3 |
F4 |
| S1 | | 1.5859 | 1.6622 | 1.6622 |
F2 | 1.5859 | | 2.2592 | 2.2592 | F3 | 1.6622 | 2.2592 | | 3.2517 | F4 | 1.6622 | 2.2592 | 3.2517 | |
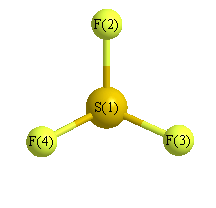 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| F2 |
S1 |
F3 |
88.110 |
|
F2 |
S1 |
F4 |
88.110 |
| F3 |
S1 |
F4 |
156.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at wB97X-D/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
1.050 |
|
|
|
| 2 |
F |
-0.302 |
|
|
|
| 3 |
F |
-0.374 |
|
|
|
| 4 |
F |
-0.374 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.683 |
-0.606 |
0.000 |
0.913 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.944 |
0.004 |
0.000 |
| y |
0.004 |
-25.181 |
0.000 |
| z |
0.000 |
0.000 |
-31.061 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.176 |
0.004 |
0.000 |
| y |
0.004 |
3.322 |
0.000 |
| z |
0.000 |
0.000 |
-5.498 |
|
| Polar |
|---|
| 3z2-r2 | -10.997 |
|---|
| x2-y2 | -0.764 |
|---|
| xy | 0.004 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.767 |
0.049 |
0.000 |
| y |
0.049 |
2.092 |
0.000 |
| z |
0.000 |
0.000 |
4.279 |
<r2> (average value of r
2) Å
2
| <r2> |
83.828 |
| (<r2>)1/2 |
9.156 |
Jump to
S1C1
Energy calculated at wB97X-D/6-31G*
| | hartrees |
|---|
| Energy at 0K | -697.500969 |
| Energy at 298.15K | |
| HF Energy | -697.500969 |
| Nuclear repulsion energy | |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at wB97X-D/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
881 |
835 |
96.99 |
|
|
|
| 2 |
A1 |
605 |
574 |
2.49 |
|
|
|
| 3 |
A1 |
369 |
350 |
14.28 |
|
|
|
| 4 |
B1 |
161i |
153i |
4.85 |
|
|
|
| 5 |
B2 |
792 |
752 |
470.15 |
|
|
|
| 6 |
B2 |
474 |
449 |
1.83 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 1479.5 cm
-1
Scaled (by 0.9485) Zero Point Vibrational Energy (zpe) 1403.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at wB97X-D/6-31G*
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
0.000 |
0.395 |
| F2 |
0.000 |
0.000 |
-1.195 |
| F3 |
0.000 |
1.669 |
0.247 |
| F4 |
0.000 |
-1.669 |
0.247 |
Atom - Atom Distances (Å)
| |
S1 |
F2 |
F3 |
F4 |
| S1 | | 1.5897 | 1.6759 | 1.6759 |
F2 | 1.5897 | | 2.2057 | 2.2057 | F3 | 1.6759 | 2.2057 | | 3.3387 | F4 | 1.6759 | 2.2057 | 3.3387 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| F2 |
S1 |
F3 |
84.932 |
|
F2 |
S1 |
F4 |
84.932 |
| F3 |
S1 |
F4 |
169.865 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at wB97X-D/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
1.066 |
|
|
|
| 2 |
F |
-0.294 |
|
|
|
| 3 |
F |
-0.386 |
|
|
|
| 4 |
F |
-0.386 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.755 |
0.755 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.882 |
0.000 |
0.000 |
| y |
0.000 |
-31.741 |
0.000 |
| z |
0.000 |
0.000 |
-25.872 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.925 |
0.000 |
0.000 |
| y |
0.000 |
-6.365 |
0.000 |
| z |
0.000 |
0.000 |
2.440 |
|
| Polar |
|---|
| 3z2-r2 | 4.879 |
|---|
| x2-y2 | 6.860 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.107 |
0.000 |
0.000 |
| y |
0.000 |
4.408 |
0.000 |
| z |
0.000 |
0.000 |
2.763 |
<r2> (average value of r
2) Å
2
| <r2> |
84.431 |
| (<r2>)1/2 |
9.189 |
