return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H6 (1-Methylcyclopropene)

using model chemistry: CID/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at CID/6-31G*
 hartrees
Energy at 0K-155.341421
Energy at 298.15K 
HF Energy-154.867710
Nuclear repulsion energy108.570214
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3404 3144 0.05      
2 A' 3239 2992 16.73      
3 A' 3157 2916 65.72      
4 A' 3148 2908 25.68      
5 A' 1968 1818 10.57      
6 A' 1619 1495 7.92      
7 A' 1573 1453 7.85      
8 A' 1497 1383 0.02      
9 A' 1282 1184 2.24      
10 A' 1164 1076 15.72      
11 A' 1148 1061 24.61      
12 A' 1038 959 5.98      
13 A' 1002 926 14.44      
14 A' 725 669 0.26      
15 A' 340 314 1.45      
16 A" 3219 2973 34.93      
17 A" 3217 2971 40.26      
18 A" 1567 1447 7.61      
19 A" 1185 1094 1.46      
20 A" 1108 1023 0.71      
21 A" 1043 963 1.93      
22 A" 760 702 34.03      
23 A" 311 287 11.29      
24 A" 155 143 0.41      

Unscaled Zero Point Vibrational Energy (zpe) 19432.7 cm-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 17950.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CID/6-31G*
ABC
0.69646 0.21294 0.17391

See section I.F.4 to change rotational constant units
Geometric Data calculated at CID/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.175 0.000
C2 -0.027 -1.324 0.000
C3 1.154 -0.399 0.000
C4 -0.855 1.381 0.000
H5 -0.298 -1.862 0.908
H6 -0.298 -1.862 -0.908
H7 2.226 -0.340 0.000
H8 -0.255 2.289 0.000
H9 -1.501 1.387 -0.879
H10 -1.501 1.387 0.879

Atom - Atom Distances (Å)
  C1 C2 C3 C4 H5 H6 H7 H8 H9 H10
C11.49961.28871.47822.24992.24992.28512.12952.12022.1202
C21.49961.50022.82881.08931.08932.45913.62073.20883.2088
C31.28871.50022.68392.25172.25171.07423.03513.31853.3185
C41.47822.82882.68393.41323.41323.52941.08891.09081.0908
H52.24991.08932.25173.41321.81603.08384.24953.89843.4649
H62.24991.08932.25173.41321.81603.08384.24953.46493.8984
H72.28512.45911.07423.52943.08383.08383.61534.20134.2013
H82.12953.62073.03511.08894.24954.24953.61531.77201.7720
H92.12023.20883.31851.09083.89843.46494.20131.77201.7576
H102.12023.20883.31851.09083.46493.89844.20131.77201.7576

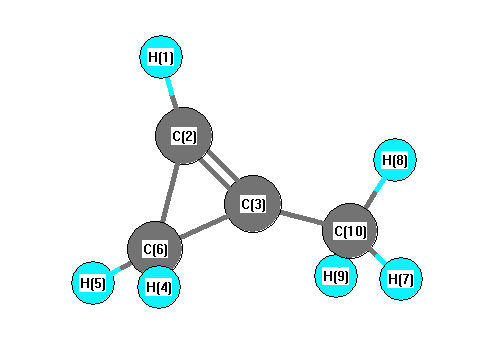
picture of 1-Methylcyclopropene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
H1 C2 C3 50.883 H1 C2 C6 119.867
C2 C3 C6 24.775 C2 C3 C10 72.666
C2 C6 C3 35.250 C2 C6 H4 49.450
C2 C6 H5 33.530 C3 C2 C6 119.975
C3 C6 H4 51.748 C3 C6 H5 66.218
C3 C10 H7 9.401 C3 C10 H8 65.160
C3 C10 H9 74.644 H4 C6 H5 74.571
C6 C3 C10 86.605 H7 C10 H8 58.765
H7 C10 H9 77.926 H8 C10 H9 60.268
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability